Jump to
S1C2
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -190.567557 |
| Energy at 298.15K | |
| HF Energy | -190.567557 |
| Nuclear repulsion energy | 87.513818 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3011 |
3011 |
46.03 |
265.53 |
0.11 |
0.20 |
| 2 |
A1 |
2264 |
2264 |
818.05 |
38.80 |
0.55 |
0.71 |
| 3 |
A1 |
1737 |
1737 |
34.22 |
4.08 |
0.74 |
0.85 |
| 4 |
A1 |
1439 |
1439 |
4.44 |
13.02 |
0.53 |
0.69 |
| 5 |
A1 |
904 |
904 |
7.61 |
50.84 |
0.16 |
0.28 |
| 6 |
B1 |
988 |
988 |
23.02 |
1.03 |
0.75 |
0.86 |
| 7 |
B1 |
593 |
593 |
14.05 |
0.06 |
0.75 |
0.86 |
| 8 |
B1 |
197 |
197 |
0.56 |
1.92 |
0.75 |
0.86 |
| 9 |
B2 |
3081 |
3081 |
14.30 |
173.80 |
0.75 |
0.86 |
| 10 |
B2 |
1028 |
1028 |
1.85 |
0.04 |
0.75 |
0.86 |
| 11 |
B2 |
401 |
401 |
3.52 |
2.42 |
0.75 |
0.86 |
| 12 |
B2 |
214i |
214i |
15.15 |
0.13 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7713.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7713.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.891 |
| C2 |
0.000 |
0.000 |
-0.562 |
| C3 |
0.000 |
0.000 |
0.726 |
| O4 |
0.000 |
0.000 |
1.915 |
| H5 |
0.000 |
0.932 |
-2.478 |
| H6 |
0.000 |
-0.932 |
-2.478 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3298 | 2.6172 | 3.8063 | 1.1011 | 1.1011 |
C2 | 1.3298 | | 1.2874 | 2.4765 | 2.1306 | 2.1306 | C3 | 2.6172 | 1.2874 | | 1.1891 | 3.3362 | 3.3362 | O4 | 3.8063 | 2.4765 | 1.1891 | | 4.4902 | 4.4902 | H5 | 1.1011 | 2.1306 | 3.3362 | 4.4902 | | 1.8644 | H6 | 1.1011 | 2.1306 | 3.3362 | 4.4902 | 1.8644 | |
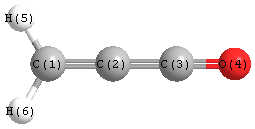 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.160 |
| C2 |
C1 |
H6 |
122.160 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.680 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.491 |
|
|
|
| 2 |
C |
0.361 |
|
|
|
| 3 |
C |
0.274 |
|
|
|
| 4 |
O |
-0.396 |
|
|
|
| 5 |
H |
-0.365 |
|
|
|
| 6 |
H |
-0.365 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.771 |
2.771 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.778 |
0.000 |
0.000 |
| y |
0.000 |
-22.688 |
0.000 |
| z |
0.000 |
0.000 |
-22.152 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.358 |
0.000 |
0.000 |
| y |
0.000 |
-0.224 |
0.000 |
| z |
0.000 |
0.000 |
0.581 |
|
| Polar |
|---|
| 3z2-r2 | 1.163 |
|---|
| x2-y2 | -0.090 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.799 |
0.000 |
0.000 |
| y |
0.000 |
4.355 |
0.000 |
| z |
0.000 |
0.000 |
11.687 |
<r2> (average value of r
2) Å
2
| <r2> |
83.943 |
| (<r2>)1/2 |
9.162 |
Jump to
S1C1
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -190.570334 |
| Energy at 298.15K | |
| HF Energy | -190.570334 |
| Nuclear repulsion energy | 87.837593 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3115 |
3115 |
10.92 |
128.09 |
0.70 |
0.83 |
| 2 |
A' |
3031 |
3031 |
38.80 |
193.42 |
0.12 |
0.21 |
| 3 |
A' |
2131 |
2131 |
794.60 |
49.74 |
0.55 |
0.71 |
| 4 |
A' |
1690 |
1690 |
6.65 |
5.20 |
0.21 |
0.35 |
| 5 |
A' |
1440 |
1440 |
1.24 |
13.02 |
0.58 |
0.73 |
| 6 |
A' |
1045 |
1045 |
20.57 |
4.38 |
0.03 |
0.06 |
| 7 |
A' |
931 |
931 |
3.10 |
39.69 |
0.10 |
0.18 |
| 8 |
A' |
492 |
492 |
8.97 |
4.78 |
0.73 |
0.85 |
| 9 |
A' |
196 |
196 |
19.26 |
6.69 |
0.69 |
0.82 |
| 10 |
A" |
995 |
995 |
23.61 |
1.39 |
0.75 |
0.86 |
| 11 |
A" |
691 |
691 |
7.06 |
0.17 |
0.75 |
0.86 |
| 12 |
A" |
269 |
269 |
2.05 |
2.40 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8012.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8012.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.160 |
-1.813 |
0.000 |
| C2 |
-0.371 |
-0.580 |
0.000 |
| C3 |
0.000 |
0.688 |
0.000 |
| O4 |
0.063 |
1.867 |
0.000 |
| H5 |
1.244 |
-2.002 |
0.000 |
| H6 |
-0.482 |
-2.704 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3426 | 2.5056 | 3.6813 | 1.1004 | 1.0981 |
C2 | 1.3426 | | 1.3205 | 2.4850 | 2.1517 | 2.1271 | C3 | 2.5056 | 1.3205 | | 1.1812 | 2.9630 | 3.4256 | O4 | 3.6813 | 2.4850 | 1.1812 | | 4.0450 | 4.6034 | H5 | 1.1004 | 2.1517 | 2.9630 | 4.0450 | | 1.8633 | H6 | 1.0981 | 2.1271 | 3.4256 | 4.6034 | 1.8633 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
140.394 |
|
C2 |
C1 |
H5 |
123.163 |
| C2 |
C1 |
H6 |
120.946 |
|
C2 |
C3 |
O4 |
166.732 |
| H5 |
C1 |
H6 |
115.891 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.508 |
|
|
|
| 2 |
C |
0.495 |
|
|
|
| 3 |
C |
0.180 |
|
|
|
| 4 |
O |
-0.337 |
|
|
|
| 5 |
H |
-0.445 |
|
|
|
| 6 |
H |
-0.400 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.929 |
-2.219 |
0.000 |
2.406 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.198 |
-0.963 |
0.000 |
| y |
-0.963 |
-22.084 |
0.000 |
| z |
0.000 |
0.000 |
-22.992 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.660 |
-0.963 |
0.000 |
| y |
-0.963 |
1.011 |
0.000 |
| z |
0.000 |
0.000 |
-0.352 |
|
| Polar |
|---|
| 3z2-r2 | -0.703 |
|---|
| x2-y2 | -1.114 |
|---|
| xy | -0.963 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.848 |
-0.027 |
0.000 |
| y |
-0.027 |
10.967 |
0.000 |
| z |
0.000 |
0.000 |
3.934 |
<r2> (average value of r
2) Å
2
| <r2> |
80.785 |
| (<r2>)1/2 |
8.988 |
