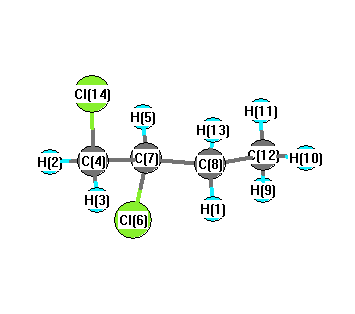
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3202 |
3069 |
1.21 |
|
|
|
| 2 |
A |
3156 |
3025 |
17.85 |
|
|
|
| 3 |
A |
3144 |
3013 |
32.00 |
|
|
|
| 4 |
A |
3131 |
3000 |
6.80 |
|
|
|
| 5 |
A |
3121 |
2991 |
1.51 |
|
|
|
| 6 |
A |
3110 |
2980 |
1.47 |
|
|
|
| 7 |
A |
3066 |
2938 |
16.96 |
|
|
|
| 8 |
A |
3062 |
2934 |
19.18 |
|
|
|
| 9 |
A |
1484 |
1423 |
7.79 |
|
|
|
| 10 |
A |
1476 |
1414 |
7.70 |
|
|
|
| 11 |
A |
1464 |
1403 |
5.10 |
|
|
|
| 12 |
A |
1458 |
1397 |
4.92 |
|
|
|
| 13 |
A |
1405 |
1346 |
2.54 |
|
|
|
| 14 |
A |
1393 |
1335 |
1.98 |
|
|
|
| 15 |
A |
1324 |
1269 |
4.47 |
|
|
|
| 16 |
A |
1311 |
1257 |
4.50 |
|
|
|
| 17 |
A |
1282 |
1228 |
5.99 |
|
|
|
| 18 |
A |
1234 |
1183 |
6.48 |
|
|
|
| 19 |
A |
1193 |
1144 |
15.11 |
|
|
|
| 20 |
A |
1143 |
1095 |
0.96 |
|
|
|
| 21 |
A |
1094 |
1048 |
1.45 |
|
|
|
| 22 |
A |
1071 |
1026 |
1.90 |
|
|
|
| 23 |
A |
1054 |
1010 |
3.13 |
|
|
|
| 24 |
A |
947 |
907 |
8.86 |
|
|
|
| 25 |
A |
841 |
806 |
5.69 |
|
|
|
| 26 |
A |
812 |
778 |
8.57 |
|
|
|
| 27 |
A |
749 |
718 |
57.53 |
|
|
|
| 28 |
A |
669 |
641 |
36.00 |
|
|
|
| 29 |
A |
463 |
444 |
1.74 |
|
|
|
| 30 |
A |
386 |
370 |
1.58 |
|
|
|
| 31 |
A |
298 |
285 |
0.06 |
|
|
|
| 32 |
A |
238 |
228 |
0.40 |
|
|
|
| 33 |
A |
216 |
207 |
3.02 |
|
|
|
| 34 |
A |
187 |
179 |
3.35 |
|
|
|
| 35 |
A |
112 |
107 |
3.50 |
|
|
|
| 36 |
A |
90 |
87 |
1.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25192.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 24141.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.243 |
|
|
|
| 2 |
H |
-0.048 |
|
|
|
| 3 |
H |
-0.019 |
|
|
|
| 4 |
C |
0.309 |
|
|
|
| 5 |
H |
-0.285 |
|
|
|
| 6 |
Cl |
-0.124 |
|
|
|
| 7 |
C |
-0.140 |
|
|
|
| 8 |
C |
0.738 |
|
|
|
| 9 |
H |
-0.112 |
|
|
|
| 10 |
H |
-0.117 |
|
|
|
| 11 |
H |
-0.109 |
|
|
|
| 12 |
C |
0.445 |
|
|
|
| 13 |
H |
-0.231 |
|
|
|
| 14 |
Cl |
-0.064 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.295 |
0.670 |
0.252 |
0.775 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-57.645 |
3.313 |
-0.205 |
| y |
3.313 |
-51.175 |
-0.177 |
| z |
-0.205 |
-0.177 |
-50.714 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.701 |
3.313 |
-0.205 |
| y |
3.313 |
3.005 |
-0.177 |
| z |
-0.205 |
-0.177 |
3.696 |
|
| Polar |
|---|
| 3z2-r2 | 7.392 |
|---|
| x2-y2 | -6.470 |
|---|
| xy | 3.313 |
|---|
| xz | -0.205 |
|---|
| yz | -0.177 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.774 |
-0.715 |
0.061 |
| y |
-0.715 |
11.943 |
-0.120 |
| z |
0.061 |
-0.120 |
9.626 |
<r2> (average value of r
2) Å
2
| <r2> |
321.975 |
| (<r2>)1/2 |
17.944 |