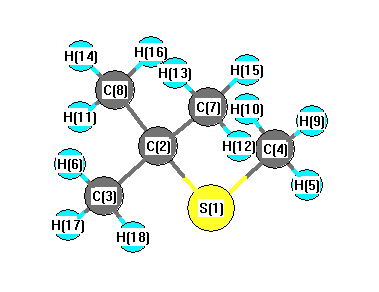
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3167 |
3035 |
7.91 |
|
|
|
| 2 |
A |
3153 |
3022 |
27.79 |
|
|
|
| 3 |
A |
3135 |
3005 |
31.59 |
|
|
|
| 4 |
A |
3127 |
2997 |
35.85 |
|
|
|
| 5 |
A |
3061 |
2933 |
28.53 |
|
|
|
| 6 |
A |
3055 |
2927 |
38.04 |
|
|
|
| 7 |
A |
3049 |
2922 |
24.41 |
|
|
|
| 8 |
A |
1500 |
1437 |
8.76 |
|
|
|
| 9 |
A |
1484 |
1422 |
9.41 |
|
|
|
| 10 |
A |
1468 |
1407 |
0.63 |
|
|
|
| 11 |
A |
1451 |
1390 |
9.25 |
|
|
|
| 12 |
A |
1413 |
1355 |
4.16 |
|
|
|
| 13 |
A |
1387 |
1329 |
11.25 |
|
|
|
| 14 |
A |
1325 |
1270 |
1.26 |
|
|
|
| 15 |
A |
1253 |
1200 |
0.42 |
|
|
|
| 16 |
A |
1191 |
1141 |
44.01 |
|
|
|
| 17 |
A |
1044 |
1000 |
2.61 |
|
|
|
| 18 |
A |
959 |
919 |
6.59 |
|
|
|
| 19 |
A |
947 |
908 |
0.94 |
|
|
|
| 20 |
A |
835 |
800 |
0.24 |
|
|
|
| 21 |
A |
741 |
710 |
0.51 |
|
|
|
| 22 |
A |
593 |
569 |
3.59 |
|
|
|
| 23 |
A |
423 |
406 |
0.01 |
|
|
|
| 24 |
A |
366 |
350 |
0.66 |
|
|
|
| 25 |
A |
320 |
307 |
0.01 |
|
|
|
| 26 |
A |
277 |
265 |
0.02 |
|
|
|
| 27 |
A |
219 |
210 |
0.47 |
|
|
|
| 28 |
A |
3156 |
3024 |
3.10 |
|
|
|
| 29 |
A |
3151 |
3020 |
29.61 |
|
|
|
| 30 |
A |
3146 |
3015 |
6.60 |
|
|
|
| 31 |
A |
3123 |
2993 |
2.29 |
|
|
|
| 32 |
A |
3046 |
2919 |
20.67 |
|
|
|
| 33 |
A |
1485 |
1423 |
3.85 |
|
|
|
| 34 |
A |
1466 |
1405 |
1.90 |
|
|
|
| 35 |
A |
1458 |
1397 |
1.79 |
|
|
|
| 36 |
A |
1440 |
1380 |
6.69 |
|
|
|
| 37 |
A |
1386 |
1328 |
13.84 |
|
|
|
| 38 |
A |
1247 |
1195 |
2.33 |
|
|
|
| 39 |
A |
1037 |
994 |
0.20 |
|
|
|
| 40 |
A |
958 |
918 |
0.63 |
|
|
|
| 41 |
A |
955 |
915 |
1.35 |
|
|
|
| 42 |
A |
939 |
900 |
0.23 |
|
|
|
| 43 |
A |
404 |
387 |
0.30 |
|
|
|
| 44 |
A |
305 |
293 |
0.59 |
|
|
|
| 45 |
A |
274 |
262 |
0.08 |
|
|
|
| 46 |
A |
224 |
215 |
0.07 |
|
|
|
| 47 |
A |
166 |
159 |
0.93 |
|
|
|
| 48 |
A |
43 |
41 |
1.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35173.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 33707.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.204 |
|
|
|
| 2 |
C |
-2.100 |
|
|
|
| 3 |
C |
1.139 |
|
|
|
| 4 |
C |
0.227 |
|
|
|
| 5 |
H |
-0.176 |
|
|
|
| 6 |
H |
-0.092 |
|
|
|
| 7 |
C |
1.055 |
|
|
|
| 8 |
C |
1.055 |
|
|
|
| 9 |
H |
-0.097 |
|
|
|
| 10 |
H |
-0.097 |
|
|
|
| 11 |
H |
-0.123 |
|
|
|
| 12 |
H |
-0.123 |
|
|
|
| 13 |
H |
-0.090 |
|
|
|
| 14 |
H |
-0.090 |
|
|
|
| 15 |
H |
-0.226 |
|
|
|
| 16 |
H |
-0.226 |
|
|
|
| 17 |
H |
-0.120 |
|
|
|
| 18 |
H |
-0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.516 |
0.704 |
0.000 |
1.672 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.364 |
-2.714 |
0.000 |
| y |
-2.714 |
-45.044 |
0.000 |
| z |
0.000 |
0.000 |
-48.565 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.559 |
-2.714 |
0.000 |
| y |
-2.714 |
3.421 |
0.000 |
| z |
0.000 |
0.000 |
-1.862 |
|
| Polar |
|---|
| 3z2-r2 | -3.723 |
|---|
| x2-y2 | -3.320 |
|---|
| xy | -2.714 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.339 |
-0.464 |
0.000 |
| y |
-0.464 |
14.385 |
0.000 |
| z |
0.000 |
0.000 |
11.549 |
<r2> (average value of r
2) Å
2
| <r2> |
225.836 |
| (<r2>)1/2 |
15.028 |