Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -277.717152 |
| Energy at 298.15K | |
| HF Energy | -276.961805 |
| Nuclear repulsion energy | 123.941685 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3084 |
3013 |
|
|
|
|
| 2 |
Ag |
1569 |
1533 |
|
|
|
|
| 3 |
Ag |
1491 |
1456 |
|
|
|
|
| 4 |
Ag |
1121 |
1095 |
|
|
|
|
| 5 |
Ag |
1084 |
1059 |
|
|
|
|
| 6 |
Ag |
461 |
450 |
|
|
|
|
| 7 |
Au |
3158 |
3085 |
|
|
|
|
| 8 |
Au |
1246 |
1217 |
|
|
|
|
| 9 |
Au |
827 |
808 |
|
|
|
|
| 10 |
Au |
137 |
134 |
|
|
|
|
| 11 |
Bg |
3135 |
3062 |
|
|
|
|
| 12 |
Bg |
1317 |
1287 |
|
|
|
|
| 13 |
Bg |
1203 |
1176 |
|
|
|
|
| 14 |
Bu |
3088 |
3017 |
|
|
|
|
| 15 |
Bu |
1577 |
1541 |
|
|
|
|
| 16 |
Bu |
1379 |
1347 |
|
|
|
|
| 17 |
Bu |
1094 |
1069 |
|
|
|
|
| 18 |
Bu |
274 |
267 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13622.2 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 13307.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.434 |
0.626 |
0.000 |
| C2 |
-0.434 |
-0.626 |
0.000 |
| F3 |
-0.434 |
1.746 |
0.000 |
| F4 |
0.434 |
-1.746 |
0.000 |
| H5 |
1.062 |
0.681 |
0.905 |
| H6 |
1.062 |
0.681 |
-0.905 |
| H7 |
-1.062 |
-0.681 |
0.905 |
| H8 |
-1.062 |
-0.681 |
-0.905 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5248 | 1.4173 | 2.3726 | 1.1029 | 1.1029 | 2.1839 | 2.1839 |
C2 | 1.5248 | | 2.3726 | 1.4173 | 2.1839 | 2.1839 | 1.1029 | 1.1029 | F3 | 1.4173 | 2.3726 | | 3.5987 | 2.0477 | 2.0477 | 2.6656 | 2.6656 | F4 | 2.3726 | 1.4173 | 3.5987 | | 2.6656 | 2.6656 | 2.0477 | 2.0477 | H5 | 1.1029 | 2.1839 | 2.0477 | 2.6656 | | 1.8106 | 2.5236 | 3.1059 | H6 | 1.1029 | 2.1839 | 2.0477 | 2.6656 | 1.8106 | | 3.1059 | 2.5236 | H7 | 2.1839 | 1.1029 | 2.6656 | 2.0477 | 2.5236 | 3.1059 | | 1.8106 | H8 | 2.1839 | 1.1029 | 2.6656 | 2.0477 | 3.1059 | 2.5236 | 1.8106 | |
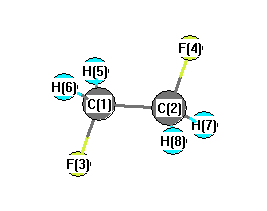 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.442 |
|
C1 |
C2 |
H7 |
111.414 |
| C1 |
C2 |
H8 |
111.414 |
|
C2 |
C1 |
F3 |
107.442 |
| C2 |
C1 |
H5 |
111.414 |
|
C2 |
C1 |
H6 |
111.414 |
| F3 |
C1 |
H5 |
108.039 |
|
F3 |
C1 |
H6 |
108.039 |
| F4 |
C2 |
H7 |
108.039 |
|
F4 |
C2 |
H8 |
108.039 |
| H5 |
C1 |
H6 |
110.330 |
|
H7 |
C2 |
H8 |
110.330 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -277.718189 |
| Energy at 298.15K | |
| HF Energy | -276.962359 |
| Nuclear repulsion energy | 126.124316 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3125 |
3053 |
|
|
|
|
| 2 |
A |
3073 |
3002 |
|
|
|
|
| 3 |
A |
1550 |
1515 |
|
|
|
|
| 4 |
A |
1477 |
1443 |
|
|
|
|
| 5 |
A |
1314 |
1284 |
|
|
|
|
| 6 |
A |
1167 |
1140 |
|
|
|
|
| 7 |
A |
1139 |
1113 |
|
|
|
|
| 8 |
A |
901 |
881 |
|
|
|
|
| 9 |
A |
329 |
321 |
|
|
|
|
| 10 |
A |
148 |
145 |
|
|
|
|
| 11 |
B |
3138 |
3065 |
|
|
|
|
| 12 |
B |
3062 |
2991 |
|
|
|
|
| 13 |
B |
1548 |
1512 |
|
|
|
|
| 14 |
B |
1442 |
1409 |
|
|
|
|
| 15 |
B |
1271 |
1242 |
|
|
|
|
| 16 |
B |
1110 |
1085 |
|
|
|
|
| 17 |
B |
925 |
904 |
|
|
|
|
| 18 |
B |
502 |
491 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13611.6 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 13297.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.271 |
0.707 |
0.525 |
| C2 |
-0.271 |
-0.707 |
0.525 |
| F3 |
-0.271 |
1.434 |
-0.558 |
| F4 |
0.271 |
-1.434 |
-0.558 |
| H5 |
-0.024 |
1.223 |
1.457 |
| H6 |
1.369 |
0.711 |
0.414 |
| H7 |
0.024 |
-1.223 |
1.457 |
| H8 |
-1.369 |
-0.711 |
0.414 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5137 | 1.4131 | 2.3993 | 1.1058 | 1.1037 | 2.1569 | 2.1709 |
C2 | 1.5137 | | 2.3993 | 1.4131 | 2.1569 | 2.1709 | 1.1058 | 1.1037 | F3 | 1.4131 | 2.3993 | | 2.9194 | 2.0416 | 2.0395 | 3.3480 | 2.5988 | F4 | 2.3993 | 1.4131 | 2.9194 | | 3.3480 | 2.5988 | 2.0416 | 2.0395 | H5 | 1.1058 | 2.1569 | 2.0416 | 3.3480 | | 1.8142 | 2.4458 | 2.5761 | H6 | 1.1037 | 2.1709 | 2.0395 | 2.5988 | 1.8142 | | 2.5761 | 3.0857 | H7 | 2.1569 | 1.1058 | 3.3480 | 2.0416 | 2.4458 | 2.5761 | | 1.8142 | H8 | 2.1709 | 1.1037 | 2.5988 | 2.0395 | 2.5761 | 3.0857 | 1.8142 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.075 |
|
C1 |
C2 |
H7 |
109.877 |
| C1 |
C2 |
H8 |
111.114 |
|
C2 |
C1 |
F3 |
110.075 |
| C2 |
C1 |
H5 |
109.877 |
|
C2 |
C1 |
H6 |
111.114 |
| F3 |
C1 |
H5 |
107.667 |
|
F3 |
C1 |
H6 |
107.627 |
| F4 |
C2 |
H7 |
107.667 |
|
F4 |
C2 |
H8 |
107.627 |
| H5 |
C1 |
H6 |
110.395 |
|
H7 |
C2 |
H8 |
110.395 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
