return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSH (Hydrogen methyl disulfide)

using model chemistry: QCISD(T)=FULL/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD(T)=FULL/aug-cc-pVDZ
 hartrees
Energy at 0K-835.786922
Energy at 298.15K 
HF Energy-835.265542
Nuclear repulsion energy146.690820
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3149 3055        
2 A 3133 3039        
3 A 3034 2943        
4 A 2651 2571        
5 A 1455 1411        
6 A 1439 1396        
7 A 1326 1286        
8 A 963 934        
9 A 959 931        
10 A 865 839        
11 A 697 676        
12 A 487 472        
13 A 308 299        
14 A 235 228        
15 A 168 163        

Unscaled Zero Point Vibrational Energy (zpe) 10432.8 cm-1
Scaled (by 0.9701) Zero Point Vibrational Energy (zpe) 10120.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)=FULL/aug-cc-pVDZ
ABC
0.52817 0.14350 0.11824

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)=FULL/aug-cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.633 0.722 -0.004
S2 -0.503 -0.726 0.012
S3 1.369 0.244 -0.087
H4 1.554 0.410 1.249
H5 -1.452 1.332 -0.902
H6 -2.659 0.314 -0.033
H7 -1.501 1.325 0.908

Atom - Atom Distances (Å)
  C1 S2 S3 H4 H5 H6 H7
C11.83673.04153.43901.10071.10421.1011
S21.83672.11162.65602.44392.39392.4504
S33.04152.11161.35913.13184.02933.2245
H43.43902.65601.35913.80974.40483.2075
H51.10072.44393.13183.80971.80231.8103
H61.10422.39394.02934.40481.80231.8020
H71.10112.45043.22453.20751.81031.8020

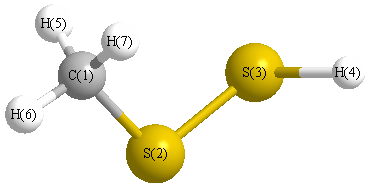
picture of Hydrogen methyl disulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 S3 100.534 S2 C1 H5 110.071
S2 C1 H6 106.294 S2 C1 H7 110.529
S2 S3 H4 97.488 H5 C1 H6 109.654
H5 C1 H7 110.605 H6 C1 H7 109.593
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability