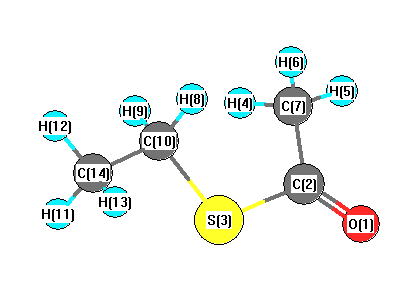
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3189 |
3056 |
6.53 |
|
|
|
| 2 |
A |
3162 |
3030 |
21.28 |
|
|
|
| 3 |
A |
3150 |
3019 |
3.66 |
|
|
|
| 4 |
A |
3148 |
3016 |
17.45 |
|
|
|
| 5 |
A |
3139 |
3008 |
0.07 |
|
|
|
| 6 |
A |
3085 |
2957 |
16.84 |
|
|
|
| 7 |
A |
3068 |
2940 |
1.31 |
|
|
|
| 8 |
A |
3064 |
2936 |
22.89 |
|
|
|
| 9 |
A |
1788 |
1714 |
356.23 |
|
|
|
| 10 |
A |
1480 |
1418 |
3.35 |
|
|
|
| 11 |
A |
1471 |
1410 |
10.60 |
|
|
|
| 12 |
A |
1459 |
1399 |
0.97 |
|
|
|
| 13 |
A |
1450 |
1390 |
13.79 |
|
|
|
| 14 |
A |
1436 |
1376 |
19.11 |
|
|
|
| 15 |
A |
1396 |
1338 |
5.13 |
|
|
|
| 16 |
A |
1360 |
1304 |
27.21 |
|
|
|
| 17 |
A |
1284 |
1230 |
18.00 |
|
|
|
| 18 |
A |
1263 |
1210 |
0.01 |
|
|
|
| 19 |
A |
1156 |
1108 |
170.02 |
|
|
|
| 20 |
A |
1082 |
1037 |
5.03 |
|
|
|
| 21 |
A |
1036 |
993 |
0.00 |
|
|
|
| 22 |
A |
1008 |
966 |
1.24 |
|
|
|
| 23 |
A |
1004 |
962 |
3.46 |
|
|
|
| 24 |
A |
954 |
914 |
32.97 |
|
|
|
| 25 |
A |
787 |
754 |
3.91 |
|
|
|
| 26 |
A |
704 |
675 |
0.24 |
|
|
|
| 27 |
A |
651 |
624 |
65.76 |
|
|
|
| 28 |
A |
517 |
495 |
1.48 |
|
|
|
| 29 |
A |
449 |
431 |
3.22 |
|
|
|
| 30 |
A |
376 |
360 |
2.22 |
|
|
|
| 31 |
A |
304 |
291 |
0.95 |
|
|
|
| 32 |
A |
250 |
239 |
0.08 |
|
|
|
| 33 |
A |
159 |
153 |
1.05 |
|
|
|
| 34 |
A |
148 |
142 |
0.02 |
|
|
|
| 35 |
A |
89 |
85 |
0.45 |
|
|
|
| 36 |
A |
52 |
50 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25057.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 24012.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.558 |
|
|
|
| 2 |
C |
0.200 |
|
|
|
| 3 |
S |
0.351 |
|
|
|
| 4 |
H |
0.075 |
|
|
|
| 5 |
H |
-0.292 |
|
|
|
| 6 |
H |
0.074 |
|
|
|
| 7 |
C |
0.218 |
|
|
|
| 8 |
H |
-0.186 |
|
|
|
| 9 |
H |
-0.186 |
|
|
|
| 10 |
C |
0.233 |
|
|
|
| 11 |
H |
-0.095 |
|
|
|
| 12 |
H |
-0.107 |
|
|
|
| 13 |
H |
-0.095 |
|
|
|
| 14 |
C |
0.368 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.857 |
3.507 |
-0.000 |
4.524 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.339 |
4.193 |
0.001 |
| y |
4.193 |
-45.011 |
0.000 |
| z |
0.001 |
0.000 |
-44.285 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.691 |
4.193 |
0.001 |
| y |
4.193 |
0.801 |
0.000 |
| z |
0.001 |
0.000 |
1.890 |
|
| Polar |
|---|
| 3z2-r2 | 3.781 |
|---|
| x2-y2 | -2.328 |
|---|
| xy | 4.193 |
|---|
| xz | 0.001 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.850 |
-0.022 |
0.000 |
| y |
-0.022 |
10.781 |
-0.000 |
| z |
0.000 |
-0.000 |
8.337 |
<r2> (average value of r
2) Å
2
| <r2> |
251.097 |
| (<r2>)1/2 |
15.846 |