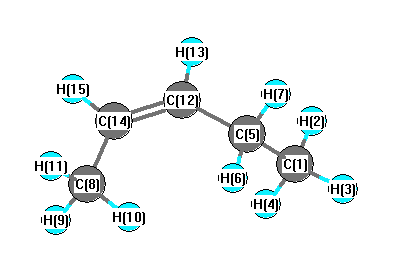
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3178 |
3046 |
46.70 |
|
|
|
| 2 |
A |
3156 |
3025 |
13.03 |
|
|
|
| 3 |
A |
3148 |
3017 |
15.27 |
|
|
|
| 4 |
A |
3141 |
3010 |
33.02 |
|
|
|
| 5 |
A |
3136 |
3005 |
29.55 |
|
|
|
| 6 |
A |
3108 |
2978 |
4.73 |
|
|
|
| 7 |
A |
3104 |
2974 |
16.94 |
|
|
|
| 8 |
A |
3053 |
2926 |
33.73 |
|
|
|
| 9 |
A |
3045 |
2918 |
30.04 |
|
|
|
| 10 |
A |
3041 |
2914 |
27.73 |
|
|
|
| 11 |
A |
1753 |
1680 |
4.72 |
|
|
|
| 12 |
A |
1483 |
1421 |
6.33 |
|
|
|
| 13 |
A |
1475 |
1414 |
7.51 |
|
|
|
| 14 |
A |
1471 |
1409 |
5.01 |
|
|
|
| 15 |
A |
1459 |
1398 |
2.88 |
|
|
|
| 16 |
A |
1454 |
1394 |
3.98 |
|
|
|
| 17 |
A |
1432 |
1372 |
8.92 |
|
|
|
| 18 |
A |
1390 |
1332 |
2.79 |
|
|
|
| 19 |
A |
1384 |
1326 |
1.83 |
|
|
|
| 20 |
A |
1320 |
1264 |
4.43 |
|
|
|
| 21 |
A |
1295 |
1241 |
0.14 |
|
|
|
| 22 |
A |
1269 |
1216 |
0.21 |
|
|
|
| 23 |
A |
1167 |
1118 |
0.19 |
|
|
|
| 24 |
A |
1090 |
1044 |
4.16 |
|
|
|
| 25 |
A |
1054 |
1010 |
3.78 |
|
|
|
| 26 |
A |
1048 |
1004 |
1.90 |
|
|
|
| 27 |
A |
1044 |
1000 |
0.46 |
|
|
|
| 28 |
A |
1001 |
959 |
0.43 |
|
|
|
| 29 |
A |
949 |
910 |
13.00 |
|
|
|
| 30 |
A |
876 |
839 |
1.33 |
|
|
|
| 31 |
A |
787 |
754 |
4.30 |
|
|
|
| 32 |
A |
713 |
684 |
35.27 |
|
|
|
| 33 |
A |
571 |
547 |
6.41 |
|
|
|
| 34 |
A |
469 |
449 |
1.44 |
|
|
|
| 35 |
A |
299 |
286 |
0.02 |
|
|
|
| 36 |
A |
255 |
245 |
0.05 |
|
|
|
| 37 |
A |
215 |
206 |
0.01 |
|
|
|
| 38 |
A |
125 |
120 |
0.41 |
|
|
|
| 39 |
A |
44 |
43 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29999.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28748.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.315 |
|
|
|
| 2 |
H |
-0.112 |
|
|
|
| 3 |
H |
-0.154 |
|
|
|
| 4 |
H |
-0.130 |
|
|
|
| 5 |
C |
0.332 |
|
|
|
| 6 |
H |
-0.239 |
|
|
|
| 7 |
H |
-0.102 |
|
|
|
| 8 |
C |
0.130 |
|
|
|
| 9 |
H |
0.022 |
|
|
|
| 10 |
H |
-0.301 |
|
|
|
| 11 |
H |
0.015 |
|
|
|
| 12 |
C |
0.991 |
|
|
|
| 13 |
H |
-0.625 |
|
|
|
| 14 |
C |
0.426 |
|
|
|
| 15 |
H |
-0.568 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.010 |
-0.297 |
0.072 |
0.306 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.124 |
0.318 |
0.915 |
| y |
0.318 |
-32.809 |
-0.280 |
| z |
0.915 |
-0.280 |
-34.676 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.618 |
0.318 |
0.915 |
| y |
0.318 |
0.592 |
-0.280 |
| z |
0.915 |
-0.280 |
-2.210 |
|
| Polar |
|---|
| 3z2-r2 | -4.420 |
|---|
| x2-y2 | 0.684 |
|---|
| xy | 0.318 |
|---|
| xz | 0.915 |
|---|
| yz | -0.280 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.493 |
0.288 |
0.381 |
| y |
0.288 |
8.836 |
-0.075 |
| z |
0.381 |
-0.075 |
8.038 |
<r2> (average value of r
2) Å
2
| <r2> |
165.935 |
| (<r2>)1/2 |
12.882 |