Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -554.172624 |
| Energy at 298.15K | -554.177691 |
| Nuclear repulsion energy | 198.342313 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3278 |
3141 |
6.41 |
|
|
|
| 2 |
A |
3200 |
3067 |
4.05 |
|
|
|
| 3 |
A |
3178 |
3045 |
3.06 |
|
|
|
| 4 |
A |
1680 |
1610 |
5.38 |
|
|
|
| 5 |
A |
1416 |
1357 |
9.05 |
|
|
|
| 6 |
A |
1305 |
1251 |
0.24 |
|
|
|
| 7 |
A |
1055 |
1011 |
0.15 |
|
|
|
| 8 |
A |
975 |
934 |
0.00 |
|
|
|
| 9 |
A |
911 |
873 |
0.00 |
|
|
|
| 10 |
A |
731 |
700 |
0.62 |
|
|
|
| 11 |
A |
623 |
597 |
0.00 |
|
|
|
| 12 |
A |
408 |
391 |
0.53 |
|
|
|
| 13 |
A |
174 |
167 |
0.17 |
|
|
|
| 14 |
A |
60 |
57 |
0.00 |
|
|
|
| 15 |
A |
3278 |
3141 |
0.43 |
|
|
|
| 16 |
A |
3202 |
3068 |
1.33 |
|
|
|
| 17 |
A |
3178 |
3046 |
0.02 |
|
|
|
| 18 |
A |
1656 |
1587 |
186.25 |
|
|
|
| 19 |
A |
1410 |
1351 |
6.71 |
|
|
|
| 20 |
A |
1272 |
1219 |
14.34 |
|
|
|
| 21 |
A |
1022 |
979 |
28.23 |
|
|
|
| 22 |
A |
980 |
939 |
44.46 |
|
|
|
| 23 |
A |
912 |
874 |
82.09 |
|
|
|
| 24 |
A |
748 |
717 |
47.50 |
|
|
|
| 25 |
A |
561 |
537 |
25.54 |
|
|
|
| 26 |
A |
388 |
372 |
4.92 |
|
|
|
| 27 |
A |
19 |
18 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18809.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 18025.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.622 |
| C2 |
0.000 |
1.364 |
-0.491 |
| C3 |
0.000 |
-1.364 |
-0.491 |
| C4 |
0.001 |
2.641 |
-0.099 |
| C5 |
-0.001 |
-2.641 |
-0.099 |
| H6 |
-0.001 |
1.096 |
-1.548 |
| H7 |
0.001 |
-1.096 |
-1.548 |
| H8 |
0.000 |
3.437 |
-0.841 |
| H9 |
0.002 |
2.931 |
0.950 |
| H10 |
-0.000 |
-3.437 |
-0.841 |
| H11 |
-0.002 |
-2.931 |
0.950 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7609 | 1.7609 | 2.7374 | 2.7374 | 2.4316 | 2.4316 | 3.7352 | 2.9492 | 3.7352 | 2.9492 |
C2 | 1.7609 | | 2.7287 | 1.3350 | 4.0241 | 1.0910 | 2.6778 | 2.1017 | 2.1287 | 4.8138 | 4.5306 | C3 | 1.7609 | 2.7287 | | 4.0241 | 1.3350 | 2.6778 | 1.0910 | 4.8138 | 4.5306 | 2.1017 | 2.1287 | C4 | 2.7374 | 1.3350 | 4.0241 | | 5.2813 | 2.1181 | 4.0076 | 1.0879 | 1.0890 | 6.1225 | 5.6695 | C5 | 2.7374 | 4.0241 | 1.3350 | 5.2813 | | 4.0076 | 2.1181 | 6.1225 | 5.6695 | 1.0879 | 1.0890 | H6 | 2.4316 | 1.0910 | 2.6778 | 2.1181 | 4.0076 | | 2.1917 | 2.4455 | 3.1001 | 4.5875 | 4.7390 | H7 | 2.4316 | 2.6778 | 1.0910 | 4.0076 | 2.1181 | 2.1917 | | 4.5875 | 4.7390 | 2.4455 | 3.1001 | H8 | 3.7352 | 2.1017 | 4.8138 | 1.0879 | 6.1225 | 2.4455 | 4.5875 | | 1.8611 | 6.8735 | 6.6147 | H9 | 2.9492 | 2.1287 | 4.5306 | 1.0890 | 5.6695 | 3.1001 | 4.7390 | 1.8611 | | 6.6147 | 5.8618 | H10 | 3.7352 | 4.8138 | 2.1017 | 6.1225 | 1.0879 | 4.5875 | 2.4455 | 6.8735 | 6.6147 | | 1.8611 | H11 | 2.9492 | 4.5306 | 2.1287 | 5.6695 | 1.0890 | 4.7390 | 3.1001 | 6.6147 | 5.8618 | 1.8611 | |
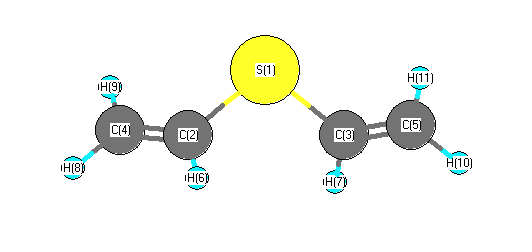 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.728 |
|
S1 |
C2 |
H6 |
114.964 |
| S1 |
C3 |
C5 |
123.728 |
|
S1 |
C3 |
H7 |
114.964 |
| C2 |
S1 |
C3 |
101.578 |
|
C2 |
C4 |
H8 |
119.976 |
| C2 |
C4 |
H9 |
122.519 |
|
C3 |
C5 |
H10 |
119.976 |
| C3 |
C5 |
H11 |
122.519 |
|
C4 |
C2 |
H6 |
121.308 |
| C5 |
C3 |
H7 |
121.308 |
|
H8 |
C4 |
H9 |
117.505 |
| H10 |
C5 |
H11 |
117.505 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.310 |
|
|
|
| 2 |
C |
0.623 |
|
|
|
| 3 |
C |
0.623 |
|
|
|
| 4 |
C |
0.613 |
|
|
|
| 5 |
C |
0.613 |
|
|
|
| 6 |
H |
-0.683 |
|
|
|
| 7 |
H |
-0.683 |
|
|
|
| 8 |
H |
-0.336 |
|
|
|
| 9 |
H |
-0.373 |
|
|
|
| 10 |
H |
-0.336 |
|
|
|
| 11 |
H |
-0.373 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.112 |
1.112 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.865 |
0.001 |
0.000 |
| y |
0.001 |
-33.851 |
0.000 |
| z |
0.000 |
0.000 |
-35.228 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.326 |
0.001 |
0.000 |
| y |
0.001 |
5.195 |
0.000 |
| z |
0.000 |
0.000 |
3.131 |
|
| Polar |
|---|
| 3z2-r2 | 6.261 |
|---|
| x2-y2 | -9.014 |
|---|
| xy | 0.001 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.269 |
0.004 |
0.000 |
| y |
0.004 |
18.547 |
0.000 |
| z |
0.000 |
0.000 |
9.013 |
<r2> (average value of r
2) Å
2
| <r2> |
189.744 |
| (<r2>)1/2 |
13.775 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -554.173189 |
| Energy at 298.15K | -554.178677 |
| HF Energy | -554.173189 |
| Nuclear repulsion energy | 201.603679 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3277 |
3141 |
4.91 |
|
|
|
| 2 |
A |
3275 |
3138 |
3.40 |
|
|
|
| 3 |
A |
3208 |
3074 |
2.50 |
|
|
|
| 4 |
A |
3194 |
3061 |
3.98 |
|
|
|
| 5 |
A |
3179 |
3046 |
0.97 |
|
|
|
| 6 |
A |
3173 |
3041 |
1.31 |
|
|
|
| 7 |
A |
1671 |
1602 |
54.90 |
|
|
|
| 8 |
A |
1662 |
1593 |
44.22 |
|
|
|
| 9 |
A |
1415 |
1356 |
5.33 |
|
|
|
| 10 |
A |
1411 |
1352 |
6.04 |
|
|
|
| 11 |
A |
1297 |
1243 |
1.95 |
|
|
|
| 12 |
A |
1280 |
1226 |
7.80 |
|
|
|
| 13 |
A |
1053 |
1010 |
11.34 |
|
|
|
| 14 |
A |
1025 |
982 |
10.05 |
|
|
|
| 15 |
A |
993 |
952 |
24.80 |
|
|
|
| 16 |
A |
982 |
941 |
31.12 |
|
|
|
| 17 |
A |
944 |
904 |
33.77 |
|
|
|
| 18 |
A |
909 |
871 |
44.69 |
|
|
|
| 19 |
A |
752 |
721 |
16.95 |
|
|
|
| 20 |
A |
705 |
676 |
5.31 |
|
|
|
| 21 |
A |
627 |
601 |
14.88 |
|
|
|
| 22 |
A |
608 |
583 |
16.20 |
|
|
|
| 23 |
A |
464 |
445 |
0.46 |
|
|
|
| 24 |
A |
366 |
350 |
0.56 |
|
|
|
| 25 |
A |
220 |
211 |
0.85 |
|
|
|
| 26 |
A |
151 |
145 |
4.19 |
|
|
|
| 27 |
A |
88 |
84 |
1.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18963.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 18172.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.025 |
-0.963 |
0.010 |
| C2 |
1.081 |
0.410 |
0.334 |
| C3 |
-1.569 |
-0.216 |
-0.018 |
| C4 |
2.297 |
0.533 |
-0.203 |
| C5 |
-1.868 |
1.081 |
-0.131 |
| H6 |
0.703 |
1.115 |
1.077 |
| H7 |
-2.355 |
-0.969 |
0.071 |
| H8 |
2.958 |
1.339 |
0.112 |
| H9 |
2.671 |
-0.165 |
-0.951 |
| H10 |
-1.110 |
1.852 |
-0.253 |
| H11 |
-2.909 |
1.399 |
-0.112 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7624 | 1.7607 | 2.7290 | 2.7898 | 2.4324 | 2.3801 | 3.7303 | 2.9268 | 3.0465 | 3.7690 |
C2 | 1.7624 | | 2.7460 | 1.3348 | 3.0602 | 1.0918 | 3.7113 | 2.1061 | 2.1235 | 2.6874 | 4.1350 | C3 | 1.7607 | 2.7460 | | 3.9427 | 1.3361 | 2.8522 | 1.0911 | 4.7890 | 4.3423 | 2.1316 | 2.1010 | C4 | 2.7290 | 1.3348 | 3.9427 | | 4.2019 | 2.1254 | 4.8956 | 1.0891 | 1.0890 | 3.6536 | 5.2786 | C5 | 2.7898 | 3.0602 | 1.3361 | 4.2019 | | 2.8409 | 2.1163 | 4.8396 | 4.7782 | 1.0883 | 1.0886 | H6 | 2.4324 | 1.0918 | 2.8522 | 2.1254 | 2.8409 | | 3.8345 | 2.4629 | 3.1020 | 2.3655 | 3.8131 | H7 | 2.3801 | 3.7113 | 1.0911 | 4.8956 | 2.1163 | 3.8345 | | 5.7925 | 5.1913 | 3.0999 | 2.4391 | H8 | 3.7303 | 2.1061 | 4.7890 | 1.0891 | 4.8396 | 2.4629 | 5.7925 | | 1.8636 | 4.1165 | 5.8719 | H9 | 2.9268 | 2.1235 | 4.3423 | 1.0890 | 4.7782 | 3.1020 | 5.1913 | 1.8636 | | 4.3416 | 5.8558 | H10 | 3.0465 | 2.6874 | 2.1316 | 3.6536 | 1.0883 | 2.3655 | 3.0999 | 4.1165 | 4.3416 | | 1.8606 | H11 | 3.7690 | 4.1350 | 2.1010 | 5.2786 | 1.0886 | 3.8131 | 2.4391 | 5.8719 | 5.8558 | 1.8606 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.956 |
|
S1 |
C2 |
H6 |
114.866 |
| S1 |
C3 |
C5 |
128.015 |
|
S1 |
C3 |
H7 |
110.958 |
| C2 |
S1 |
C3 |
102.418 |
|
C2 |
C4 |
H8 |
120.316 |
| C2 |
C4 |
H9 |
122.023 |
|
C3 |
C5 |
H10 |
122.768 |
| C3 |
C5 |
H11 |
119.766 |
|
C4 |
C2 |
H6 |
121.974 |
| C5 |
C3 |
H7 |
121.026 |
|
H8 |
C4 |
H9 |
117.655 |
| H10 |
C5 |
H11 |
117.464 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.303 |
|
|
|
| 2 |
C |
0.642 |
|
|
|
| 3 |
C |
0.665 |
|
|
|
| 4 |
C |
0.485 |
|
|
|
| 5 |
C |
0.405 |
|
|
|
| 6 |
H |
-0.525 |
|
|
|
| 7 |
H |
-0.521 |
|
|
|
| 8 |
H |
-0.338 |
|
|
|
| 9 |
H |
-0.344 |
|
|
|
| 10 |
H |
-0.406 |
|
|
|
| 11 |
H |
-0.366 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.058 |
0.942 |
0.148 |
0.956 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.243 |
1.661 |
-0.705 |
| y |
1.661 |
-37.461 |
1.222 |
| z |
-0.705 |
1.222 |
-41.065 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.020 |
1.661 |
-0.705 |
| y |
1.661 |
-0.307 |
1.222 |
| z |
-0.705 |
1.222 |
-5.713 |
|
| Polar |
|---|
| 3z2-r2 | -11.425 |
|---|
| x2-y2 | 4.218 |
|---|
| xy | 1.661 |
|---|
| xz | -0.705 |
|---|
| yz | 1.222 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.984 |
-0.270 |
-0.450 |
| y |
-0.270 |
10.811 |
0.014 |
| z |
-0.450 |
0.014 |
7.759 |
<r2> (average value of r
2) Å
2
| <r2> |
166.710 |
| (<r2>)1/2 |
12.912 |
