return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H7NO (Ethoxyacetonitrile)

using model chemistry: mPW1PW91/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at mPW1PW91/aug-cc-pVDZ
 hartrees
Energy at 0K-286.538568
Energy at 298.15K-286.546354
HF Energy-286.538568
Nuclear repulsion energy216.233081
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3159 3027 17.93      
2 A' 3071 2943 13.58      
3 A' 3007 2882 60.67      
4 A' 3000 2875 21.23      
5 A' 2389 2289 1.05      
6 A' 1506 1443 5.58      
7 A' 1480 1418 2.47      
8 A' 1472 1410 4.92      
9 A' 1436 1376 8.03      
10 A' 1390 1332 18.34      
11 A' 1362 1305 43.38      
12 A' 1181 1132 215.63      
13 A' 1153 1105 17.68      
14 A' 1074 1029 19.79      
15 A' 967 926 4.50      
16 A' 909 871 5.12      
17 A' 558 535 0.97      
18 A' 422 405 1.09      
19 A' 300 287 2.34      
20 A' 132 127 3.59      
21 A" 3164 3032 20.27      
22 A" 3044 2917 1.71      
23 A" 3038 2911 58.83      
24 A" 1462 1401 7.02      
25 A" 1295 1241 2.45      
26 A" 1248 1196 3.23      
27 A" 1175 1126 5.53      
28 A" 1018 976 0.77      
29 A" 820 786 0.85      
30 A" 370 354 1.60      
31 A" 243 233 0.18      
32 A" 116 111 6.49      
33 A" 77 74 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 23518.2 cm-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 22537.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at mPW1PW91/aug-cc-pVDZ
ABC
0.53053 0.04978 0.04672

See section I.F.4 to change rotational constant units
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ

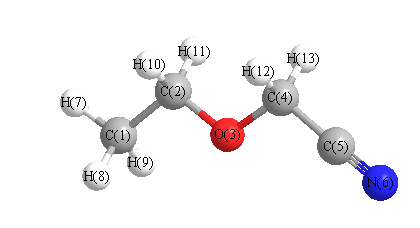
Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.005 2.519 0.000
C2 -1.254 1.030 0.000
O3 0.000 0.363 0.000
C4 -0.151 -1.029 0.000
C5 1.169 -1.665 0.000
N6 2.194 -2.202 0.000
H7 -1.962 3.055 0.000
H8 -0.439 2.816 0.890
H9 -0.439 2.816 -0.890
H10 -1.832 0.727 -0.891
H11 -1.832 0.727 0.891
H12 -0.702 -1.379 -0.891
H13 -0.702 -1.379 0.891

Atom - Atom Distances (Å)
  C1 C2 O3 C4 C5 N6 H7 H8 H9 H10 H11 H12 H13
C11.50942.37843.64944.71455.70261.09671.09591.09592.16472.16474.00964.0096
C21.50941.42042.33623.62364.72592.14532.15592.15591.10431.10432.62702.6270
O32.37841.42041.40052.34043.37543.33102.64632.64632.06962.06962.07882.0788
C43.64942.33621.40051.46432.62154.46793.95783.95782.58992.58991.10461.1046
C54.71453.62362.34041.46431.15765.66364.84324.84323.93963.93962.09162.0916
N65.70264.72593.37542.62151.15766.70135.73665.73665.05815.05813.13963.1396
H71.09672.14533.33104.46795.66366.70131.77991.77992.49572.49574.69484.6948
H81.09592.15592.64633.95784.84325.73661.77991.77963.07832.51084.56524.2035
H91.09592.15592.64633.95784.84325.73661.77991.77962.51083.07834.20354.5652
H102.16471.10432.06962.58993.93965.05812.49573.07832.51081.78222.39042.9817
H112.16471.10432.06962.58993.93965.05812.49572.51083.07831.78222.98172.3904
H124.00962.62702.07881.10462.09163.13964.69484.56524.20352.39042.98171.7823
H134.00962.62702.07881.10462.09163.13964.69484.20354.56522.98172.39041.7823

picture of Ethoxyacetonitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 108.507 C1 C2 H10 110.874
C1 C2 H11 110.874 C2 C1 H7 109.791
C2 C1 H8 110.680 C2 C1 H9 110.680
C2 O3 C4 111.819 O3 C2 H10 109.484
O3 C2 H11 109.484 O3 C4 C5 109.541
O3 C4 H12 111.611 O3 C4 H13 111.611
C4 C5 N6 178.057 C5 C4 H12 108.197
C5 C4 H13 108.197 H7 C1 H8 108.533
H7 C1 H9 108.533 H8 C1 H9 108.564
H10 C2 H11 107.601 H12 C4 H13 107.558
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability