Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -189.360713 |
| Energy at 298.15K | -189.363403 |
| HF Energy | -188.793637 |
| Nuclear repulsion energy | 69.217089 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3655 |
3571 |
|
|
|
|
| 2 |
A' |
3107 |
3035 |
|
|
|
|
| 3 |
A' |
1804 |
1762 |
|
|
|
|
| 4 |
A' |
1419 |
1387 |
|
|
|
|
| 5 |
A' |
1332 |
1301 |
|
|
|
|
| 6 |
A' |
1137 |
1111 |
|
|
|
|
| 7 |
A' |
617 |
603 |
|
|
|
|
| 8 |
A" |
1044 |
1020 |
|
|
|
|
| 9 |
A" |
705 |
689 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7410.0 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 7238.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.428 |
0.000 |
| O2 |
-1.048 |
-0.447 |
0.000 |
| O3 |
1.178 |
0.111 |
0.000 |
| H4 |
-0.385 |
1.465 |
0.000 |
| H5 |
-0.658 |
-1.342 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3648 | 1.2198 | 1.1069 | 1.8875 |
O2 | 1.3648 | | 2.2947 | 2.0242 | 0.9758 | O3 | 1.2198 | 2.2947 | | 2.0681 | 2.3411 | H4 | 1.1069 | 2.0242 | 2.0681 | | 2.8203 | H5 | 1.8875 | 0.9758 | 2.3411 | 2.8203 | |
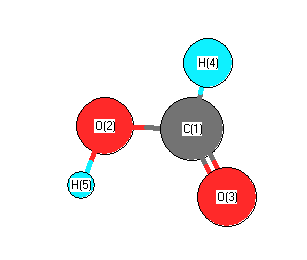 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.301 |
|
O2 |
C1 |
O3 |
125.109 |
| O2 |
C1 |
H4 |
109.511 |
|
O3 |
C1 |
H4 |
125.380 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -189.354079 |
| Energy at 298.15K | |
| HF Energy | -188.785943 |
| Nuclear repulsion energy | 69.018721 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3713 |
3627 |
|
|
|
|
| 2 |
A' |
2995 |
2926 |
|
|
|
|
| 3 |
A' |
1845 |
1803 |
|
|
|
|
| 4 |
A' |
1444 |
1411 |
|
|
|
|
| 5 |
A' |
1306 |
1276 |
|
|
|
|
| 6 |
A' |
1127 |
1101 |
|
|
|
|
| 7 |
A' |
645 |
630 |
|
|
|
|
| 8 |
A" |
1027 |
1003 |
|
|
|
|
| 9 |
A" |
525 |
513 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7313.1 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 7144.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.393 |
0.000 |
| O2 |
-0.912 |
-0.631 |
0.000 |
| O3 |
1.196 |
0.192 |
0.000 |
| H4 |
-0.466 |
1.405 |
0.000 |
| H5 |
-1.806 |
-0.251 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3717 | 1.2131 | 1.1134 | 1.9173 |
O2 | 1.3717 | | 2.2633 | 2.0841 | 0.9712 | O3 | 1.2131 | 2.2633 | | 2.0575 | 3.0345 | H4 | 1.1134 | 2.0841 | 2.0575 | | 2.1299 | H5 | 1.9173 | 0.9712 | 3.0345 | 2.1299 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.632 |
|
O2 |
C1 |
O3 |
122.118 |
| O2 |
C1 |
H4 |
113.587 |
|
O3 |
C1 |
H4 |
124.295 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
