Jump to
S1C2
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -189.690177 |
| Energy at 298.15K | -189.692865 |
| HF Energy | -189.690177 |
| Nuclear repulsion energy | 69.499049 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3624 |
3624 |
39.43 |
85.94 |
0.17 |
0.30 |
| 2 |
A' |
2997 |
2997 |
51.50 |
136.84 |
0.25 |
0.40 |
| 3 |
A' |
1755 |
1755 |
339.91 |
13.23 |
0.11 |
0.21 |
| 4 |
A' |
1359 |
1359 |
1.41 |
5.45 |
0.68 |
0.81 |
| 5 |
A' |
1265 |
1265 |
3.54 |
2.01 |
0.20 |
0.34 |
| 6 |
A' |
1079 |
1079 |
241.49 |
2.94 |
0.34 |
0.50 |
| 7 |
A' |
606 |
606 |
35.68 |
3.81 |
0.33 |
0.49 |
| 8 |
A" |
1012 |
1012 |
4.46 |
1.30 |
0.75 |
0.86 |
| 9 |
A" |
677 |
677 |
124.17 |
0.54 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7185.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7185.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.425 |
0.000 |
| O2 |
-1.041 |
-0.449 |
0.000 |
| O3 |
1.172 |
0.117 |
0.000 |
| H4 |
-0.392 |
1.462 |
0.000 |
| H5 |
-0.654 |
-1.347 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3594 | 1.2116 | 1.1083 | 1.8890 |
O2 | 1.3594 | | 2.2841 | 2.0183 | 0.9775 | O3 | 1.2116 | 2.2841 | | 2.0624 | 2.3405 | H4 | 1.1083 | 2.0183 | 2.0624 | | 2.8210 | H5 | 1.8890 | 0.9775 | 2.3405 | 2.8210 | |
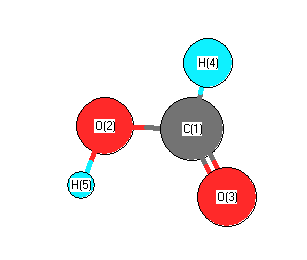 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.722 |
|
O2 |
C1 |
O3 |
125.244 |
| O2 |
C1 |
H4 |
109.326 |
|
O3 |
C1 |
H4 |
125.430 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.823 |
|
|
|
| 2 |
O |
-0.290 |
|
|
|
| 3 |
O |
-0.401 |
|
|
|
| 4 |
H |
-0.240 |
|
|
|
| 5 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.476 |
-0.286 |
0.000 |
1.503 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.219 |
-0.042 |
0.000 |
| y |
-0.042 |
-13.089 |
0.000 |
| z |
0.000 |
0.000 |
-17.157 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.096 |
-0.042 |
0.000 |
| y |
-0.042 |
6.599 |
0.000 |
| z |
0.000 |
0.000 |
0.497 |
|
| Polar |
|---|
| 3z2-r2 | 0.993 |
|---|
| x2-y2 | -9.130 |
|---|
| xy | -0.042 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.276 |
0.118 |
0.000 |
| y |
0.118 |
3.769 |
0.000 |
| z |
0.000 |
0.000 |
2.429 |
<r2> (average value of r
2) Å
2
| <r2> |
37.918 |
| (<r2>)1/2 |
6.158 |
Jump to
S1C1
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -189.684118 |
| Energy at 298.15K | |
| HF Energy | -189.684118 |
| Nuclear repulsion energy | 69.309089 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3679 |
3679 |
32.37 |
135.51 |
0.21 |
0.34 |
| 2 |
A' |
2901 |
2901 |
92.28 |
129.85 |
0.25 |
0.40 |
| 3 |
A' |
1797 |
1797 |
272.49 |
19.78 |
0.20 |
0.34 |
| 4 |
A' |
1365 |
1365 |
0.26 |
3.76 |
0.60 |
0.75 |
| 5 |
A' |
1244 |
1244 |
264.14 |
3.45 |
0.56 |
0.72 |
| 6 |
A' |
1062 |
1062 |
65.09 |
10.74 |
0.43 |
0.60 |
| 7 |
A' |
631 |
631 |
8.59 |
0.30 |
0.55 |
0.71 |
| 8 |
A" |
987 |
987 |
0.26 |
1.64 |
0.75 |
0.86 |
| 9 |
A" |
543 |
543 |
75.45 |
1.24 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7105.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7105.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.391 |
0.000 |
| O2 |
-0.905 |
-0.633 |
0.000 |
| O3 |
1.189 |
0.197 |
0.000 |
| H4 |
-0.472 |
1.402 |
0.000 |
| H5 |
-1.804 |
-0.261 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3662 | 1.2049 | 1.1154 | 1.9181 |
O2 | 1.3662 | | 2.2523 | 2.0798 | 0.9729 | O3 | 1.2049 | 2.2523 | | 2.0519 | 3.0279 | H4 | 1.1154 | 2.0798 | 2.0519 | | 2.1304 | H5 | 1.9181 | 0.9729 | 3.0279 | 2.1304 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.017 |
|
O2 |
C1 |
O3 |
122.210 |
| O2 |
C1 |
H4 |
113.495 |
|
O3 |
C1 |
H4 |
124.295 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.853 |
|
|
|
| 2 |
O |
-0.348 |
|
|
|
| 3 |
O |
-0.417 |
|
|
|
| 4 |
H |
-0.252 |
|
|
|
| 5 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.486 |
1.474 |
0.000 |
3.785 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.166 |
-1.160 |
0.000 |
| y |
-1.160 |
-17.590 |
0.000 |
| z |
0.000 |
0.000 |
-17.163 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.210 |
-1.160 |
0.000 |
| y |
-1.160 |
-0.925 |
0.000 |
| z |
0.000 |
0.000 |
-0.285 |
|
| Polar |
|---|
| 3z2-r2 | -0.569 |
|---|
| x2-y2 | 1.424 |
|---|
| xy | -1.160 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.852 |
0.134 |
0.000 |
| y |
0.134 |
3.477 |
0.000 |
| z |
0.000 |
0.000 |
2.440 |
<r2> (average value of r
2) Å
2
| <r2> |
38.402 |
| (<r2>)1/2 |
6.197 |
