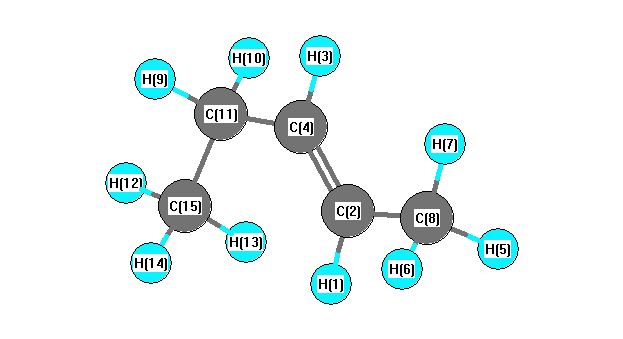
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3178 |
3045 |
25.95 |
|
|
|
| 2 |
A' |
3149 |
3018 |
23.74 |
|
|
|
| 3 |
A' |
3143 |
3011 |
32.70 |
|
|
|
| 4 |
A' |
3138 |
3007 |
14.47 |
|
|
|
| 5 |
A' |
3057 |
2929 |
23.38 |
|
|
|
| 6 |
A' |
3040 |
2913 |
40.11 |
|
|
|
| 7 |
A' |
3032 |
2906 |
34.15 |
|
|
|
| 8 |
A' |
1763 |
1690 |
0.08 |
|
|
|
| 9 |
A' |
1486 |
1424 |
5.21 |
|
|
|
| 10 |
A' |
1471 |
1410 |
12.54 |
|
|
|
| 11 |
A' |
1461 |
1400 |
3.64 |
|
|
|
| 12 |
A' |
1406 |
1347 |
8.60 |
|
|
|
| 13 |
A' |
1393 |
1335 |
0.03 |
|
|
|
| 14 |
A' |
1365 |
1308 |
5.32 |
|
|
|
| 15 |
A' |
1333 |
1277 |
1.00 |
|
|
|
| 16 |
A' |
1316 |
1261 |
0.08 |
|
|
|
| 17 |
A' |
1153 |
1104 |
0.36 |
|
|
|
| 18 |
A' |
1112 |
1066 |
7.91 |
|
|
|
| 19 |
A' |
1029 |
986 |
2.67 |
|
|
|
| 20 |
A' |
962 |
922 |
8.89 |
|
|
|
| 21 |
A' |
872 |
836 |
0.06 |
|
|
|
| 22 |
A' |
587 |
562 |
0.72 |
|
|
|
| 23 |
A' |
366 |
351 |
0.29 |
|
|
|
| 24 |
A' |
204 |
196 |
0.36 |
|
|
|
| 25 |
A" |
3135 |
3004 |
31.60 |
|
|
|
| 26 |
A" |
3102 |
2973 |
18.85 |
|
|
|
| 27 |
A" |
3059 |
2931 |
15.21 |
|
|
|
| 28 |
A" |
1478 |
1416 |
7.76 |
|
|
|
| 29 |
A" |
1455 |
1394 |
7.02 |
|
|
|
| 30 |
A" |
1281 |
1228 |
0.37 |
|
|
|
| 31 |
A" |
1098 |
1052 |
2.47 |
|
|
|
| 32 |
A" |
1058 |
1014 |
1.41 |
|
|
|
| 33 |
A" |
1003 |
961 |
34.63 |
|
|
|
| 34 |
A" |
829 |
794 |
2.01 |
|
|
|
| 35 |
A" |
712 |
682 |
0.57 |
|
|
|
| 36 |
A" |
300 |
288 |
2.12 |
|
|
|
| 37 |
A" |
230 |
221 |
0.06 |
|
|
|
| 38 |
A" |
205 |
197 |
3.03 |
|
|
|
| 39 |
A" |
133 |
127 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30043.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28791.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.840 |
|
|
|
| 2 |
C |
0.549 |
|
|
|
| 3 |
H |
-0.709 |
|
|
|
| 4 |
C |
0.839 |
|
|
|
| 5 |
H |
0.014 |
|
|
|
| 6 |
H |
0.014 |
|
|
|
| 7 |
H |
-0.203 |
|
|
|
| 8 |
C |
0.048 |
|
|
|
| 9 |
H |
-0.072 |
|
|
|
| 10 |
H |
-0.072 |
|
|
|
| 11 |
C |
0.284 |
|
|
|
| 12 |
H |
-0.253 |
|
|
|
| 13 |
H |
-0.033 |
|
|
|
| 14 |
H |
-0.033 |
|
|
|
| 15 |
C |
0.467 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.109 |
0.015 |
0.000 |
0.110 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.055 |
0.539 |
0.000 |
| y |
0.539 |
-32.566 |
0.000 |
| z |
0.000 |
0.000 |
-34.609 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.532 |
0.539 |
0.000 |
| y |
0.539 |
0.766 |
0.000 |
| z |
0.000 |
0.000 |
-2.298 |
|
| Polar |
|---|
| 3z2-r2 | -4.597 |
|---|
| x2-y2 | 0.511 |
|---|
| xy | 0.539 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.153 |
0.464 |
0.000 |
| y |
0.464 |
9.299 |
0.000 |
| z |
0.000 |
0.000 |
7.637 |
<r2> (average value of r
2) Å
2
| <r2> |
171.581 |
| (<r2>)1/2 |
13.099 |