Jump to
S1C2
S1C3
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -268.319591 |
| Energy at 298.15K | -268.327905 |
| Nuclear repulsion energy | 194.611762 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3159 |
3027 |
23.83 |
|
|
|
| 2 |
A |
3139 |
3008 |
32.82 |
|
|
|
| 3 |
A |
3116 |
2986 |
33.88 |
|
|
|
| 4 |
A |
3067 |
2939 |
57.72 |
|
|
|
| 5 |
A |
3033 |
2907 |
50.96 |
|
|
|
| 6 |
A |
2980 |
2855 |
112.75 |
|
|
|
| 7 |
A |
1533 |
1469 |
1.83 |
|
|
|
| 8 |
A |
1516 |
1453 |
0.86 |
|
|
|
| 9 |
A |
1501 |
1439 |
2.91 |
|
|
|
| 10 |
A |
1412 |
1353 |
5.61 |
|
|
|
| 11 |
A |
1373 |
1316 |
0.11 |
|
|
|
| 12 |
A |
1337 |
1282 |
1.08 |
|
|
|
| 13 |
A |
1271 |
1218 |
6.68 |
|
|
|
| 14 |
A |
1231 |
1179 |
4.47 |
|
|
|
| 15 |
A |
1206 |
1155 |
5.84 |
|
|
|
| 16 |
A |
1191 |
1141 |
105.32 |
|
|
|
| 17 |
A |
1154 |
1106 |
26.31 |
|
|
|
| 18 |
A |
1125 |
1078 |
73.77 |
|
|
|
| 19 |
A |
1087 |
1042 |
36.35 |
|
|
|
| 20 |
A |
1008 |
966 |
23.79 |
|
|
|
| 21 |
A |
980 |
939 |
38.48 |
|
|
|
| 22 |
A |
958 |
918 |
48.98 |
|
|
|
| 23 |
A |
874 |
838 |
9.52 |
|
|
|
| 24 |
A |
745 |
714 |
0.47 |
|
|
|
| 25 |
A |
687 |
659 |
5.39 |
|
|
|
| 26 |
A |
279 |
267 |
11.10 |
|
|
|
| 27 |
A |
55 |
53 |
4.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20507.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 19652.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.171 |
-0.084 |
0.145 |
| C2 |
-1.030 |
-0.664 |
-0.054 |
| C3 |
-0.847 |
0.843 |
0.150 |
| O4 |
0.301 |
-1.177 |
-0.063 |
| O5 |
0.489 |
1.066 |
-0.273 |
| H6 |
1.433 |
-0.011 |
1.219 |
| H7 |
2.071 |
-0.210 |
-0.466 |
| H8 |
-1.501 |
-0.901 |
-1.017 |
| H9 |
-0.963 |
1.122 |
1.211 |
| H10 |
-1.512 |
1.459 |
-0.462 |
| H11 |
-1.608 |
-1.131 |
0.755 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 2.2847 | 2.2204 | 1.4119 | 1.4010 | 1.1072 | 1.0954 | 3.0261 | 2.6727 | 3.1537 | 3.0313 |
C2 | 2.2847 | | 1.5318 | 1.4262 | 2.3125 | 2.8484 | 3.1610 | 1.0980 | 2.1898 | 2.2151 | 1.0983 | C3 | 2.2204 | 1.5318 | | 2.3325 | 1.4188 | 2.6585 | 3.1624 | 2.1981 | 1.1024 | 1.0941 | 2.1996 | O4 | 1.4119 | 1.4262 | 2.3325 | | 2.2599 | 2.0695 | 2.0567 | 2.0576 | 2.9161 | 3.2235 | 2.0775 | O5 | 1.4010 | 2.3125 | 1.4188 | 2.2599 | | 2.0679 | 2.0415 | 2.8948 | 2.0766 | 2.0474 | 3.2058 | H6 | 1.1072 | 2.8484 | 2.6585 | 2.0695 | 2.0679 | | 1.8124 | 3.7947 | 2.6505 | 3.6955 | 3.2737 | H7 | 1.0954 | 3.1610 | 3.1624 | 2.0567 | 2.0415 | 1.8124 | | 3.6796 | 3.7135 | 3.9521 | 3.9843 | H8 | 3.0261 | 1.0980 | 2.1981 | 2.0576 | 2.8948 | 3.7947 | 3.6796 | | 3.0570 | 2.4242 | 1.7904 | H9 | 2.6727 | 2.1898 | 1.1024 | 2.9161 | 2.0766 | 2.6505 | 3.7135 | 3.0570 | | 1.7922 | 2.3870 | H10 | 3.1537 | 2.2151 | 1.0941 | 3.2235 | 2.0474 | 3.6955 | 3.9521 | 2.4242 | 1.7922 | | 2.8628 | H11 | 3.0313 | 1.0983 | 2.1996 | 2.0775 | 3.2058 | 3.2737 | 3.9843 | 1.7904 | 2.3870 | 2.8628 | |
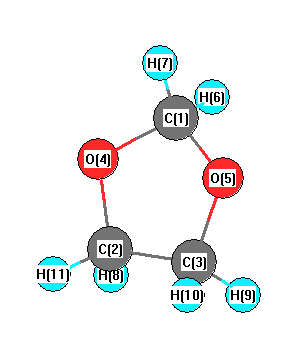 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C2 |
107.229 |
|
C1 |
O5 |
C3 |
103.890 |
| C2 |
C3 |
O5 |
103.142 |
|
C2 |
C3 |
H9 |
111.421 |
| C2 |
C3 |
H10 |
114.000 |
|
C3 |
C2 |
O4 |
104.039 |
| C3 |
C2 |
H8 |
112.356 |
|
C3 |
C2 |
H11 |
112.460 |
| O4 |
C1 |
O5 |
106.916 |
|
O4 |
C1 |
H6 |
109.891 |
| O4 |
C1 |
H7 |
109.579 |
|
O4 |
C2 |
H8 |
108.514 |
| O4 |
C2 |
H11 |
110.086 |
|
O5 |
C1 |
H6 |
110.522 |
| O5 |
C1 |
H7 |
109.107 |
|
O5 |
C3 |
H9 |
110.274 |
| O5 |
C3 |
H10 |
108.440 |
|
H6 |
C1 |
H7 |
110.743 |
| H8 |
C2 |
H11 |
109.215 |
|
H9 |
C3 |
H10 |
109.358 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.209 |
|
|
|
| 2 |
C |
0.975 |
|
|
|
| 3 |
C |
0.782 |
|
|
|
| 4 |
O |
-0.657 |
|
|
|
| 5 |
O |
-0.622 |
|
|
|
| 6 |
H |
-0.251 |
|
|
|
| 7 |
H |
-0.316 |
|
|
|
| 8 |
H |
-0.298 |
|
|
|
| 9 |
H |
-0.286 |
|
|
|
| 10 |
H |
-0.286 |
|
|
|
| 11 |
H |
-0.249 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.182 |
0.356 |
0.762 |
1.450 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.268 |
-0.763 |
0.499 |
| y |
-0.763 |
-34.487 |
0.553 |
| z |
0.499 |
0.553 |
-29.805 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.878 |
-0.763 |
0.499 |
| y |
-0.763 |
-6.451 |
0.553 |
| z |
0.499 |
0.553 |
0.573 |
|
| Polar |
|---|
| 3z2-r2 | 1.146 |
|---|
| x2-y2 | 8.219 |
|---|
| xy | -0.763 |
|---|
| xz | 0.499 |
|---|
| yz | 0.553 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.401 |
-0.054 |
0.058 |
| y |
-0.054 |
6.346 |
0.072 |
| z |
0.058 |
0.072 |
5.923 |
<r2> (average value of r
2) Å
2
| <r2> |
93.305 |
| (<r2>)1/2 |
9.659 |
