Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -751.300917 |
| Energy at 298.15K | -751.302080 |
| HF Energy | -751.300917 |
| Nuclear repulsion energy | 561.897084 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1866 |
1789 |
0.00 |
|
|
|
| 2 |
Ag |
1450 |
1389 |
0.00 |
|
|
|
| 3 |
Ag |
1337 |
1282 |
0.00 |
|
|
|
| 4 |
Ag |
1193 |
1144 |
0.00 |
|
|
|
| 5 |
Ag |
722 |
692 |
0.00 |
|
|
|
| 6 |
Ag |
588 |
563 |
0.00 |
|
|
|
| 7 |
Ag |
380 |
364 |
0.00 |
|
|
|
| 8 |
Ag |
352 |
337 |
0.00 |
|
|
|
| 9 |
Ag |
217 |
208 |
0.00 |
|
|
|
| 10 |
Au |
580 |
556 |
2.21 |
|
|
|
| 11 |
Au |
345 |
331 |
3.45 |
|
|
|
| 12 |
Au |
109 |
105 |
0.07 |
|
|
|
| 13 |
Au |
12 |
12 |
0.14 |
|
|
|
| 14 |
Bg |
651 |
624 |
0.00 |
|
|
|
| 15 |
Bg |
472 |
452 |
0.00 |
|
|
|
| 16 |
Bg |
180 |
172 |
0.00 |
|
|
|
| 17 |
Bu |
1809 |
1733 |
402.42 |
|
|
|
| 18 |
Bu |
1355 |
1298 |
447.10 |
|
|
|
| 19 |
Bu |
1227 |
1176 |
280.59 |
|
|
|
| 20 |
Bu |
966 |
926 |
301.08 |
|
|
|
| 21 |
Bu |
621 |
595 |
4.84 |
|
|
|
| 22 |
Bu |
494 |
473 |
3.70 |
|
|
|
| 23 |
Bu |
288 |
276 |
4.61 |
|
|
|
| 24 |
Bu |
133 |
128 |
0.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8674.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 8312.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.151 |
1.860 |
0.000 |
| C2 |
0.462 |
0.558 |
0.000 |
| C3 |
-0.462 |
-0.558 |
0.000 |
| C4 |
-0.151 |
-1.860 |
0.000 |
| F5 |
1.066 |
2.804 |
0.000 |
| F6 |
-1.066 |
2.353 |
0.000 |
| F7 |
1.766 |
0.238 |
0.000 |
| F8 |
-1.766 |
-0.238 |
0.000 |
| F9 |
1.066 |
-2.353 |
0.000 |
| F10 |
-1.066 |
-2.804 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3382 | 2.4944 | 3.7315 | 1.3152 | 1.3128 | 2.2887 | 2.8417 | 4.3110 | 4.8199 |
C2 | 1.3382 | | 1.4496 | 2.4944 | 2.3256 | 2.3573 | 1.3425 | 2.3664 | 2.9733 | 3.6934 | C3 | 2.4944 | 1.4496 | | 1.3382 | 3.6934 | 2.9733 | 2.3664 | 1.3425 | 2.3573 | 2.3256 | C4 | 3.7315 | 2.4944 | 1.3382 | | 4.8199 | 4.3110 | 2.8417 | 2.2887 | 1.3128 | 1.3152 | F5 | 1.3152 | 2.3256 | 3.6934 | 4.8199 | | 2.1789 | 2.6597 | 4.1565 | 5.1573 | 5.9998 | F6 | 1.3128 | 2.3573 | 2.9733 | 4.3110 | 2.1789 | | 3.5344 | 2.6843 | 5.1665 | 5.1573 | F7 | 2.2887 | 1.3425 | 2.3664 | 2.8417 | 2.6597 | 3.5344 | | 3.5642 | 2.6843 | 4.1565 | F8 | 2.8417 | 2.3664 | 1.3425 | 2.2887 | 4.1565 | 2.6843 | 3.5642 | | 3.5344 | 2.6597 | F9 | 4.3110 | 2.9733 | 2.3573 | 1.3128 | 5.1573 | 5.1665 | 2.6843 | 3.5344 | | 2.1789 | F10 | 4.8199 | 3.6934 | 2.3256 | 1.3152 | 5.9998 | 5.1573 | 4.1565 | 2.6597 | 2.1789 | |
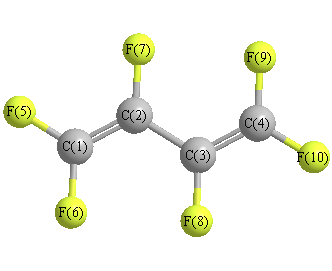 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.907 |
|
C1 |
C2 |
F7 |
117.252 |
| C2 |
C1 |
F5 |
122.441 |
|
C2 |
C1 |
F6 |
125.545 |
| C2 |
C3 |
C4 |
126.907 |
|
C2 |
C3 |
F8 |
115.841 |
| C3 |
C2 |
F7 |
115.841 |
|
C3 |
C4 |
F9 |
125.545 |
| C3 |
C4 |
F10 |
122.441 |
|
C4 |
C3 |
F8 |
117.252 |
| F5 |
C1 |
F6 |
112.014 |
|
F9 |
C4 |
F10 |
112.014 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.808 |
|
|
|
| 2 |
C |
0.696 |
|
|
|
| 3 |
C |
0.696 |
|
|
|
| 4 |
C |
0.808 |
|
|
|
| 5 |
F |
-0.455 |
|
|
|
| 6 |
F |
-0.458 |
|
|
|
| 7 |
F |
-0.590 |
|
|
|
| 8 |
F |
-0.590 |
|
|
|
| 9 |
F |
-0.458 |
|
|
|
| 10 |
F |
-0.455 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-55.844 |
-0.267 |
0.000 |
| y |
-0.267 |
-53.146 |
0.000 |
| z |
0.000 |
0.000 |
-49.663 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.439 |
-0.267 |
0.000 |
| y |
-0.267 |
-0.393 |
0.000 |
| z |
0.000 |
0.000 |
4.832 |
|
| Polar |
|---|
| 3z2-r2 | 9.663 |
|---|
| x2-y2 | -2.698 |
|---|
| xy | -0.267 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.533 |
-0.006 |
0.000 |
| y |
-0.006 |
13.430 |
0.000 |
| z |
0.000 |
0.000 |
5.055 |
<r2> (average value of r
2) Å
2
| <r2> |
420.377 |
| (<r2>)1/2 |
20.503 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -751.303736 |
| Energy at 298.15K | -751.304893 |
| HF Energy | -751.303736 |
| Nuclear repulsion energy | 564.801305 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
1869 |
1791 |
69.92 |
|
|
|
| 2 |
A |
1409 |
1351 |
18.51 |
|
|
|
| 3 |
A |
1349 |
1293 |
172.85 |
|
|
|
| 4 |
A |
1149 |
1101 |
289.06 |
|
|
|
| 5 |
A |
721 |
691 |
0.32 |
|
|
|
| 6 |
A |
670 |
642 |
0.90 |
|
|
|
| 7 |
A |
542 |
519 |
0.01 |
|
|
|
| 8 |
A |
472 |
452 |
0.23 |
|
|
|
| 9 |
A |
380 |
364 |
0.76 |
|
|
|
| 10 |
A |
258 |
247 |
0.30 |
|
|
|
| 11 |
A |
180 |
173 |
0.36 |
|
|
|
| 12 |
A |
92 |
88 |
0.19 |
|
|
|
| 13 |
A |
40 |
38 |
0.00 |
|
|
|
| 14 |
B |
1834 |
1757 |
300.88 |
|
|
|
| 15 |
B |
1347 |
1291 |
293.31 |
|
|
|
| 16 |
B |
1201 |
1151 |
121.06 |
|
|
|
| 17 |
B |
986 |
945 |
204.53 |
|
|
|
| 18 |
B |
641 |
614 |
2.01 |
|
|
|
| 19 |
B |
625 |
599 |
7.22 |
|
|
|
| 20 |
B |
563 |
539 |
3.59 |
|
|
|
| 21 |
B |
425 |
408 |
3.85 |
|
|
|
| 22 |
B |
295 |
283 |
4.47 |
|
|
|
| 23 |
B |
204 |
195 |
2.44 |
|
|
|
| 24 |
B |
105 |
101 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8677.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 8316.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.188 |
1.574 |
-0.366 |
| C2 |
0.159 |
0.706 |
0.585 |
| C3 |
-0.159 |
-0.706 |
0.585 |
| C4 |
0.188 |
-1.574 |
-0.366 |
| F5 |
0.159 |
2.838 |
-0.365 |
| F6 |
-0.903 |
1.262 |
-1.424 |
| F7 |
0.859 |
1.167 |
1.642 |
| F8 |
-0.859 |
-1.167 |
1.642 |
| F9 |
0.903 |
-1.262 |
-1.424 |
| F10 |
-0.159 |
-2.838 |
-0.365 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3334 | 2.4698 | 3.1695 | 1.3111 | 1.3142 | 2.3009 | 3.4627 | 3.2167 | 4.4115 |
C2 | 1.3334 | | 1.4468 | 2.4698 | 2.3344 | 2.3392 | 1.3491 | 2.3791 | 2.9083 | 3.6826 | C3 | 2.4698 | 1.4468 | | 1.3334 | 3.6826 | 2.9083 | 2.3791 | 1.3491 | 2.3392 | 2.3344 | C4 | 3.1695 | 2.4698 | 1.3334 | | 4.4115 | 3.2167 | 3.4627 | 2.3009 | 1.3142 | 1.3111 | F5 | 1.3111 | 2.3344 | 3.6826 | 4.4115 | | 2.1753 | 2.7043 | 4.5938 | 4.2987 | 5.6847 | F6 | 1.3142 | 2.3392 | 2.9083 | 3.2167 | 2.1753 | | 3.5374 | 3.9112 | 3.1027 | 4.2987 | F7 | 2.3009 | 1.3491 | 2.3791 | 3.4627 | 2.7043 | 3.5374 | | 2.8977 | 3.9112 | 4.5938 | F8 | 3.4627 | 2.3791 | 1.3491 | 2.3009 | 4.5938 | 3.9112 | 2.8977 | | 3.5374 | 2.7043 | F9 | 3.2167 | 2.9083 | 2.3392 | 1.3142 | 4.2987 | 3.1027 | 3.9112 | 3.5374 | | 2.1753 | F10 | 4.4115 | 3.6826 | 2.3344 | 1.3111 | 5.6847 | 4.2987 | 4.5938 | 2.7043 | 2.1753 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.283 |
|
C1 |
C2 |
F7 |
118.136 |
| C2 |
C1 |
F5 |
123.947 |
|
C2 |
C1 |
F6 |
124.139 |
| C2 |
C3 |
C4 |
125.283 |
|
C2 |
C3 |
F8 |
116.580 |
| C3 |
C2 |
F7 |
116.580 |
|
C3 |
C4 |
F9 |
124.139 |
| C3 |
C4 |
F10 |
123.947 |
|
C4 |
C3 |
F8 |
118.136 |
| F5 |
C1 |
F6 |
111.908 |
|
F9 |
C4 |
F10 |
111.908 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.071 |
|
|
|
| 2 |
C |
0.385 |
|
|
|
| 3 |
C |
0.385 |
|
|
|
| 4 |
C |
1.071 |
|
|
|
| 5 |
F |
-0.439 |
|
|
|
| 6 |
F |
-0.454 |
|
|
|
| 7 |
F |
-0.563 |
|
|
|
| 8 |
F |
-0.563 |
|
|
|
| 9 |
F |
-0.454 |
|
|
|
| 10 |
F |
-0.439 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.929 |
0.929 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.443 |
-1.573 |
0.000 |
| y |
-1.573 |
-53.156 |
0.000 |
| z |
0.000 |
0.000 |
-54.554 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.413 |
-1.573 |
0.000 |
| y |
-1.573 |
-0.158 |
0.000 |
| z |
0.000 |
0.000 |
-2.255 |
|
| Polar |
|---|
| 3z2-r2 | -4.510 |
|---|
| x2-y2 | 1.714 |
|---|
| xy | -1.573 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.920 |
-0.107 |
0.000 |
| y |
-0.107 |
10.482 |
0.000 |
| z |
0.000 |
0.000 |
8.071 |
<r2> (average value of r
2) Å
2
| <r2> |
389.217 |
| (<r2>)1/2 |
19.729 |
