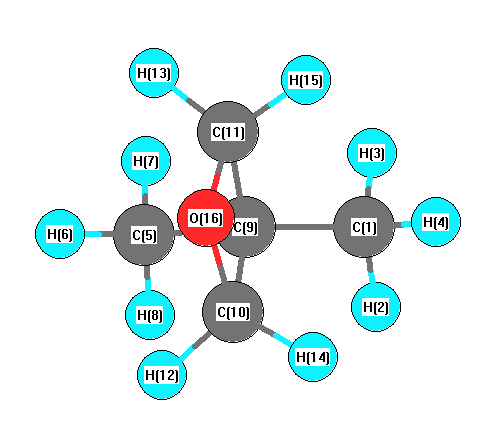
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3136 |
3006 |
32.02 |
|
|
|
| 2 |
A' |
3134 |
3003 |
28.29 |
|
|
|
| 3 |
A' |
3095 |
2966 |
69.51 |
|
|
|
| 4 |
A' |
3050 |
2923 |
23.22 |
|
|
|
| 5 |
A' |
3043 |
2916 |
26.58 |
|
|
|
| 6 |
A' |
3039 |
2912 |
15.76 |
|
|
|
| 7 |
A' |
1525 |
1461 |
2.37 |
|
|
|
| 8 |
A' |
1488 |
1426 |
3.88 |
|
|
|
| 9 |
A' |
1471 |
1410 |
3.03 |
|
|
|
| 10 |
A' |
1408 |
1350 |
4.19 |
|
|
|
| 11 |
A' |
1395 |
1337 |
10.84 |
|
|
|
| 12 |
A' |
1379 |
1322 |
1.06 |
|
|
|
| 13 |
A' |
1305 |
1251 |
0.45 |
|
|
|
| 14 |
A' |
1271 |
1218 |
7.20 |
|
|
|
| 15 |
A' |
1162 |
1114 |
1.07 |
|
|
|
| 16 |
A' |
1037 |
994 |
8.24 |
|
|
|
| 17 |
A' |
990 |
949 |
12.46 |
|
|
|
| 18 |
A' |
958 |
918 |
3.16 |
|
|
|
| 19 |
A' |
942 |
903 |
5.25 |
|
|
|
| 20 |
A' |
855 |
820 |
11.05 |
|
|
|
| 21 |
A' |
650 |
623 |
0.69 |
|
|
|
| 22 |
A' |
412 |
395 |
0.02 |
|
|
|
| 23 |
A' |
344 |
330 |
0.11 |
|
|
|
| 24 |
A' |
75 |
72 |
5.85 |
|
|
|
| 25 |
A" |
3126 |
2995 |
54.17 |
|
|
|
| 26 |
A" |
3122 |
2991 |
0.02 |
|
|
|
| 27 |
A" |
3094 |
2965 |
9.01 |
|
|
|
| 28 |
A" |
3030 |
2904 |
133.38 |
|
|
|
| 29 |
A" |
1504 |
1441 |
2.69 |
|
|
|
| 30 |
A" |
1486 |
1424 |
9.33 |
|
|
|
| 31 |
A" |
1466 |
1405 |
0.01 |
|
|
|
| 32 |
A" |
1300 |
1246 |
0.27 |
|
|
|
| 33 |
A" |
1221 |
1171 |
0.16 |
|
|
|
| 34 |
A" |
1154 |
1106 |
0.08 |
|
|
|
| 35 |
A" |
1059 |
1015 |
81.46 |
|
|
|
| 36 |
A" |
1051 |
1008 |
0.38 |
|
|
|
| 37 |
A" |
932 |
893 |
0.05 |
|
|
|
| 38 |
A" |
895 |
858 |
3.41 |
|
|
|
| 39 |
A" |
390 |
373 |
0.71 |
|
|
|
| 40 |
A" |
314 |
301 |
0.17 |
|
|
|
| 41 |
A" |
260 |
249 |
0.08 |
|
|
|
| 42 |
A" |
211 |
202 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31388.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 30079.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.522 |
|
|
|
| 2 |
H |
-0.044 |
|
|
|
| 3 |
H |
-0.044 |
|
|
|
| 4 |
H |
-0.181 |
|
|
|
| 5 |
C |
0.596 |
|
|
|
| 6 |
H |
-0.141 |
|
|
|
| 7 |
H |
-0.072 |
|
|
|
| 8 |
H |
-0.072 |
|
|
|
| 9 |
C |
-0.901 |
|
|
|
| 10 |
C |
1.230 |
|
|
|
| 11 |
C |
1.230 |
|
|
|
| 12 |
H |
-0.321 |
|
|
|
| 13 |
H |
-0.321 |
|
|
|
| 14 |
H |
-0.382 |
|
|
|
| 15 |
H |
-0.382 |
|
|
|
| 16 |
O |
-0.716 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.325 |
2.007 |
0.000 |
2.033 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.031 |
0.752 |
0.000 |
| y |
0.752 |
-43.845 |
0.000 |
| z |
0.000 |
0.000 |
-36.301 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.042 |
0.752 |
0.000 |
| y |
0.752 |
-6.679 |
0.000 |
| z |
0.000 |
0.000 |
4.637 |
|
| Polar |
|---|
| 3z2-r2 | 9.274 |
|---|
| x2-y2 | 5.814 |
|---|
| xy | 0.752 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.414 |
-0.014 |
0.000 |
| y |
-0.014 |
9.709 |
0.000 |
| z |
0.000 |
0.000 |
9.589 |
<r2> (average value of r
2) Å
2
| <r2> |
158.619 |
| (<r2>)1/2 |
12.594 |