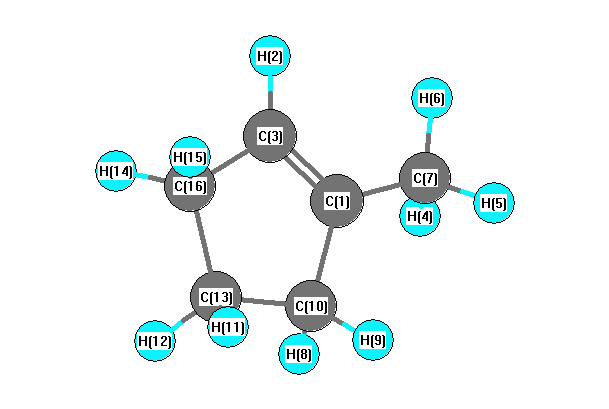
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3207 |
3073 |
17.70 |
|
|
|
| 2 |
A |
3146 |
3015 |
22.09 |
|
|
|
| 3 |
A |
3133 |
3003 |
40.74 |
|
|
|
| 4 |
A |
3098 |
2969 |
19.27 |
|
|
|
| 5 |
A |
3091 |
2962 |
55.38 |
|
|
|
| 6 |
A |
3081 |
2953 |
31.98 |
|
|
|
| 7 |
A |
3070 |
2942 |
10.51 |
|
|
|
| 8 |
A |
3038 |
2912 |
47.76 |
|
|
|
| 9 |
A |
3030 |
2903 |
57.25 |
|
|
|
| 10 |
A |
3026 |
2900 |
36.79 |
|
|
|
| 11 |
A |
1741 |
1669 |
4.05 |
|
|
|
| 12 |
A |
1497 |
1435 |
2.14 |
|
|
|
| 13 |
A |
1476 |
1414 |
4.24 |
|
|
|
| 14 |
A |
1470 |
1408 |
6.17 |
|
|
|
| 15 |
A |
1467 |
1406 |
1.58 |
|
|
|
| 16 |
A |
1458 |
1397 |
8.66 |
|
|
|
| 17 |
A |
1402 |
1344 |
4.09 |
|
|
|
| 18 |
A |
1363 |
1306 |
1.12 |
|
|
|
| 19 |
A |
1328 |
1272 |
1.09 |
|
|
|
| 20 |
A |
1320 |
1265 |
1.52 |
|
|
|
| 21 |
A |
1270 |
1217 |
2.07 |
|
|
|
| 22 |
A |
1251 |
1199 |
0.73 |
|
|
|
| 23 |
A |
1224 |
1173 |
1.66 |
|
|
|
| 24 |
A |
1179 |
1130 |
0.40 |
|
|
|
| 25 |
A |
1148 |
1100 |
1.49 |
|
|
|
| 26 |
A |
1078 |
1033 |
1.57 |
|
|
|
| 27 |
A |
1055 |
1011 |
2.54 |
|
|
|
| 28 |
A |
1024 |
981 |
7.09 |
|
|
|
| 29 |
A |
1012 |
970 |
6.07 |
|
|
|
| 30 |
A |
950 |
911 |
2.23 |
|
|
|
| 31 |
A |
930 |
891 |
0.99 |
|
|
|
| 32 |
A |
905 |
867 |
0.21 |
|
|
|
| 33 |
A |
884 |
847 |
0.54 |
|
|
|
| 34 |
A |
831 |
797 |
5.52 |
|
|
|
| 35 |
A |
799 |
766 |
11.52 |
|
|
|
| 36 |
A |
657 |
630 |
1.70 |
|
|
|
| 37 |
A |
584 |
560 |
0.06 |
|
|
|
| 38 |
A |
442 |
424 |
4.47 |
|
|
|
| 39 |
A |
320 |
307 |
0.68 |
|
|
|
| 40 |
A |
242 |
232 |
2.62 |
|
|
|
| 41 |
A |
183 |
175 |
0.74 |
|
|
|
| 42 |
A |
129 |
123 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31770.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 30445.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.137 |
|
|
|
| 2 |
H |
-0.569 |
|
|
|
| 3 |
C |
0.327 |
|
|
|
| 4 |
H |
0.001 |
|
|
|
| 5 |
H |
0.005 |
|
|
|
| 6 |
H |
-0.195 |
|
|
|
| 7 |
C |
0.367 |
|
|
|
| 8 |
H |
-0.139 |
|
|
|
| 9 |
H |
-0.185 |
|
|
|
| 10 |
C |
0.206 |
|
|
|
| 11 |
H |
-0.254 |
|
|
|
| 12 |
H |
-0.210 |
|
|
|
| 13 |
C |
0.650 |
|
|
|
| 14 |
H |
-0.168 |
|
|
|
| 15 |
H |
-0.150 |
|
|
|
| 16 |
C |
0.452 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.128 |
-0.220 |
0.073 |
0.264 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.587 |
0.143 |
0.068 |
| y |
0.143 |
-37.782 |
-0.039 |
| z |
0.068 |
-0.039 |
-38.993 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.801 |
0.143 |
0.068 |
| y |
0.143 |
0.508 |
-0.039 |
| z |
0.068 |
-0.039 |
-1.309 |
|
| Polar |
|---|
| 3z2-r2 | -2.617 |
|---|
| x2-y2 | 0.195 |
|---|
| xy | 0.143 |
|---|
| xz | 0.068 |
|---|
| yz | -0.039 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.375 |
0.814 |
0.040 |
| y |
0.814 |
11.206 |
-0.084 |
| z |
0.040 |
-0.084 |
8.456 |
<r2> (average value of r
2) Å
2
| <r2> |
170.220 |
| (<r2>)1/2 |
13.047 |