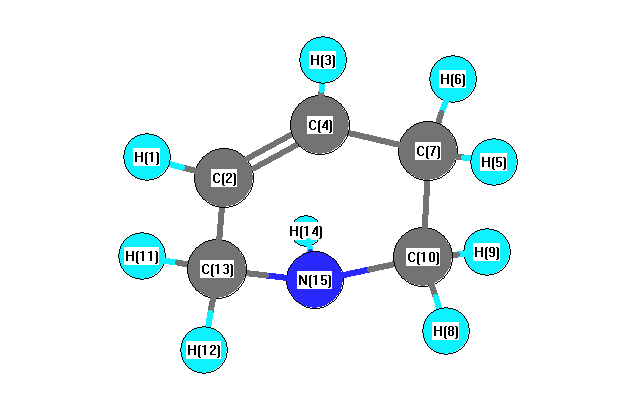
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3742 |
3407 |
0.99 |
|
|
|
| 2 |
A |
3321 |
3024 |
37.46 |
|
|
|
| 3 |
A |
3295 |
3000 |
14.69 |
|
|
|
| 4 |
A |
3225 |
2937 |
60.06 |
|
|
|
| 5 |
A |
3206 |
2919 |
38.30 |
|
|
|
| 6 |
A |
3191 |
2905 |
81.14 |
|
|
|
| 7 |
A |
3172 |
2889 |
19.40 |
|
|
|
| 8 |
A |
3154 |
2872 |
55.82 |
|
|
|
| 9 |
A |
3144 |
2863 |
45.88 |
|
|
|
| 10 |
A |
1856 |
1690 |
3.51 |
|
|
|
| 11 |
A |
1623 |
1478 |
11.26 |
|
|
|
| 12 |
A |
1606 |
1463 |
1.93 |
|
|
|
| 13 |
A |
1594 |
1451 |
3.73 |
|
|
|
| 14 |
A |
1586 |
1444 |
2.76 |
|
|
|
| 15 |
A |
1528 |
1391 |
2.12 |
|
|
|
| 16 |
A |
1514 |
1379 |
11.71 |
|
|
|
| 17 |
A |
1484 |
1351 |
3.88 |
|
|
|
| 18 |
A |
1455 |
1325 |
0.18 |
|
|
|
| 19 |
A |
1427 |
1300 |
1.45 |
|
|
|
| 20 |
A |
1353 |
1232 |
4.56 |
|
|
|
| 21 |
A |
1303 |
1186 |
19.12 |
|
|
|
| 22 |
A |
1286 |
1171 |
6.40 |
|
|
|
| 23 |
A |
1224 |
1115 |
16.38 |
|
|
|
| 24 |
A |
1200 |
1093 |
2.11 |
|
|
|
| 25 |
A |
1118 |
1018 |
4.06 |
|
|
|
| 26 |
A |
1095 |
997 |
0.74 |
|
|
|
| 27 |
A |
1084 |
987 |
9.61 |
|
|
|
| 28 |
A |
1057 |
963 |
10.32 |
|
|
|
| 29 |
A |
978 |
891 |
13.19 |
|
|
|
| 30 |
A |
951 |
866 |
7.32 |
|
|
|
| 31 |
A |
911 |
829 |
21.80 |
|
|
|
| 32 |
A |
870 |
793 |
68.03 |
|
|
|
| 33 |
A |
813 |
741 |
42.20 |
|
|
|
| 34 |
A |
709 |
646 |
25.92 |
|
|
|
| 35 |
A |
564 |
513 |
0.98 |
|
|
|
| 36 |
A |
519 |
473 |
0.55 |
|
|
|
| 37 |
A |
432 |
393 |
0.67 |
|
|
|
| 38 |
A |
310 |
282 |
7.90 |
|
|
|
| 39 |
A |
186 |
169 |
1.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31541.8 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 28722.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.379 |
|
|
|
| 2 |
C |
0.170 |
|
|
|
| 3 |
H |
-0.407 |
|
|
|
| 4 |
C |
0.025 |
|
|
|
| 5 |
H |
-0.023 |
|
|
|
| 6 |
H |
-0.010 |
|
|
|
| 7 |
C |
0.198 |
|
|
|
| 8 |
H |
-0.163 |
|
|
|
| 9 |
H |
-0.153 |
|
|
|
| 10 |
C |
0.780 |
|
|
|
| 11 |
H |
-0.062 |
|
|
|
| 12 |
H |
-0.054 |
|
|
|
| 13 |
C |
0.673 |
|
|
|
| 14 |
H |
-0.033 |
|
|
|
| 15 |
N |
-0.564 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.523 |
0.780 |
-0.464 |
1.047 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.942 |
-1.941 |
0.183 |
| y |
-1.941 |
-41.036 |
0.903 |
| z |
0.183 |
0.903 |
-37.531 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.342 |
-1.941 |
0.183 |
| y |
-1.941 |
-4.299 |
0.903 |
| z |
0.183 |
0.903 |
0.958 |
|
| Polar |
|---|
| 3z2-r2 | 1.916 |
|---|
| x2-y2 | 5.094 |
|---|
| xy | -1.941 |
|---|
| xz | 0.183 |
|---|
| yz | 0.903 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
146.940 |
| (<r2>)1/2 |
12.122 |