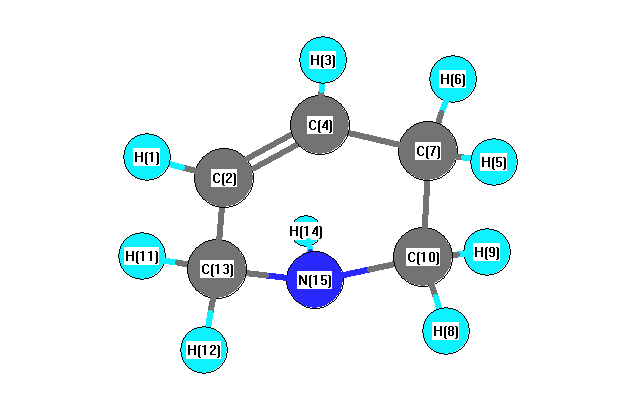
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3544 |
3396 |
0.55 |
|
|
|
| 2 |
A |
3194 |
3061 |
33.33 |
|
|
|
| 3 |
A |
3170 |
3038 |
9.71 |
|
|
|
| 4 |
A |
3114 |
2984 |
38.07 |
|
|
|
| 5 |
A |
3086 |
2957 |
23.34 |
|
|
|
| 6 |
A |
3075 |
2947 |
48.65 |
|
|
|
| 7 |
A |
3054 |
2927 |
23.57 |
|
|
|
| 8 |
A |
3027 |
2901 |
37.06 |
|
|
|
| 9 |
A |
3016 |
2890 |
44.23 |
|
|
|
| 10 |
A |
1738 |
1665 |
3.60 |
|
|
|
| 11 |
A |
1492 |
1430 |
7.74 |
|
|
|
| 12 |
A |
1477 |
1415 |
3.66 |
|
|
|
| 13 |
A |
1462 |
1401 |
11.55 |
|
|
|
| 14 |
A |
1458 |
1397 |
1.73 |
|
|
|
| 15 |
A |
1415 |
1356 |
0.76 |
|
|
|
| 16 |
A |
1387 |
1329 |
5.38 |
|
|
|
| 17 |
A |
1363 |
1307 |
4.65 |
|
|
|
| 18 |
A |
1338 |
1283 |
0.22 |
|
|
|
| 19 |
A |
1321 |
1266 |
0.48 |
|
|
|
| 20 |
A |
1252 |
1200 |
4.22 |
|
|
|
| 21 |
A |
1221 |
1171 |
16.52 |
|
|
|
| 22 |
A |
1190 |
1140 |
6.01 |
|
|
|
| 23 |
A |
1145 |
1097 |
11.44 |
|
|
|
| 24 |
A |
1102 |
1057 |
1.75 |
|
|
|
| 25 |
A |
1060 |
1016 |
5.83 |
|
|
|
| 26 |
A |
1009 |
967 |
3.21 |
|
|
|
| 27 |
A |
992 |
951 |
0.14 |
|
|
|
| 28 |
A |
977 |
936 |
4.93 |
|
|
|
| 29 |
A |
932 |
893 |
6.21 |
|
|
|
| 30 |
A |
908 |
870 |
12.80 |
|
|
|
| 31 |
A |
865 |
829 |
2.66 |
|
|
|
| 32 |
A |
804 |
771 |
80.99 |
|
|
|
| 33 |
A |
752 |
721 |
42.83 |
|
|
|
| 34 |
A |
646 |
619 |
27.73 |
|
|
|
| 35 |
A |
529 |
506 |
1.24 |
|
|
|
| 36 |
A |
487 |
466 |
0.39 |
|
|
|
| 37 |
A |
400 |
384 |
0.76 |
|
|
|
| 38 |
A |
289 |
277 |
7.81 |
|
|
|
| 39 |
A |
165 |
158 |
1.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29727.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28487.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.495 |
|
|
|
| 2 |
C |
0.430 |
|
|
|
| 3 |
H |
-0.519 |
|
|
|
| 4 |
C |
0.239 |
|
|
|
| 5 |
H |
-0.133 |
|
|
|
| 6 |
H |
-0.127 |
|
|
|
| 7 |
C |
0.449 |
|
|
|
| 8 |
H |
-0.257 |
|
|
|
| 9 |
H |
-0.229 |
|
|
|
| 10 |
C |
0.719 |
|
|
|
| 11 |
H |
-0.158 |
|
|
|
| 12 |
H |
-0.153 |
|
|
|
| 13 |
C |
0.564 |
|
|
|
| 14 |
H |
-0.020 |
|
|
|
| 15 |
N |
-0.310 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.522 |
0.693 |
-0.463 |
0.984 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.747 |
-1.999 |
0.227 |
| y |
-1.999 |
-40.504 |
0.899 |
| z |
0.227 |
0.899 |
-37.108 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.059 |
-1.999 |
0.227 |
| y |
-1.999 |
-4.077 |
0.899 |
| z |
0.227 |
0.899 |
1.017 |
|
| Polar |
|---|
| 3z2-r2 | 2.035 |
|---|
| x2-y2 | 4.757 |
|---|
| xy | -1.999 |
|---|
| xz | 0.227 |
|---|
| yz | 0.899 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.755 |
0.038 |
-0.204 |
| y |
0.038 |
10.073 |
0.014 |
| z |
-0.204 |
0.014 |
8.211 |
<r2> (average value of r
2) Å
2
| <r2> |
147.067 |
| (<r2>)1/2 |
12.127 |