Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -194.332997 |
| Energy at 298.15K | |
| HF Energy | -194.332997 |
| Nuclear repulsion energy | 132.619952 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3895 |
3733 |
34.33 |
105.74 |
0.22 |
0.36 |
| 2 |
A |
3154 |
3023 |
24.18 |
36.30 |
0.75 |
0.86 |
| 3 |
A |
3133 |
3002 |
46.45 |
53.67 |
0.56 |
0.72 |
| 4 |
A |
3104 |
2974 |
18.37 |
84.15 |
0.72 |
0.83 |
| 5 |
A |
3064 |
2936 |
58.56 |
148.05 |
0.05 |
0.10 |
| 6 |
A |
3055 |
2928 |
31.33 |
166.12 |
0.02 |
0.03 |
| 7 |
A |
3044 |
2917 |
23.39 |
94.16 |
0.68 |
0.81 |
| 8 |
A |
3004 |
2879 |
64.49 |
130.81 |
0.08 |
0.14 |
| 9 |
A |
1501 |
1438 |
2.57 |
3.30 |
0.70 |
0.82 |
| 10 |
A |
1486 |
1424 |
9.34 |
1.47 |
0.75 |
0.86 |
| 11 |
A |
1472 |
1411 |
7.26 |
6.60 |
0.74 |
0.85 |
| 12 |
A |
1459 |
1398 |
2.76 |
7.18 |
0.74 |
0.85 |
| 13 |
A |
1440 |
1380 |
3.39 |
0.70 |
0.20 |
0.33 |
| 14 |
A |
1399 |
1341 |
4.71 |
0.43 |
0.74 |
0.85 |
| 15 |
A |
1362 |
1305 |
0.62 |
0.78 |
0.68 |
0.81 |
| 16 |
A |
1319 |
1264 |
22.41 |
4.85 |
0.73 |
0.85 |
| 17 |
A |
1266 |
1213 |
1.99 |
3.14 |
0.70 |
0.82 |
| 18 |
A |
1246 |
1194 |
33.03 |
1.74 |
0.22 |
0.36 |
| 19 |
A |
1157 |
1109 |
5.72 |
1.38 |
0.15 |
0.26 |
| 20 |
A |
1131 |
1084 |
28.49 |
3.84 |
0.47 |
0.64 |
| 21 |
A |
1087 |
1041 |
24.92 |
2.82 |
0.69 |
0.81 |
| 22 |
A |
994 |
953 |
46.51 |
2.82 |
0.72 |
0.84 |
| 23 |
A |
920 |
881 |
1.79 |
0.25 |
0.45 |
0.62 |
| 24 |
A |
882 |
846 |
1.41 |
11.22 |
0.09 |
0.16 |
| 25 |
A |
768 |
736 |
0.68 |
0.24 |
0.41 |
0.59 |
| 26 |
A |
476 |
456 |
7.72 |
0.23 |
0.48 |
0.65 |
| 27 |
A |
328 |
314 |
10.82 |
0.40 |
0.38 |
0.55 |
| 28 |
A |
259 |
248 |
98.85 |
1.59 |
0.72 |
0.84 |
| 29 |
A |
220 |
211 |
1.95 |
0.07 |
0.36 |
0.52 |
| 30 |
A |
139 |
133 |
7.84 |
0.15 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 23881.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 22885.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.544 |
-0.511 |
0.128 |
| C2 |
-0.631 |
0.634 |
-0.291 |
| C3 |
0.764 |
0.541 |
0.294 |
| O4 |
1.388 |
-0.632 |
-0.215 |
| H5 |
-2.540 |
-0.404 |
-0.320 |
| H6 |
-1.670 |
-0.537 |
1.219 |
| H7 |
-1.131 |
-1.476 |
-0.187 |
| H8 |
-0.546 |
0.667 |
-1.386 |
| H9 |
-1.063 |
1.597 |
0.022 |
| H10 |
0.708 |
0.501 |
1.395 |
| H11 |
1.345 |
1.436 |
0.016 |
| H12 |
2.261 |
-0.712 |
0.174 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5234 | 2.5424 | 2.9548 | 1.0971 | 1.0989 | 1.0959 | 2.1627 | 2.1646 | 2.7753 | 3.4861 | 3.8115 |
C2 | 1.5234 | | 1.5158 | 2.3848 | 2.1731 | 2.1750 | 2.1710 | 1.0993 | 1.0997 | 2.1572 | 2.1548 | 3.2246 | C3 | 2.5424 | 1.5158 | | 1.4225 | 3.4911 | 2.8186 | 2.8085 | 2.1337 | 2.1274 | 1.1041 | 1.1026 | 1.9560 | O4 | 2.9548 | 2.3848 | 1.4225 | | 3.9360 | 3.3789 | 2.6563 | 2.6073 | 3.3209 | 2.0831 | 2.0817 | 0.9597 | H5 | 1.0971 | 2.1731 | 3.4911 | 3.9360 | | 1.7736 | 1.7756 | 2.5020 | 2.5106 | 3.7830 | 4.3120 | 4.8367 | H6 | 1.0989 | 2.1750 | 2.8186 | 3.3789 | 1.7736 | | 1.7745 | 3.0825 | 2.5210 | 2.6003 | 3.7988 | 4.0717 | H7 | 1.0959 | 2.1710 | 2.8085 | 2.6563 | 1.7756 | 1.7745 | | 2.5244 | 3.0802 | 3.1287 | 3.8275 | 3.4956 | H8 | 2.1627 | 1.0993 | 2.1337 | 2.6073 | 2.5020 | 3.0825 | 2.5244 | | 1.7645 | 3.0554 | 2.4766 | 3.4954 | H9 | 2.1646 | 1.0997 | 2.1274 | 3.3209 | 2.5106 | 2.5210 | 3.0802 | 1.7645 | | 2.4944 | 2.4129 | 4.0501 | H10 | 2.7753 | 2.1572 | 1.1041 | 2.0831 | 3.7830 | 2.6003 | 3.1287 | 3.0554 | 2.4944 | | 1.7840 | 2.3186 | H11 | 3.4861 | 2.1548 | 1.1026 | 2.0817 | 4.3120 | 3.7988 | 3.8275 | 2.4766 | 2.4129 | 1.7840 | | 2.3411 | H12 | 3.8115 | 3.2246 | 1.9560 | 0.9597 | 4.8367 | 4.0717 | 3.4956 | 3.4954 | 4.0501 | 2.3186 | 2.3411 | |
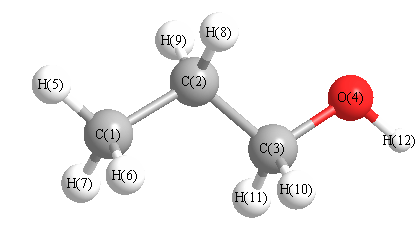 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.553 |
|
C1 |
C2 |
H8 |
110.044 |
| C1 |
C2 |
H9 |
110.170 |
|
C2 |
C1 |
H5 |
111.003 |
| C2 |
C1 |
H6 |
111.044 |
|
C2 |
C1 |
H7 |
110.904 |
| C2 |
C3 |
O4 |
108.475 |
|
C2 |
C3 |
H10 |
109.858 |
| C2 |
C3 |
H11 |
109.755 |
|
C3 |
C2 |
H8 |
108.304 |
| C3 |
C2 |
H9 |
107.795 |
|
C3 |
O4 |
H12 |
108.835 |
| O4 |
C3 |
H10 |
110.441 |
|
O4 |
C3 |
H11 |
110.417 |
| H5 |
C1 |
H6 |
107.730 |
|
H5 |
C1 |
H7 |
108.126 |
| H6 |
C1 |
H7 |
107.896 |
|
H8 |
C2 |
H9 |
106.716 |
| H10 |
C3 |
H11 |
107.891 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.412 |
|
|
|
| 2 |
C |
0.387 |
|
|
|
| 3 |
C |
0.857 |
|
|
|
| 4 |
O |
-0.498 |
|
|
|
| 5 |
H |
-0.135 |
|
|
|
| 6 |
H |
-0.133 |
|
|
|
| 7 |
H |
-0.092 |
|
|
|
| 8 |
H |
-0.223 |
|
|
|
| 9 |
H |
-0.183 |
|
|
|
| 10 |
H |
-0.261 |
|
|
|
| 11 |
H |
-0.229 |
|
|
|
| 12 |
H |
0.098 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.615 |
0.960 |
0.944 |
1.480 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.168 |
-0.127 |
1.974 |
| y |
-0.127 |
-27.408 |
-0.556 |
| z |
1.974 |
-0.556 |
-26.993 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.032 |
-0.127 |
1.974 |
| y |
-0.127 |
-2.328 |
-0.556 |
| z |
1.974 |
-0.556 |
-1.705 |
|
| Polar |
|---|
| 3z2-r2 | -3.409 |
|---|
| x2-y2 | 4.240 |
|---|
| xy | -0.127 |
|---|
| xz | 1.974 |
|---|
| yz | -0.556 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.327 |
0.088 |
0.078 |
| y |
0.088 |
6.625 |
-0.008 |
| z |
0.078 |
-0.008 |
6.219 |
<r2> (average value of r
2) Å
2
| <r2> |
95.531 |
| (<r2>)1/2 |
9.774 |
