Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -79.814223 |
| Energy at 298.15K | -79.820132 |
| HF Energy | -79.814223 |
| Nuclear repulsion energy | 42.173500 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3058 |
2931 |
0.00 |
344.46 |
0.00 |
0.01 |
| 2 |
A1g |
1411 |
1352 |
0.00 |
0.00 |
0.09 |
0.16 |
| 3 |
A1g |
1033 |
990 |
0.00 |
12.05 |
0.17 |
0.29 |
| 4 |
A1u |
313 |
300 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
3057 |
2929 |
60.64 |
0.00 |
0.00 |
0.00 |
| 6 |
A2u |
1379 |
1321 |
1.47 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3123 |
2993 |
0.00 |
123.63 |
0.75 |
0.86 |
| 7 |
Eg |
3123 |
2993 |
0.00 |
123.60 |
0.75 |
0.86 |
| 8 |
Eg |
1479 |
1417 |
0.00 |
8.84 |
0.75 |
0.86 |
| 8 |
Eg |
1479 |
1417 |
0.00 |
8.84 |
0.75 |
0.86 |
| 9 |
Eg |
1207 |
1157 |
0.00 |
0.07 |
0.75 |
0.86 |
| 9 |
Eg |
1207 |
1157 |
0.00 |
0.07 |
0.75 |
0.86 |
| 10 |
Eu |
3146 |
3015 |
62.11 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3146 |
3015 |
62.11 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1479 |
1417 |
9.58 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1479 |
1417 |
9.58 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
817 |
783 |
4.05 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
817 |
783 |
4.05 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16375.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 15692.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.761 |
| C2 |
0.000 |
0.000 |
-0.761 |
| H3 |
0.000 |
1.022 |
1.161 |
| H4 |
-0.885 |
-0.511 |
1.161 |
| H5 |
0.885 |
-0.511 |
1.161 |
| H6 |
0.000 |
-1.022 |
-1.161 |
| H7 |
-0.885 |
0.511 |
-1.161 |
| H8 |
0.885 |
0.511 |
-1.161 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5225 | 1.0977 | 1.0977 | 1.0977 | 2.1773 | 2.1773 | 2.1773 |
C2 | 1.5225 | | 2.1773 | 2.1773 | 2.1773 | 1.0977 | 1.0977 | 1.0977 | H3 | 1.0977 | 2.1773 | | 1.7707 | 1.7707 | 3.0941 | 2.5373 | 2.5373 | H4 | 1.0977 | 2.1773 | 1.7707 | | 1.7707 | 2.5373 | 2.5373 | 3.0941 | H5 | 1.0977 | 2.1773 | 1.7707 | 1.7707 | | 2.5373 | 3.0941 | 2.5373 | H6 | 2.1773 | 1.0977 | 3.0941 | 2.5373 | 2.5373 | | 1.7707 | 1.7707 | H7 | 2.1773 | 1.0977 | 2.5373 | 2.5373 | 3.0941 | 1.7707 | | 1.7707 | H8 | 2.1773 | 1.0977 | 2.5373 | 3.0941 | 2.5373 | 1.7707 | 1.7707 | |
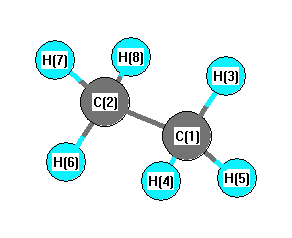 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.360 |
|
C1 |
C2 |
H7 |
111.360 |
| C1 |
C2 |
H8 |
111.360 |
|
C2 |
C1 |
H3 |
111.360 |
| C2 |
C1 |
H4 |
111.360 |
|
C2 |
C1 |
H5 |
111.360 |
| H3 |
C1 |
H4 |
107.518 |
|
H3 |
C1 |
H5 |
107.518 |
| H4 |
C1 |
H5 |
107.518 |
|
H6 |
C2 |
H7 |
107.518 |
| H6 |
C2 |
H8 |
107.518 |
|
H7 |
C2 |
H8 |
107.518 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.355 |
|
|
|
| 2 |
C |
0.355 |
|
|
|
| 3 |
H |
-0.118 |
|
|
|
| 4 |
H |
-0.118 |
|
|
|
| 5 |
H |
-0.118 |
|
|
|
| 6 |
H |
-0.118 |
|
|
|
| 7 |
H |
-0.118 |
|
|
|
| 8 |
H |
-0.118 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.875 |
0.000 |
0.000 |
| y |
0.000 |
-14.875 |
0.000 |
| z |
0.000 |
0.000 |
-15.585 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.355 |
0.000 |
0.000 |
| y |
0.000 |
0.355 |
0.000 |
| z |
0.000 |
0.000 |
-0.710 |
|
| Polar |
|---|
| 3z2-r2 | -1.420 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.076 |
0.000 |
0.000 |
| y |
0.000 |
4.076 |
0.000 |
| z |
0.000 |
0.000 |
4.651 |
<r2> (average value of r
2) Å
2
| <r2> |
30.752 |
| (<r2>)1/2 |
5.545 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -79.809724 |
| Energy at 298.15K | |
| HF Energy | -79.809724 |
| Nuclear repulsion energy | 42.015013 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3072 |
2944 |
0.00 |
|
|
|
| 2 |
A1' |
1429 |
1369 |
0.00 |
|
|
|
| 3 |
A1' |
1032 |
989 |
0.00 |
|
|
|
| 4 |
A1" |
305i |
293i |
0.00 |
|
|
|
| 5 |
A2" |
3063 |
2935 |
62.60 |
|
|
|
| 6 |
A2" |
1372 |
1314 |
1.12 |
|
|
|
| 7 |
E' |
3151 |
3020 |
51.72 |
|
|
|
| 7 |
E' |
3151 |
3020 |
51.73 |
|
|
|
| 8 |
E' |
1484 |
1422 |
11.09 |
|
|
|
| 8 |
E' |
1484 |
1422 |
11.09 |
|
|
|
| 9 |
E' |
894 |
856 |
3.02 |
|
|
|
| 9 |
E' |
894 |
856 |
3.02 |
|
|
|
| 10 |
E" |
3130 |
2999 |
0.00 |
|
|
|
| 10 |
E" |
3130 |
2999 |
0.00 |
|
|
|
| 11 |
E" |
1476 |
1414 |
0.00 |
|
|
|
| 11 |
E" |
1476 |
1414 |
0.00 |
|
|
|
| 12 |
E" |
1156 |
1107 |
0.00 |
|
|
|
| 12 |
E" |
1156 |
1107 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16120.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 15447.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.768 |
| C2 |
0.000 |
0.000 |
-0.768 |
| H3 |
0.000 |
1.017 |
1.177 |
| H4 |
-0.881 |
-0.509 |
1.177 |
| H5 |
0.881 |
-0.509 |
1.177 |
| H6 |
0.000 |
1.017 |
-1.177 |
| H7 |
0.881 |
-0.509 |
-1.177 |
| H8 |
-0.881 |
-0.509 |
-1.177 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5351 | 1.0968 | 1.0968 | 1.0968 | 2.1947 | 2.1947 | 2.1947 |
C2 | 1.5351 | | 2.1947 | 2.1947 | 2.1947 | 1.0968 | 1.0968 | 1.0968 | H3 | 1.0968 | 2.1947 | | 1.7623 | 1.7623 | 2.3541 | 2.9407 | 2.9407 | H4 | 1.0968 | 2.1947 | 1.7623 | | 1.7623 | 2.9407 | 2.9407 | 2.3541 | H5 | 1.0968 | 2.1947 | 1.7623 | 1.7623 | | 2.9407 | 2.3541 | 2.9407 | H6 | 2.1947 | 1.0968 | 2.3541 | 2.9407 | 2.9407 | | 1.7623 | 1.7623 | H7 | 2.1947 | 1.0968 | 2.9407 | 2.9407 | 2.3541 | 1.7623 | | 1.7623 | H8 | 2.1947 | 1.0968 | 2.9407 | 2.3541 | 2.9407 | 1.7623 | 1.7623 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.923 |
|
C1 |
C2 |
H7 |
111.923 |
| C1 |
C2 |
H8 |
111.923 |
|
C2 |
C1 |
H3 |
111.923 |
| C2 |
C1 |
H4 |
111.923 |
|
C2 |
C1 |
H5 |
111.923 |
| H3 |
C1 |
H4 |
106.912 |
|
H3 |
C1 |
H5 |
106.912 |
| H4 |
C1 |
H5 |
106.912 |
|
H6 |
C2 |
H7 |
106.912 |
| H6 |
C2 |
H8 |
106.912 |
|
H7 |
C2 |
H8 |
106.912 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.485 |
|
|
|
| 2 |
C |
0.485 |
|
|
|
| 3 |
H |
-0.162 |
|
|
|
| 4 |
H |
-0.162 |
|
|
|
| 5 |
H |
-0.162 |
|
|
|
| 6 |
H |
-0.162 |
|
|
|
| 7 |
H |
-0.162 |
|
|
|
| 8 |
H |
-0.162 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.879 |
0.000 |
0.000 |
| y |
0.000 |
-14.879 |
0.000 |
| z |
0.000 |
0.000 |
-15.530 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.325 |
0.000 |
0.000 |
| y |
0.000 |
0.325 |
0.000 |
| z |
0.000 |
0.000 |
-0.651 |
|
| Polar |
|---|
| 3z2-r2 | -1.302 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.046 |
0.000 |
0.000 |
| y |
0.000 |
4.046 |
0.000 |
| z |
0.000 |
0.000 |
4.666 |
<r2> (average value of r
2) Å
2
| <r2> |
31.023 |
| (<r2>)1/2 |
5.570 |
