Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -79.596584 |
| Energy at 298.15K | -79.602486 |
| HF Energy | -79.236431 |
| Nuclear repulsion energy | 41.821281 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3061 |
2990 |
|
|
|
|
| 2 |
A1g |
1477 |
1443 |
|
|
|
|
| 3 |
A1g |
1035 |
1011 |
|
|
|
|
| 4 |
A1u |
324 |
316 |
|
|
|
|
| 5 |
A2u |
3060 |
2990 |
|
|
|
|
| 6 |
A2u |
1449 |
1416 |
|
|
|
|
| 7 |
Eg |
3127 |
3055 |
|
|
|
|
| 7 |
Eg |
3127 |
3055 |
|
|
|
|
| 8 |
Eg |
1548 |
1513 |
|
|
|
|
| 8 |
Eg |
1548 |
1512 |
|
|
|
|
| 9 |
Eg |
1257 |
1228 |
|
|
|
|
| 9 |
Eg |
1257 |
1228 |
|
|
|
|
| 10 |
Eu |
3148 |
3075 |
|
|
|
|
| 10 |
Eu |
3148 |
3075 |
|
|
|
|
| 11 |
Eu |
1554 |
1518 |
|
|
|
|
| 11 |
Eu |
1553 |
1517 |
|
|
|
|
| 12 |
Eu |
839 |
820 |
|
|
|
|
| 12 |
Eu |
839 |
820 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16674.8 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 16289.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.770 |
| C2 |
0.000 |
0.000 |
-0.770 |
| H3 |
0.000 |
1.031 |
1.168 |
| H4 |
-0.893 |
-0.515 |
1.168 |
| H5 |
0.893 |
-0.515 |
1.168 |
| H6 |
0.000 |
-1.031 |
-1.168 |
| H7 |
-0.893 |
0.515 |
-1.168 |
| H8 |
0.893 |
0.515 |
-1.168 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5400 | 1.1050 | 1.1050 | 1.1050 | 2.1954 | 2.1954 | 2.1954 |
C2 | 1.5400 | | 2.1954 | 2.1954 | 2.1954 | 1.1050 | 1.1050 | 1.1050 | H3 | 1.1050 | 2.1954 | | 1.7851 | 1.7851 | 3.1161 | 2.5541 | 2.5541 | H4 | 1.1050 | 2.1954 | 1.7851 | | 1.7851 | 2.5541 | 2.5541 | 3.1161 | H5 | 1.1050 | 2.1954 | 1.7851 | 1.7851 | | 2.5541 | 3.1161 | 2.5541 | H6 | 2.1954 | 1.1050 | 3.1161 | 2.5541 | 2.5541 | | 1.7851 | 1.7851 | H7 | 2.1954 | 1.1050 | 2.5541 | 2.5541 | 3.1161 | 1.7851 | | 1.7851 | H8 | 2.1954 | 1.1050 | 2.5541 | 3.1161 | 2.5541 | 1.7851 | 1.7851 | |
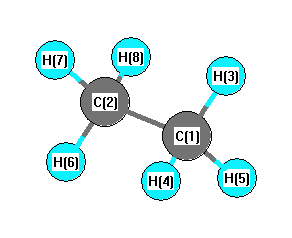 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.138 |
|
C1 |
C2 |
H7 |
111.138 |
| C1 |
C2 |
H8 |
111.138 |
|
C2 |
C1 |
H3 |
111.138 |
| C2 |
C1 |
H4 |
111.138 |
|
C2 |
C1 |
H5 |
111.138 |
| H3 |
C1 |
H4 |
107.754 |
|
H3 |
C1 |
H5 |
107.754 |
| H4 |
C1 |
H5 |
107.754 |
|
H6 |
C2 |
H7 |
107.754 |
| H6 |
C2 |
H8 |
107.754 |
|
H7 |
C2 |
H8 |
107.754 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -79.591934 |
| Energy at 298.15K | |
| HF Energy | -79.231278 |
| Nuclear repulsion energy | 41.652916 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3073 |
3002 |
|
|
|
|
| 2 |
A1' |
1496 |
1462 |
|
|
|
|
| 3 |
A1' |
1030 |
1006 |
|
|
|
|
| 4 |
A1" |
318i |
310i |
|
|
|
|
| 5 |
A2" |
3067 |
2996 |
|
|
|
|
| 6 |
A2" |
1448 |
1415 |
|
|
|
|
| 7 |
E' |
3153 |
3081 |
|
|
|
|
| 7 |
E' |
3153 |
3080 |
|
|
|
|
| 8 |
E' |
1559 |
1523 |
|
|
|
|
| 8 |
E' |
1558 |
1522 |
|
|
|
|
| 9 |
E' |
924 |
903 |
|
|
|
|
| 9 |
E' |
924 |
903 |
|
|
|
|
| 10 |
E" |
3134 |
3062 |
|
|
|
|
| 10 |
E" |
3134 |
3062 |
|
|
|
|
| 11 |
E" |
1545 |
1510 |
|
|
|
|
| 11 |
E" |
1545 |
1509 |
|
|
|
|
| 12 |
E" |
1203 |
1175 |
|
|
|
|
| 12 |
E" |
1203 |
1175 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16416.5 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 16037.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.777 |
| C2 |
0.000 |
0.000 |
-0.777 |
| H3 |
0.000 |
1.026 |
1.185 |
| H4 |
-0.889 |
-0.513 |
1.185 |
| H5 |
0.889 |
-0.513 |
1.185 |
| H6 |
0.000 |
1.026 |
-1.185 |
| H7 |
0.889 |
-0.513 |
-1.185 |
| H8 |
-0.889 |
-0.513 |
-1.185 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5536 | 1.1041 | 1.1041 | 1.1041 | 2.2137 | 2.2137 | 2.2137 |
C2 | 1.5536 | | 2.2137 | 2.2137 | 2.2137 | 1.1041 | 1.1041 | 1.1041 | H3 | 1.1041 | 2.2137 | | 1.7770 | 1.7770 | 2.3696 | 2.9619 | 2.9619 | H4 | 1.1041 | 2.2137 | 1.7770 | | 1.7770 | 2.9619 | 2.9619 | 2.3696 | H5 | 1.1041 | 2.2137 | 1.7770 | 1.7770 | | 2.9619 | 2.3696 | 2.9619 | H6 | 2.2137 | 1.1041 | 2.3696 | 2.9619 | 2.9619 | | 1.7770 | 1.7770 | H7 | 2.2137 | 1.1041 | 2.9619 | 2.9619 | 2.3696 | 1.7770 | | 1.7770 | H8 | 2.2137 | 1.1041 | 2.9619 | 2.3696 | 2.9619 | 1.7770 | 1.7770 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.686 |
|
C1 |
C2 |
H7 |
111.686 |
| C1 |
C2 |
H8 |
111.686 |
|
C2 |
C1 |
H3 |
111.686 |
| C2 |
C1 |
H4 |
111.686 |
|
C2 |
C1 |
H5 |
111.686 |
| H3 |
C1 |
H4 |
107.168 |
|
H3 |
C1 |
H5 |
107.168 |
| H4 |
C1 |
H5 |
107.168 |
|
H6 |
C2 |
H7 |
107.168 |
| H6 |
C2 |
H8 |
107.168 |
|
H7 |
C2 |
H8 |
107.168 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
