return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8 (Propane)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-118.802141
Energy at 298.15K-118.810489
HF Energy-118.275068
Nuclear repulsion energy81.826548
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3135 3062        
2 A1 3051 2981        
3 A1 3050 2979        
4 A1 1560 1524        
5 A1 1539 1503        
6 A1 1466 1432        
7 A1 1208 1180        
8 A1 900 880        
9 A1 375 366        
10 A2 3124 3052        
11 A2 1537 1501        
12 A2 1344 1313        
13 A2 929 908        
14 A2 232 226        
15 B1 3133 3060        
16 B1 3085 3014        
17 B1 1554 1518        
18 B1 1253 1224        
19 B1 763 745        
20 B1 285 278        
21 B2 3133 3060        
22 B2 3049 2978        
23 B2 1544 1508        
24 B2 1454 1421        
25 B2 1403 1370        
26 B2 1095 1070        
27 B2 949 927        

Unscaled Zero Point Vibrational Energy (zpe) 23073.1 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 22540.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.96089 0.27814 0.24596

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.595
C2 0.000 1.278 -0.263
C3 0.000 -1.278 -0.263
H4 0.887 0.000 1.258
H5 -0.887 0.000 1.258
H6 0.000 2.188 0.364
H7 0.000 -2.188 0.364
H8 0.894 1.314 -0.914
H9 -0.894 1.314 -0.914
H10 -0.894 -1.314 -0.914
H11 0.894 -1.314 -0.914

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10 H11
C11.53941.53941.10741.10742.19982.19982.19212.19212.19212.1921
C21.53942.55582.17562.17561.10523.52191.10651.10652.81822.8182
C31.53942.55582.17562.17563.52191.10522.81822.81821.10651.1065
H41.10742.17562.17561.77462.52412.52412.53893.10133.10132.5389
H51.10742.17562.17561.77462.52412.52413.10132.53892.53893.1013
H62.19981.10523.52192.52412.52414.37521.78811.78813.83383.8338
H72.19983.52191.10522.52412.52414.37523.83383.83381.78811.7881
H82.19211.10652.81822.53893.10131.78813.83381.78753.17872.6285
H92.19211.10652.81823.10132.53891.78813.83381.78752.62853.1787
H102.19212.81821.10653.10132.53893.83381.78813.17872.62851.7875
H112.19212.81821.10652.53893.10133.83381.78812.62853.17871.7875

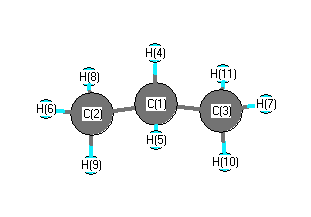
picture of Propane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H6 111.511 C1 C2 H8 110.825
C1 C2 H9 110.825 C1 C3 H7 111.511
C1 C3 H10 110.825 C1 C3 H11 110.825
C2 C1 C3 112.223 C2 C1 H4 109.487
C2 C1 H5 109.487 C3 C1 H4 109.487
C3 C1 H5 109.487 H4 C1 H5 106.506
H6 C2 H8 107.892 H6 C2 H9 107.892
H7 C3 H10 107.892 H7 C3 H11 107.892
H8 C2 H9 107.746 H10 C3 H11 107.746
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability