Jump to
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -134.272593 |
| Energy at 298.15K | -134.280909 |
| Nuclear repulsion energy | 83.043705 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3714 |
3382 |
0.97 |
|
|
|
| 2 |
A' |
3222 |
2934 |
72.88 |
|
|
|
| 3 |
A' |
3181 |
2896 |
33.69 |
|
|
|
| 4 |
A' |
3151 |
2870 |
33.35 |
|
|
|
| 5 |
A' |
1780 |
1621 |
27.38 |
|
|
|
| 6 |
A' |
1607 |
1463 |
1.48 |
|
|
|
| 7 |
A' |
1584 |
1443 |
0.65 |
|
|
|
| 8 |
A' |
1521 |
1385 |
15.01 |
|
|
|
| 9 |
A' |
1482 |
1349 |
10.56 |
|
|
|
| 10 |
A' |
1227 |
1117 |
12.35 |
|
|
|
| 11 |
A' |
1151 |
1048 |
37.49 |
|
|
|
| 12 |
A' |
953 |
867 |
21.53 |
|
|
|
| 13 |
A' |
921 |
838 |
138.67 |
|
|
|
| 14 |
A' |
431 |
392 |
6.62 |
|
|
|
| 15 |
A" |
3794 |
3455 |
1.35 |
|
|
|
| 16 |
A" |
3231 |
2942 |
98.88 |
|
|
|
| 17 |
A" |
3202 |
2915 |
0.11 |
|
|
|
| 18 |
A" |
1586 |
1444 |
6.41 |
|
|
|
| 19 |
A" |
1488 |
1355 |
0.18 |
|
|
|
| 20 |
A" |
1363 |
1241 |
0.06 |
|
|
|
| 21 |
A" |
1071 |
976 |
0.90 |
|
|
|
| 22 |
A" |
829 |
755 |
0.44 |
|
|
|
| 23 |
A" |
304 |
277 |
25.77 |
|
|
|
| 24 |
A" |
263 |
239 |
22.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21525.8 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 19601.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.299 |
-0.085 |
0.000 |
| C2 |
0.000 |
0.573 |
0.000 |
| C3 |
1.210 |
-0.359 |
0.000 |
| H4 |
2.141 |
0.212 |
0.000 |
| H5 |
1.209 |
-1.002 |
0.883 |
| H6 |
1.209 |
-1.002 |
-0.883 |
| H7 |
0.042 |
1.225 |
-0.873 |
| H8 |
0.042 |
1.225 |
0.873 |
| H9 |
-1.405 |
-0.673 |
0.806 |
| H10 |
-1.405 |
-0.673 |
-0.806 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4562 | 2.5246 | 3.4533 | 2.8133 | 2.8133 | 2.0682 | 2.0682 | 1.0035 | 1.0035 |
C2 | 1.4562 | | 1.5276 | 2.1714 | 2.1731 | 2.1731 | 1.0910 | 1.0910 | 2.0434 | 2.0434 | C3 | 2.5246 | 1.5276 | | 1.0921 | 1.0924 | 1.0924 | 2.1537 | 2.1537 | 2.7545 | 2.7545 | H4 | 3.4533 | 2.1714 | 1.0921 | | 1.7669 | 1.7669 | 2.4894 | 2.4894 | 3.7427 | 3.7427 | H5 | 2.8133 | 2.1731 | 1.0924 | 1.7669 | | 1.7667 | 3.0675 | 2.5146 | 2.6359 | 3.1299 | H6 | 2.8133 | 2.1731 | 1.0924 | 1.7669 | 1.7667 | | 2.5146 | 3.0675 | 3.1299 | 2.6359 | H7 | 2.0682 | 1.0910 | 2.1537 | 2.4894 | 3.0675 | 2.5146 | | 1.7469 | 2.9183 | 2.3875 | H8 | 2.0682 | 1.0910 | 2.1537 | 2.4894 | 2.5146 | 3.0675 | 1.7469 | | 2.3875 | 2.9183 | H9 | 1.0035 | 2.0434 | 2.7545 | 3.7427 | 2.6359 | 3.1299 | 2.9183 | 2.3875 | | 1.6122 | H10 | 1.0035 | 2.0434 | 2.7545 | 3.7427 | 3.1299 | 2.6359 | 2.3875 | 2.9183 | 1.6122 | |
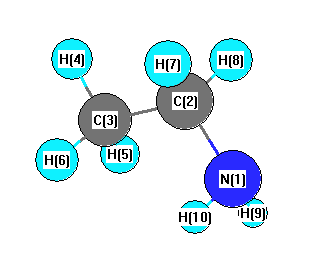 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.559 |
|
N1 |
C2 |
H7 |
107.709 |
| N1 |
C2 |
H8 |
107.709 |
|
C2 |
N1 |
H9 |
111.011 |
| C2 |
N1 |
H10 |
111.011 |
|
C2 |
C3 |
H4 |
110.874 |
| C2 |
C3 |
H5 |
110.992 |
|
C2 |
C3 |
H6 |
110.992 |
| C3 |
C2 |
H7 |
109.541 |
|
C3 |
C2 |
H8 |
109.541 |
| H4 |
C3 |
H5 |
107.965 |
|
H4 |
C3 |
H6 |
107.965 |
| H5 |
C3 |
H6 |
107.920 |
|
H7 |
C2 |
H8 |
106.376 |
| H9 |
N1 |
H10 |
106.895 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.410 |
0.000 |
|
|
| 2 |
C |
0.550 |
0.000 |
|
|
| 3 |
C |
0.328 |
0.000 |
|
|
| 4 |
H |
-0.072 |
0.000 |
|
|
| 5 |
H |
-0.062 |
0.000 |
|
|
| 6 |
H |
-0.062 |
0.000 |
|
|
| 7 |
H |
-0.123 |
0.000 |
|
|
| 8 |
H |
-0.123 |
0.000 |
|
|
| 9 |
H |
-0.013 |
0.000 |
|
|
| 10 |
H |
-0.013 |
0.000 |
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.995 |
-0.919 |
0.000 |
1.355 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.700 |
-0.041 |
-0.013 |
| y |
-0.041 |
5.138 |
0.041 |
| z |
-0.013 |
0.041 |
4.851 |
<r2> (average value of r
2) Å
2
| <r2> |
58.568 |
| (<r2>)1/2 |
7.653 |
Jump to
S1C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -134.272930 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
