Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -135.159561 |
| Energy at 298.15K | -135.167688 |
| Nuclear repulsion energy | 82.784801 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3522 |
3375 |
1.93 |
|
|
|
| 2 |
A' |
3126 |
2996 |
44.33 |
|
|
|
| 3 |
A' |
3064 |
2936 |
34.39 |
|
|
|
| 4 |
A' |
3042 |
2915 |
24.74 |
|
|
|
| 5 |
A' |
1650 |
1581 |
22.17 |
|
|
|
| 6 |
A' |
1484 |
1422 |
2.88 |
|
|
|
| 7 |
A' |
1466 |
1404 |
0.29 |
|
|
|
| 8 |
A' |
1395 |
1337 |
15.54 |
|
|
|
| 9 |
A' |
1364 |
1307 |
7.50 |
|
|
|
| 10 |
A' |
1155 |
1107 |
11.28 |
|
|
|
| 11 |
A' |
1091 |
1045 |
24.48 |
|
|
|
| 12 |
A' |
900 |
862 |
3.59 |
|
|
|
| 13 |
A' |
846 |
810 |
152.02 |
|
|
|
| 14 |
A' |
403 |
387 |
6.86 |
|
|
|
| 15 |
A" |
3607 |
3457 |
0.70 |
|
|
|
| 16 |
A" |
3131 |
3000 |
56.68 |
|
|
|
| 17 |
A" |
3100 |
2970 |
4.25 |
|
|
|
| 18 |
A" |
1470 |
1409 |
8.37 |
|
|
|
| 19 |
A" |
1383 |
1325 |
0.07 |
|
|
|
| 20 |
A" |
1261 |
1208 |
0.06 |
|
|
|
| 21 |
A" |
1002 |
960 |
0.95 |
|
|
|
| 22 |
A" |
775 |
742 |
1.28 |
|
|
|
| 23 |
A" |
280 |
268 |
24.77 |
|
|
|
| 24 |
A" |
242 |
232 |
19.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20377.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 19528.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.303 |
-0.085 |
0.000 |
| C2 |
0.000 |
0.571 |
0.000 |
| C3 |
1.211 |
-0.355 |
0.000 |
| H4 |
2.150 |
0.216 |
0.000 |
| H5 |
1.212 |
-1.003 |
0.888 |
| H6 |
1.212 |
-1.003 |
-0.888 |
| H7 |
0.037 |
1.231 |
-0.878 |
| H8 |
0.037 |
1.231 |
0.878 |
| H9 |
-1.396 |
-0.687 |
0.813 |
| H10 |
-1.396 |
-0.687 |
-0.813 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4589 | 2.5286 | 3.4662 | 2.8206 | 2.8206 | 2.0735 | 2.0735 | 1.0158 | 1.0158 |
C2 | 1.4589 | | 1.5245 | 2.1792 | 2.1758 | 2.1758 | 1.0990 | 1.0990 | 2.0477 | 2.0477 | C3 | 2.5286 | 1.5245 | | 1.0990 | 1.0991 | 1.0991 | 2.1597 | 2.1597 | 2.7509 | 2.7509 | H4 | 3.4662 | 2.1792 | 1.0990 | | 1.7759 | 1.7759 | 2.5031 | 2.5031 | 3.7485 | 3.7485 | H5 | 2.8206 | 2.1758 | 1.0991 | 1.7759 | | 1.7756 | 3.0803 | 2.5241 | 2.6280 | 3.1292 | H6 | 2.8206 | 2.1758 | 1.0991 | 1.7759 | 1.7756 | | 2.5241 | 3.0803 | 3.1292 | 2.6280 | H7 | 2.0735 | 1.0990 | 2.1597 | 2.5031 | 3.0803 | 2.5241 | | 1.7556 | 2.9314 | 2.3958 | H8 | 2.0735 | 1.0990 | 2.1597 | 2.5031 | 2.5241 | 3.0803 | 1.7556 | | 2.3958 | 2.9314 | H9 | 1.0158 | 2.0477 | 2.7509 | 3.7485 | 2.6280 | 3.1292 | 2.9314 | 2.3958 | | 1.6251 | H10 | 1.0158 | 2.0477 | 2.7509 | 3.7485 | 3.1292 | 2.6280 | 2.3958 | 2.9314 | 1.6251 | |
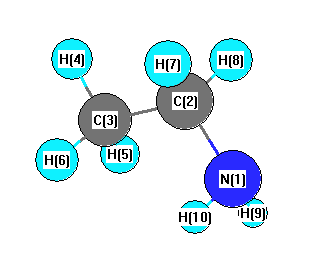 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.876 |
|
N1 |
C2 |
H7 |
107.477 |
| N1 |
C2 |
H8 |
107.477 |
|
C2 |
N1 |
H9 |
110.390 |
| C2 |
N1 |
H10 |
110.390 |
|
C2 |
C3 |
H4 |
111.296 |
| C2 |
C3 |
H5 |
111.021 |
|
C2 |
C3 |
H6 |
111.021 |
| C3 |
C2 |
H7 |
109.758 |
|
C3 |
C2 |
H8 |
109.758 |
| H4 |
C3 |
H5 |
107.791 |
|
H4 |
C3 |
H6 |
107.791 |
| H5 |
C3 |
H6 |
107.760 |
|
H7 |
C2 |
H8 |
106.017 |
| H9 |
N1 |
H10 |
106.238 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.278 |
|
|
|
| 2 |
C |
0.638 |
|
|
|
| 3 |
C |
0.448 |
|
|
|
| 4 |
H |
-0.123 |
|
|
|
| 5 |
H |
-0.121 |
|
|
|
| 6 |
H |
-0.121 |
|
|
|
| 7 |
H |
-0.202 |
|
|
|
| 8 |
H |
-0.202 |
|
|
|
| 9 |
H |
-0.020 |
|
|
|
| 10 |
H |
-0.020 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.970 |
-0.883 |
0.000 |
1.311 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.231 |
2.587 |
0.000 |
| y |
2.587 |
-20.581 |
0.000 |
| z |
0.000 |
0.000 |
-19.230 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.325 |
2.587 |
0.000 |
| y |
2.587 |
1.149 |
0.000 |
| z |
0.000 |
0.000 |
3.176 |
|
| Polar |
|---|
| 3z2-r2 | 6.352 |
|---|
| x2-y2 | -3.649 |
|---|
| xy | 2.587 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.311 |
-0.046 |
0.000 |
| y |
-0.046 |
5.571 |
0.000 |
| z |
0.000 |
0.000 |
5.173 |
<r2> (average value of r
2) Å
2
| <r2> |
58.716 |
| (<r2>)1/2 |
7.663 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -135.159636 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
