Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -478.027770 |
| Energy at 298.15K | -478.033965 |
| HF Energy | -478.027770 |
| Nuclear repulsion energy | 107.172462 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3146 |
3015 |
20.66 |
|
|
|
| 2 |
A' |
3088 |
2959 |
21.08 |
|
|
|
| 3 |
A' |
3062 |
2934 |
20.36 |
|
|
|
| 4 |
A' |
2703 |
2590 |
3.42 |
|
|
|
| 5 |
A' |
1479 |
1417 |
2.74 |
|
|
|
| 6 |
A' |
1467 |
1406 |
2.43 |
|
|
|
| 7 |
A' |
1394 |
1336 |
5.05 |
|
|
|
| 8 |
A' |
1286 |
1232 |
29.94 |
|
|
|
| 9 |
A' |
1115 |
1069 |
1.02 |
|
|
|
| 10 |
A' |
1007 |
965 |
4.44 |
|
|
|
| 11 |
A' |
855 |
819 |
0.79 |
|
|
|
| 12 |
A' |
687 |
658 |
1.58 |
|
|
|
| 13 |
A' |
303 |
291 |
2.23 |
|
|
|
| 14 |
A" |
3159 |
3028 |
23.00 |
|
|
|
| 15 |
A" |
3136 |
3006 |
0.54 |
|
|
|
| 16 |
A" |
1468 |
1407 |
9.46 |
|
|
|
| 17 |
A" |
1260 |
1207 |
0.48 |
|
|
|
| 18 |
A" |
1038 |
994 |
0.31 |
|
|
|
| 19 |
A" |
784 |
751 |
3.44 |
|
|
|
| 20 |
A" |
251 |
240 |
0.46 |
|
|
|
| 21 |
A" |
151 |
144 |
15.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16418.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 15733.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.514 |
0.696 |
0.000 |
| C2 |
0.000 |
0.831 |
0.000 |
| S3 |
-0.754 |
-0.841 |
0.000 |
| H4 |
1.978 |
1.691 |
0.000 |
| H5 |
1.864 |
0.157 |
0.889 |
| H6 |
1.864 |
0.157 |
-0.889 |
| H7 |
-0.337 |
1.373 |
0.891 |
| H8 |
-0.337 |
1.373 |
-0.891 |
| H9 |
-2.048 |
-0.446 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5201 | 2.7403 | 1.0973 | 1.0970 | 1.0970 | 2.1628 | 2.1628 | 3.7412 |
C2 | 1.5201 | | 1.8343 | 2.1567 | 2.1723 | 2.1723 | 1.0958 | 1.0958 | 2.4138 | S3 | 2.7403 | 1.8343 | | 3.7249 | 2.9398 | 2.9398 | 2.4227 | 2.4227 | 1.3532 | H4 | 1.0973 | 2.1567 | 3.7249 | | 1.7764 | 1.7764 | 2.5006 | 2.5006 | 4.5581 | H5 | 1.0970 | 2.1723 | 2.9398 | 1.7764 | | 1.7779 | 2.5144 | 3.0806 | 4.0573 | H6 | 1.0970 | 2.1723 | 2.9398 | 1.7764 | 1.7779 | | 3.0806 | 2.5144 | 4.0573 | H7 | 2.1628 | 1.0958 | 2.4227 | 2.5006 | 2.5144 | 3.0806 | | 1.7817 | 2.6515 | H8 | 2.1628 | 1.0958 | 2.4227 | 2.5006 | 3.0806 | 2.5144 | 1.7817 | | 2.6515 | H9 | 3.7412 | 2.4138 | 1.3532 | 4.5581 | 4.0573 | 4.0573 | 2.6515 | 2.6515 | |
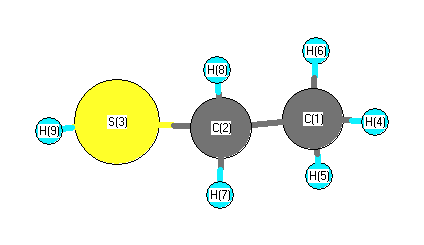 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.198 |
|
C1 |
C2 |
H7 |
110.492 |
| C1 |
C2 |
H8 |
110.492 |
|
C2 |
C1 |
H4 |
109.914 |
| C2 |
C1 |
H5 |
111.176 |
|
C2 |
C1 |
H6 |
111.176 |
| C2 |
S3 |
H9 |
97.298 |
|
S3 |
C2 |
H7 |
108.925 |
| S3 |
C2 |
H8 |
108.925 |
|
H4 |
C1 |
H5 |
108.101 |
| H4 |
C1 |
H6 |
108.101 |
|
H5 |
C1 |
H6 |
108.264 |
| H7 |
C2 |
H8 |
108.772 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.397 |
|
|
|
| 2 |
C |
0.141 |
|
|
|
| 3 |
S |
0.055 |
|
|
|
| 4 |
H |
-0.112 |
|
|
|
| 5 |
H |
-0.109 |
|
|
|
| 6 |
H |
-0.109 |
|
|
|
| 7 |
H |
-0.182 |
|
|
|
| 8 |
H |
-0.182 |
|
|
|
| 9 |
H |
0.099 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.093 |
1.721 |
0.000 |
1.723 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.005 |
-0.256 |
0.000 |
| y |
-0.256 |
-28.272 |
0.000 |
| z |
0.000 |
0.000 |
-29.082 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.672 |
-0.256 |
0.000 |
| y |
-0.256 |
-1.228 |
0.000 |
| z |
0.000 |
0.000 |
-2.443 |
|
| Polar |
|---|
| 3z2-r2 | -4.887 |
|---|
| x2-y2 | 3.266 |
|---|
| xy | -0.256 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.885 |
0.927 |
0.000 |
| y |
0.927 |
7.506 |
0.000 |
| z |
0.000 |
0.000 |
6.382 |
<r2> (average value of r
2) Å
2
| <r2> |
83.704 |
| (<r2>)1/2 |
9.149 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -478.028559 |
| Energy at 298.15K | -478.034870 |
| HF Energy | -478.028559 |
| Nuclear repulsion energy | 106.924519 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3161 |
3029 |
23.85 |
|
|
|
| 2 |
A |
3139 |
3008 |
8.51 |
|
|
|
| 3 |
A |
3131 |
3001 |
19.31 |
|
|
|
| 4 |
A |
3089 |
2960 |
13.95 |
|
|
|
| 5 |
A |
3055 |
2927 |
25.78 |
|
|
|
| 6 |
A |
2702 |
2589 |
3.31 |
|
|
|
| 7 |
A |
1474 |
1412 |
2.73 |
|
|
|
| 8 |
A |
1466 |
1405 |
10.41 |
|
|
|
| 9 |
A |
1453 |
1393 |
1.40 |
|
|
|
| 10 |
A |
1393 |
1334 |
5.30 |
|
|
|
| 11 |
A |
1293 |
1239 |
18.09 |
|
|
|
| 12 |
A |
1270 |
1217 |
4.05 |
|
|
|
| 13 |
A |
1116 |
1070 |
4.74 |
|
|
|
| 14 |
A |
1072 |
1027 |
1.04 |
|
|
|
| 15 |
A |
995 |
953 |
7.19 |
|
|
|
| 16 |
A |
865 |
829 |
5.53 |
|
|
|
| 17 |
A |
734 |
703 |
1.75 |
|
|
|
| 18 |
A |
670 |
642 |
3.74 |
|
|
|
| 19 |
A |
329 |
316 |
1.18 |
|
|
|
| 20 |
A |
260 |
250 |
1.58 |
|
|
|
| 21 |
A |
212 |
203 |
13.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16438.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 15752.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.639 |
-0.351 |
-0.054 |
| C2 |
0.500 |
0.642 |
0.092 |
| S3 |
-1.165 |
-0.099 |
-0.081 |
| H4 |
2.606 |
0.166 |
0.014 |
| H5 |
1.606 |
-1.109 |
0.740 |
| H6 |
1.587 |
-0.869 |
-1.019 |
| H7 |
0.552 |
1.175 |
1.049 |
| H8 |
0.537 |
1.398 |
-0.702 |
| H9 |
-1.086 |
-0.927 |
0.987 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5179 | 2.8151 | 1.0990 | 1.0973 | 1.0961 | 2.1740 | 2.1665 | 2.9729 |
C2 | 1.5179 | | 1.8309 | 2.1605 | 2.1695 | 2.1676 | 1.0962 | 1.0973 | 2.4041 | S3 | 2.8151 | 1.8309 | | 3.7815 | 3.0611 | 3.0076 | 2.4182 | 2.3505 | 1.3538 | H4 | 1.0990 | 2.1605 | 3.7815 | | 1.7751 | 1.7818 | 2.5117 | 2.5123 | 3.9714 | H5 | 1.0973 | 2.1695 | 3.0611 | 1.7751 | | 1.7745 | 2.5342 | 3.0829 | 2.7093 | H6 | 1.0961 | 2.1676 | 3.0076 | 1.7818 | 1.7745 | | 3.0859 | 2.5182 | 3.3423 | H7 | 2.1740 | 1.0962 | 2.4182 | 2.5117 | 2.5342 | 3.0859 | | 1.7652 | 2.6659 | H8 | 2.1665 | 1.0973 | 2.3505 | 2.5123 | 3.0829 | 2.5182 | 1.7652 | | 3.3008 | H9 | 2.9729 | 2.4041 | 1.3538 | 3.9714 | 2.7093 | 3.3423 | 2.6659 | 3.3008 | |
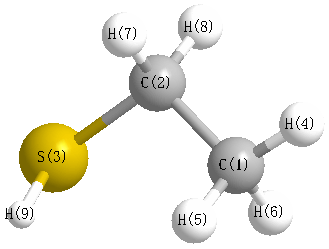 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
114.085 |
|
C1 |
C2 |
H7 |
111.514 |
| C1 |
C2 |
H8 |
110.847 |
|
C2 |
C1 |
H4 |
110.267 |
| C2 |
C1 |
H5 |
111.087 |
|
C2 |
C1 |
H6 |
111.005 |
| C2 |
S3 |
H9 |
96.888 |
|
S3 |
C2 |
H7 |
108.805 |
| S3 |
C2 |
H8 |
103.948 |
|
H4 |
C1 |
H5 |
107.842 |
| H4 |
C1 |
H6 |
108.534 |
|
H5 |
C1 |
H6 |
107.996 |
| H7 |
C2 |
H8 |
107.171 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.456 |
|
|
|
| 2 |
C |
-0.096 |
|
|
|
| 3 |
S |
0.106 |
|
|
|
| 4 |
H |
-0.125 |
|
|
|
| 5 |
H |
-0.132 |
|
|
|
| 6 |
H |
-0.091 |
|
|
|
| 7 |
H |
-0.134 |
|
|
|
| 8 |
H |
-0.085 |
|
|
|
| 9 |
H |
0.099 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.602 |
0.154 |
0.689 |
1.750 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.223 |
1.330 |
-0.655 |
| y |
1.330 |
-27.270 |
-1.730 |
| z |
-0.655 |
-1.730 |
-27.000 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.088 |
1.330 |
-0.655 |
| y |
1.330 |
0.841 |
-1.730 |
| z |
-0.655 |
-1.730 |
1.247 |
|
| Polar |
|---|
| 3z2-r2 | 2.494 |
|---|
| x2-y2 | -1.953 |
|---|
| xy | 1.330 |
|---|
| xz | -0.655 |
|---|
| yz | -1.730 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.611 |
0.227 |
-0.058 |
| y |
0.227 |
6.855 |
0.011 |
| z |
-0.058 |
0.011 |
6.419 |
<r2> (average value of r
2) Å
2
| <r2> |
84.275 |
| (<r2>)1/2 |
9.180 |
