Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.284368 |
| Energy at 298.15K | |
| HF Energy | -476.769058 |
| Nuclear repulsion energy | 106.331037 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3148 |
3076 |
|
|
|
|
| 2 |
A' |
3085 |
3014 |
|
|
|
|
| 3 |
A' |
3066 |
2995 |
|
|
|
|
| 4 |
A' |
2716 |
2653 |
|
|
|
|
| 5 |
A' |
1550 |
1514 |
|
|
|
|
| 6 |
A' |
1539 |
1504 |
|
|
|
|
| 7 |
A' |
1460 |
1427 |
|
|
|
|
| 8 |
A' |
1357 |
1325 |
|
|
|
|
| 9 |
A' |
1143 |
1116 |
|
|
|
|
| 10 |
A' |
1025 |
1001 |
|
|
|
|
| 11 |
A' |
887 |
866 |
|
|
|
|
| 12 |
A' |
699 |
683 |
|
|
|
|
| 13 |
A' |
311 |
304 |
|
|
|
|
| 14 |
A" |
3157 |
3084 |
|
|
|
|
| 15 |
A" |
3133 |
3061 |
|
|
|
|
| 16 |
A" |
1539 |
1504 |
|
|
|
|
| 17 |
A" |
1305 |
1274 |
|
|
|
|
| 18 |
A" |
1088 |
1063 |
|
|
|
|
| 19 |
A" |
810 |
791 |
|
|
|
|
| 20 |
A" |
266 |
260 |
|
|
|
|
| 21 |
A" |
177 |
173 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16729.9 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 16343.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.528 |
0.693 |
0.000 |
| C2 |
0.000 |
0.844 |
0.000 |
| S3 |
-0.762 |
-0.846 |
0.000 |
| H4 |
2.007 |
1.688 |
0.000 |
| H5 |
1.870 |
0.144 |
0.896 |
| H6 |
1.870 |
0.144 |
-0.896 |
| H7 |
-0.337 |
1.388 |
0.898 |
| H8 |
-0.337 |
1.388 |
-0.898 |
| H9 |
-2.055 |
-0.440 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5356 | 2.7590 | 1.1047 | 1.1046 | 1.1046 | 2.1835 | 2.1835 | 3.7579 |
C2 | 1.5356 | | 1.8534 | 2.1777 | 2.1880 | 2.1880 | 1.1030 | 1.1030 | 2.4230 | S3 | 2.7590 | 1.8534 | | 3.7537 | 2.9505 | 2.9505 | 2.4452 | 2.4452 | 1.3552 | H4 | 1.1047 | 2.1777 | 3.7537 | | 1.7907 | 1.7907 | 2.5280 | 2.5280 | 4.5860 | H5 | 1.1046 | 2.1880 | 2.9505 | 1.7907 | | 1.7913 | 2.5329 | 3.1038 | 4.0675 | H6 | 1.1046 | 2.1880 | 2.9505 | 1.7907 | 1.7913 | | 3.1038 | 2.5329 | 4.0675 | H7 | 2.1835 | 1.1030 | 2.4452 | 2.5280 | 2.5329 | 3.1038 | | 1.7965 | 2.6650 | H8 | 2.1835 | 1.1030 | 2.4452 | 2.5280 | 3.1038 | 2.5329 | 1.7965 | | 2.6650 | H9 | 3.7579 | 2.4230 | 1.3552 | 4.5860 | 4.0675 | 4.0675 | 2.6650 | 2.6650 | |
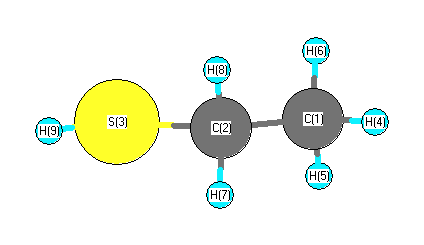 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.631 |
|
C1 |
C2 |
H7 |
110.619 |
| C1 |
C2 |
H8 |
110.619 |
|
C2 |
C1 |
H4 |
110.067 |
| C2 |
C1 |
H5 |
110.877 |
|
C2 |
C1 |
H6 |
110.877 |
| C2 |
S3 |
H9 |
96.849 |
|
S3 |
C2 |
H7 |
108.941 |
| S3 |
C2 |
H8 |
108.941 |
|
H4 |
C1 |
H5 |
108.294 |
| H4 |
C1 |
H6 |
108.294 |
|
H5 |
C1 |
H6 |
108.345 |
| H7 |
C2 |
H8 |
109.054 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.285230 |
| Energy at 298.15K | -477.291506 |
| HF Energy | -476.769554 |
| Nuclear repulsion energy | 106.098289 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3158 |
3085 |
|
|
|
|
| 2 |
A |
3138 |
3066 |
|
|
|
|
| 3 |
A |
3129 |
3057 |
|
|
|
|
| 4 |
A |
3085 |
3014 |
|
|
|
|
| 5 |
A |
3058 |
2987 |
|
|
|
|
| 6 |
A |
2709 |
2646 |
|
|
|
|
| 7 |
A |
1544 |
1509 |
|
|
|
|
| 8 |
A |
1537 |
1502 |
|
|
|
|
| 9 |
A |
1526 |
1491 |
|
|
|
|
| 10 |
A |
1458 |
1424 |
|
|
|
|
| 11 |
A |
1361 |
1330 |
|
|
|
|
| 12 |
A |
1316 |
1285 |
|
|
|
|
| 13 |
A |
1160 |
1133 |
|
|
|
|
| 14 |
A |
1099 |
1074 |
|
|
|
|
| 15 |
A |
1018 |
994 |
|
|
|
|
| 16 |
A |
897 |
876 |
|
|
|
|
| 17 |
A |
759 |
741 |
|
|
|
|
| 18 |
A |
687 |
672 |
|
|
|
|
| 19 |
A |
337 |
329 |
|
|
|
|
| 20 |
A |
273 |
267 |
|
|
|
|
| 21 |
A |
226 |
220 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16737.6 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 16351.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.648 |
-0.358 |
-0.053 |
| C2 |
0.506 |
0.656 |
0.090 |
| S3 |
-1.173 |
-0.102 |
-0.081 |
| H4 |
2.627 |
0.154 |
0.010 |
| H5 |
1.604 |
-1.115 |
0.751 |
| H6 |
1.584 |
-0.883 |
-1.022 |
| H7 |
0.558 |
1.191 |
1.054 |
| H8 |
0.547 |
1.412 |
-0.715 |
| H9 |
-1.075 |
-0.915 |
1.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5344 | 2.8327 | 1.1063 | 1.1051 | 1.1038 | 2.1939 | 2.1869 | 2.9727 |
C2 | 1.5344 | | 1.8496 | 2.1816 | 2.1860 | 2.1838 | 1.1034 | 1.1047 | 2.4071 | S3 | 2.8327 | 1.8496 | | 3.8095 | 3.0704 | 3.0162 | 2.4407 | 2.3767 | 1.3562 | H4 | 1.1063 | 2.1816 | 3.8095 | | 1.7904 | 1.7962 | 2.5389 | 2.5368 | 3.9786 | H5 | 1.1051 | 2.1860 | 3.0704 | 1.7904 | | 1.7881 | 2.5502 | 3.1062 | 2.6980 | H6 | 1.1038 | 2.1838 | 3.0162 | 1.7962 | 1.7881 | | 3.1086 | 2.5368 | 3.3414 | H7 | 2.1939 | 1.1034 | 2.4407 | 2.5389 | 2.5502 | 3.1086 | | 1.7821 | 2.6657 | H8 | 2.1869 | 1.1047 | 2.3767 | 2.5368 | 3.1062 | 2.5368 | 1.7821 | | 3.3140 | H9 | 2.9727 | 2.4071 | 1.3562 | 3.9786 | 2.6980 | 3.3414 | 2.6657 | 3.3140 | |
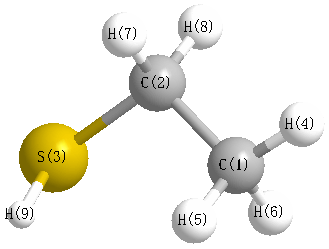 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.342 |
|
C1 |
C2 |
H7 |
111.509 |
| C1 |
C2 |
H8 |
110.869 |
|
C2 |
C1 |
H4 |
110.357 |
| C2 |
C1 |
H5 |
110.776 |
|
C2 |
C1 |
H6 |
110.684 |
| C2 |
S3 |
H9 |
96.108 |
|
S3 |
C2 |
H7 |
108.845 |
| S3 |
C2 |
H8 |
104.271 |
|
H4 |
C1 |
H5 |
108.115 |
| H4 |
C1 |
H6 |
108.728 |
|
H5 |
C1 |
H6 |
108.097 |
| H7 |
C2 |
H8 |
107.617 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
