Jump to
S1C2
Energy calculated at QCISD(T)/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -169.510686 |
| Energy at 298.15K | |
| HF Energy | -168.961275 |
| Nuclear repulsion energy | 70.489542 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3729 |
3646 |
|
|
|
|
| 2 |
A' |
3577 |
3497 |
|
|
|
|
| 3 |
A' |
2991 |
2924 |
|
|
|
|
| 4 |
A' |
1753 |
1714 |
|
|
|
|
| 5 |
A' |
1615 |
1580 |
|
|
|
|
| 6 |
A' |
1393 |
1362 |
|
|
|
|
| 7 |
A' |
1260 |
1233 |
|
|
|
|
| 8 |
A' |
1041 |
1018 |
|
|
|
|
| 9 |
A' |
553 |
540 |
|
|
|
|
| 10 |
A" |
1020 |
997 |
|
|
|
|
| 11 |
A" |
623 |
610 |
|
|
|
|
| 12 |
A" |
43i |
42i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9756.1 cm
-1
Scaled (by 0.9778) Zero Point Vibrational Energy (zpe) 9539.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.426 |
0.000 |
| O2 |
1.212 |
0.228 |
0.000 |
| N3 |
-0.951 |
-0.564 |
0.000 |
| H4 |
-0.445 |
1.447 |
0.000 |
| H5 |
-0.657 |
-1.534 |
0.000 |
| H6 |
-1.938 |
-0.340 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2282 | 1.3723 | 1.1140 | 2.0670 | 2.0831 |
O2 | 1.2282 | | 2.3033 | 2.0574 | 2.5685 | 3.2003 | N3 | 1.3723 | 2.3033 | | 2.0732 | 1.0141 | 1.0117 | H4 | 1.1140 | 2.0574 | 2.0732 | | 2.9885 | 2.3276 | H5 | 2.0670 | 2.5685 | 1.0141 | 2.9885 | | 1.7515 | H6 | 2.0831 | 3.2003 | 1.0117 | 2.3276 | 1.7515 | |
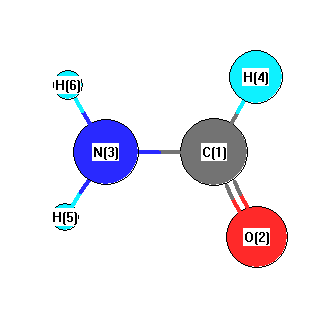 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.268 |
|
C1 |
N3 |
H6 |
121.059 |
| O2 |
C1 |
N3 |
124.588 |
|
O2 |
C1 |
H4 |
122.835 |
| N3 |
C1 |
H4 |
112.576 |
|
H5 |
N3 |
H6 |
119.673 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -169.510687 |
| Energy at 298.15K | -169.514200 |
| HF Energy | -168.961253 |
| Nuclear repulsion energy | 70.484383 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3726 |
3644 |
|
|
|
|
| 2 |
A |
3576 |
3497 |
|
|
|
|
| 3 |
A |
2992 |
2925 |
|
|
|
|
| 4 |
A |
1753 |
1714 |
|
|
|
|
| 5 |
A |
1615 |
1579 |
|
|
|
|
| 6 |
A |
1394 |
1363 |
|
|
|
|
| 7 |
A |
1260 |
1232 |
|
|
|
|
| 8 |
A |
1045 |
1022 |
|
|
|
|
| 9 |
A |
1019 |
997 |
|
|
|
|
| 10 |
A |
623 |
609 |
|
|
|
|
| 11 |
A |
553 |
541 |
|
|
|
|
| 12 |
A |
120 |
118 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9838.4 cm
-1
Scaled (by 0.9778) Zero Point Vibrational Energy (zpe) 9620.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.161 |
0.394 |
0.001 |
| O2 |
1.208 |
-0.249 |
0.003 |
| N3 |
-1.094 |
-0.161 |
-0.019 |
| H4 |
0.137 |
1.508 |
0.001 |
| H5 |
-1.187 |
-1.169 |
0.040 |
| H6 |
-1.919 |
0.419 |
0.065 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2281 | 1.3731 | 1.1139 | 2.0650 | 2.0817 |
O2 | 1.2281 | | 2.3035 | 2.0574 | 2.5655 | 3.1978 | N3 | 1.3731 | 2.3035 | | 2.0741 | 1.0142 | 1.0118 | H4 | 1.1139 | 2.0574 | 2.0741 | | 2.9869 | 2.3277 | H5 | 2.0650 | 2.5655 | 1.0142 | 2.9869 | | 1.7490 | H6 | 2.0817 | 3.1978 | 1.0118 | 2.3277 | 1.7490 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.997 |
|
C1 |
N3 |
H6 |
120.834 |
| O2 |
C1 |
N3 |
124.548 |
|
O2 |
C1 |
H4 |
122.846 |
| N3 |
C1 |
H4 |
112.600 |
|
H5 |
N3 |
H6 |
119.365 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
