return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH3 (Propane, 2-chloro-)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-577.885538
Energy at 298.15K-577.893209
HF Energy-577.206213
Nuclear repulsion energy164.470425
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3164 3091        
2 A' 3146 3073        
3 A' 3106 3035        
4 A' 3064 2993        
5 A' 1551 1515        
6 A' 1537 1501        
7 A' 1464 1431        
8 A' 1342 1311        
9 A' 1221 1193        
10 A' 1114 1089        
11 A' 924 903        
12 A' 654 639        
13 A' 432 422        
14 A' 347 339        
15 A' 282 275        
16 A" 3161 3088        
17 A" 3141 3068        
18 A" 3060 2989        
19 A" 1531 1496        
20 A" 1525 1490        
21 A" 1451 1417        
22 A" 1394 1362        
23 A" 1180 1153        
24 A" 979 957        
25 A" 960 938        
26 A" 328 321        
27 A" 255 249        

Unscaled Zero Point Vibrational Energy (zpe) 21156.3 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 20667.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.26515 0.14899 0.10494

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.906 1.016 0.000
C2 0.581 -0.062 0.000
C3 0.581 -0.903 1.277
C4 0.581 -0.903 -1.277
H5 1.428 0.645 0.000
H6 1.494 -1.528 1.305
H7 1.494 -1.528 -1.305
H8 0.565 -0.263 2.175
H9 0.565 -0.263 -2.175
H10 -0.302 -1.566 1.300
H11 -0.302 -1.566 -1.300

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
Cl11.83722.74382.74382.36333.73343.73342.92042.92042.95352.9535
C21.83721.52941.52941.10332.16462.16462.18422.18422.17512.1751
C32.74381.52942.55482.17861.10672.80951.10283.51131.10462.8037
C42.74381.52942.55482.17862.80951.10673.51131.10282.80371.1046
H52.36331.10332.17862.17862.53602.53602.50992.50993.09383.0938
H63.73342.16461.10672.80952.53602.61021.79513.81791.79663.1642
H73.73342.16462.80951.10672.53602.61023.81791.79513.16421.7966
H82.92042.18421.10283.51132.50991.79513.81794.34981.79343.8109
H92.92042.18423.51131.10282.50993.81791.79514.34983.81091.7934
H102.95352.17511.10462.80373.09381.79663.16421.79343.81092.5993
H112.95352.17512.80371.10463.09383.16421.79663.81091.79342.5993

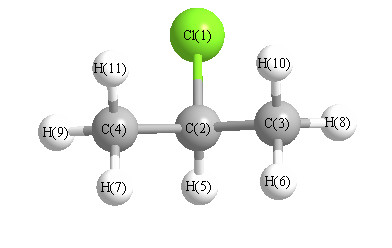
picture of Propane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 108.834 Cl1 C2 C4 108.834
Cl1 C3 H5 55.975 C2 C3 H6 109.356
C2 C3 H8 111.119 C2 C3 H10 110.293
C2 C4 H7 109.356 C2 C4 H9 111.119
C2 C4 H11 110.293 C3 C2 C4 113.280
C3 C2 H5 110.646 C4 C2 H5 110.646
H6 C3 H8 108.670 H6 C3 H10 108.677
H7 C4 H9 108.670 H7 C4 H11 108.677
H8 C3 H10 108.673 H9 C4 H11 108.673
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability