Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -233.650813 |
| Energy at 298.15K | -233.661953 |
| Nuclear repulsion energy | 199.535328 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3864 |
3703 |
15.89 |
|
|
|
| 2 |
A |
3156 |
3024 |
47.63 |
|
|
|
| 3 |
A |
3155 |
3024 |
34.67 |
|
|
|
| 4 |
A |
3149 |
3017 |
4.05 |
|
|
|
| 5 |
A |
3148 |
3016 |
6.23 |
|
|
|
| 6 |
A |
3130 |
3000 |
57.46 |
|
|
|
| 7 |
A |
3123 |
2993 |
2.28 |
|
|
|
| 8 |
A |
3065 |
2937 |
9.88 |
|
|
|
| 9 |
A |
3050 |
2923 |
30.43 |
|
|
|
| 10 |
A |
3045 |
2918 |
18.87 |
|
|
|
| 11 |
A |
1499 |
1437 |
9.45 |
|
|
|
| 12 |
A |
1491 |
1429 |
3.03 |
|
|
|
| 13 |
A |
1479 |
1417 |
2.59 |
|
|
|
| 14 |
A |
1465 |
1404 |
0.08 |
|
|
|
| 15 |
A |
1461 |
1400 |
0.07 |
|
|
|
| 16 |
A |
1451 |
1390 |
0.05 |
|
|
|
| 17 |
A |
1412 |
1353 |
7.58 |
|
|
|
| 18 |
A |
1398 |
1340 |
39.39 |
|
|
|
| 19 |
A |
1383 |
1325 |
22.78 |
|
|
|
| 20 |
A |
1356 |
1299 |
19.77 |
|
|
|
| 21 |
A |
1266 |
1213 |
17.08 |
|
|
|
| 22 |
A |
1235 |
1184 |
54.46 |
|
|
|
| 23 |
A |
1163 |
1115 |
53.77 |
|
|
|
| 24 |
A |
1039 |
995 |
0.83 |
|
|
|
| 25 |
A |
1024 |
982 |
9.11 |
|
|
|
| 26 |
A |
948 |
908 |
0.00 |
|
|
|
| 27 |
A |
945 |
905 |
32.38 |
|
|
|
| 28 |
A |
928 |
889 |
1.73 |
|
|
|
| 29 |
A |
915 |
876 |
0.20 |
|
|
|
| 30 |
A |
766 |
734 |
2.56 |
|
|
|
| 31 |
A |
467 |
448 |
8.87 |
|
|
|
| 32 |
A |
461 |
442 |
8.54 |
|
|
|
| 33 |
A |
416 |
399 |
0.43 |
|
|
|
| 34 |
A |
342 |
328 |
9.52 |
|
|
|
| 35 |
A |
341 |
326 |
1.20 |
|
|
|
| 36 |
A |
290 |
278 |
77.82 |
|
|
|
| 37 |
A |
267 |
256 |
0.10 |
|
|
|
| 38 |
A |
248 |
238 |
14.26 |
|
|
|
| 39 |
A |
186 |
178 |
3.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29763.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28522.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.006 |
-0.000 |
0.014 |
| H2 |
1.743 |
1.272 |
-0.223 |
| H3 |
0.203 |
2.154 |
-0.101 |
| H4 |
0.636 |
1.306 |
-1.607 |
| C5 |
0.683 |
1.258 |
-0.512 |
| H6 |
-1.977 |
0.886 |
0.066 |
| H7 |
-1.976 |
-0.890 |
0.065 |
| H8 |
-1.617 |
-0.001 |
-1.436 |
| C9 |
-1.484 |
-0.001 |
-0.348 |
| H10 |
0.207 |
-2.154 |
-0.101 |
| H11 |
1.745 |
-1.269 |
-0.223 |
| H12 |
0.638 |
-1.305 |
-1.607 |
| C13 |
0.685 |
-1.257 |
-0.512 |
| H14 |
0.943 |
0.001 |
1.730 |
| O15 |
0.024 |
-0.000 |
1.448 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
H4 |
C5 |
H6 |
H7 |
H8 |
C9 |
H10 |
H11 |
H12 |
C13 |
H14 |
O15 |
| C1 | | 2.1754 | 2.1672 | 2.1789 | 1.5274 | 2.1623 | 2.1623 | 2.1681 | 1.5223 | 2.1672 | 2.1754 | 2.1789 | 1.5274 | 1.9608 | 1.4339 |
H2 | 2.1754 | | 1.7788 | 1.7732 | 1.0990 | 3.7513 | 4.3112 | 3.7927 | 3.4717 | 3.7564 | 2.5411 | 3.1277 | 2.7565 | 2.4638 | 2.7138 | H3 | 2.1672 | 1.7788 | | 1.7815 | 1.0962 | 2.5275 | 3.7470 | 3.1206 | 2.7481 | 4.3077 | 3.7564 | 3.7978 | 3.4689 | 2.9223 | 2.6594 | H4 | 2.1789 | 1.7732 | 1.7815 | | 1.0976 | 3.1312 | 3.8001 | 2.6105 | 2.7914 | 3.7978 | 3.1270 | 2.6115 | 2.7876 | 3.5970 | 3.3787 | C5 | 1.5274 | 1.0990 | 1.0962 | 1.0976 | | 2.7473 | 3.4661 | 2.7801 | 2.5117 | 3.4689 | 2.7562 | 2.7879 | 2.5145 | 2.5836 | 2.4201 | H6 | 2.1623 | 3.7513 | 2.5275 | 3.1312 | 2.7473 | | 1.7762 | 1.7812 | 1.0962 | 3.7470 | 4.3112 | 3.8001 | 3.4661 | 3.4762 | 2.5885 | H7 | 2.1623 | 4.3112 | 3.7470 | 3.8001 | 3.4661 | 1.7762 | | 1.7812 | 1.0962 | 2.5276 | 3.7514 | 3.1305 | 2.7471 | 3.4764 | 2.5888 | H8 | 2.1681 | 3.7927 | 3.1206 | 2.6105 | 2.7801 | 1.7812 | 1.7812 | | 1.0966 | 3.1214 | 3.7928 | 2.6106 | 2.7805 | 4.0720 | 3.3182 | C9 | 1.5223 | 3.4717 | 2.7481 | 2.7914 | 2.5117 | 1.0962 | 1.0962 | 1.0966 | | 2.7484 | 3.4717 | 2.7912 | 2.5117 | 3.1953 | 2.3448 | H10 | 2.1672 | 3.7564 | 4.3077 | 3.7978 | 3.4689 | 3.7470 | 2.5276 | 3.1214 | 2.7484 | | 1.7788 | 1.7815 | 1.0962 | 2.9219 | 2.6592 | H11 | 2.1754 | 2.5411 | 3.7564 | 3.1270 | 2.7562 | 4.3112 | 3.7514 | 3.7928 | 3.4717 | 1.7788 | | 1.7732 | 1.0990 | 2.4640 | 2.7140 | H12 | 2.1789 | 3.1277 | 3.7978 | 2.6115 | 2.7879 | 3.8001 | 3.1305 | 2.6106 | 2.7912 | 1.7815 | 1.7732 | | 1.0976 | 3.5971 | 3.3786 | C13 | 1.5274 | 2.7565 | 3.4689 | 2.7876 | 2.5145 | 3.4661 | 2.7471 | 2.7805 | 2.5117 | 1.0962 | 1.0990 | 1.0976 | | 2.5836 | 2.4201 | H14 | 1.9608 | 2.4638 | 2.9223 | 3.5970 | 2.5836 | 3.4762 | 3.4764 | 4.0720 | 3.1953 | 2.9219 | 2.4640 | 3.5971 | 2.5836 | | 0.9617 | O15 | 1.4339 | 2.7138 | 2.6594 | 3.3787 | 2.4201 | 2.5885 | 2.5888 | 3.3182 | 2.3448 | 2.6592 | 2.7140 | 3.3786 | 2.4201 | 0.9617 | |
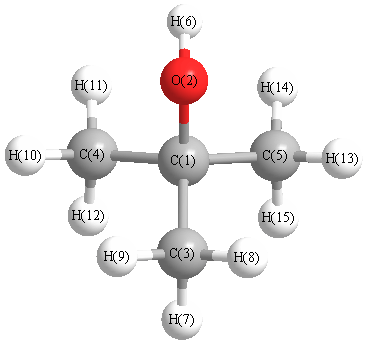 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C5 |
H2 |
110.788 |
|
C1 |
C5 |
H3 |
110.302 |
| C1 |
C5 |
H4 |
111.149 |
|
C1 |
C9 |
H6 |
110.276 |
| C1 |
C9 |
H7 |
110.279 |
|
C1 |
C9 |
H8 |
110.715 |
| C1 |
C13 |
H10 |
110.303 |
|
C1 |
C13 |
H11 |
110.788 |
| C1 |
C13 |
H12 |
111.152 |
|
C1 |
O15 |
H14 |
108.257 |
| H2 |
C5 |
H3 |
108.252 |
|
H2 |
C5 |
H4 |
107.653 |
| H3 |
C5 |
H4 |
108.595 |
|
C5 |
C1 |
C9 |
110.892 |
| C5 |
C1 |
C13 |
110.798 |
|
C5 |
C1 |
O15 |
109.579 |
| H6 |
C9 |
H7 |
108.230 |
|
H6 |
C9 |
H8 |
108.638 |
| H7 |
C9 |
H8 |
108.638 |
|
C9 |
C1 |
C13 |
110.895 |
| C9 |
C1 |
O15 |
104.931 |
|
H10 |
C13 |
H11 |
108.251 |
| H10 |
C13 |
H12 |
108.593 |
|
H11 |
C13 |
H12 |
107.652 |
| C13 |
C1 |
O15 |
109.578 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-2.040 |
|
|
|
| 2 |
H |
-0.149 |
|
|
|
| 3 |
H |
-0.069 |
|
|
|
| 4 |
H |
-0.037 |
|
|
|
| 5 |
C |
1.092 |
|
|
|
| 6 |
H |
-0.086 |
|
|
|
| 7 |
H |
-0.086 |
|
|
|
| 8 |
H |
-0.005 |
|
|
|
| 9 |
C |
0.898 |
|
|
|
| 10 |
H |
-0.069 |
|
|
|
| 11 |
H |
-0.149 |
|
|
|
| 12 |
H |
-0.037 |
|
|
|
| 13 |
C |
1.092 |
|
|
|
| 14 |
H |
0.077 |
|
|
|
| 15 |
O |
-0.435 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.147 |
0.001 |
-1.100 |
1.589 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.859 |
0.001 |
2.707 |
| y |
0.001 |
-33.722 |
0.003 |
| z |
2.707 |
0.003 |
-34.794 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.399 |
0.001 |
2.707 |
| y |
0.001 |
-0.395 |
0.003 |
| z |
2.707 |
0.003 |
-2.004 |
|
| Polar |
|---|
| 3z2-r2 | -4.007 |
|---|
| x2-y2 | 1.863 |
|---|
| xy | 0.001 |
|---|
| xz | 2.707 |
|---|
| yz | 0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.725 |
0.000 |
0.069 |
| y |
0.000 |
8.632 |
0.000 |
| z |
0.069 |
0.000 |
8.247 |
<r2> (average value of r
2) Å
2
| <r2> |
125.486 |
| (<r2>)1/2 |
11.202 |
