return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-555.707103
Energy at 298.15K-555.717827
HF Energy-554.847356
Nuclear repulsion energy242.656286
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3148 3076        
2 A' 3135 3062        
3 A' 3124 3051        
4 A' 3054 2983        
5 A' 3046 2976        
6 A' 2701 2639        
7 A' 1557 1522        
8 A' 1544 1508        
9 A' 1531 1496        
10 A' 1471 1437        
11 A' 1443 1410        
12 A' 1298 1268        
13 A' 1240 1211        
14 A' 1085 1060        
15 A' 971 948        
16 A' 896 876        
17 A' 854 834        
18 A' 608 594        
19 A' 403 393        
20 A' 376 368        
21 A' 311 304        
22 A' 292 285        
23 A" 3147 3074        
24 A" 3142 3070        
25 A" 3119 3047        
26 A" 3043 2972        
27 A" 1545 1510        
28 A" 1528 1493        
29 A" 1522 1487        
30 A" 1443 1410        
31 A" 1283 1254        
32 A" 1073 1048        
33 A" 989 966        
34 A" 962 940        
35 A" 410 400        
36 A" 311 304        
37 A" 293 287        
38 A" 262 256        
39 A" 210 205        

Unscaled Zero Point Vibrational Energy (zpe) 29185.0 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 28510.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.14751 0.09801 0.09710

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.354 -0.007 0.000
S2 -1.515 0.089 0.000
C3 0.840 1.456 0.000
C4 0.840 -0.734 1.266
C5 0.840 -0.734 -1.266
H6 -1.728 -1.252 0.000
H7 1.946 1.476 0.000
H8 0.482 1.994 -0.897
H9 0.482 1.994 0.897
H10 1.948 -0.762 1.282
H11 1.948 -0.762 -1.282
H12 0.484 -0.220 2.176
H13 0.470 -1.776 1.291
H14 0.484 -0.220 -2.176
H15 0.470 -1.776 -1.291

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.87141.54121.53921.53922.42612.17532.19602.19602.18032.18032.19082.19332.19082.1933
S21.87142.72262.79742.79741.35783.72842.90192.90193.78883.78882.97123.01372.97123.0137
C31.54122.72262.53022.53023.73181.10661.10521.10522.79122.79122.77003.50002.77003.5000
C41.53922.79742.53022.53282.90942.77723.50022.77621.10812.77931.10411.10573.49922.7861
C51.53922.79742.53022.53282.90942.77722.77623.50022.77931.10813.49922.78611.10411.1057
H62.42611.35783.73182.90942.90944.57604.02804.02803.92343.92343.27042.60213.27042.6021
H72.17533.72841.10662.77722.77724.57601.79331.79332.57912.57913.12253.79753.12253.7975
H82.19602.90191.10523.50022.77624.02801.79331.79403.80663.14473.78784.35882.55703.7904
H92.19602.90191.10522.77623.50024.02801.79331.79403.14473.80662.55703.79043.78784.3588
H102.18033.78882.79121.10812.77933.92342.57913.80663.14472.56491.79831.79233.79453.1360
H112.18033.78882.79122.77931.10813.92342.57913.14473.80662.56493.79453.13601.79831.7923
H122.19082.97122.77001.10413.49923.27043.12253.78782.55701.79833.79451.79044.35293.8005
H132.19333.01373.50001.10572.78612.60213.79754.35883.79041.79233.13601.79043.80052.5819
H142.19082.97122.77003.49921.10413.27043.12252.55703.78783.79451.79834.35293.80051.7904
H152.19333.01373.50002.78611.10572.60213.79753.79044.35883.13601.79233.80052.58191.7904

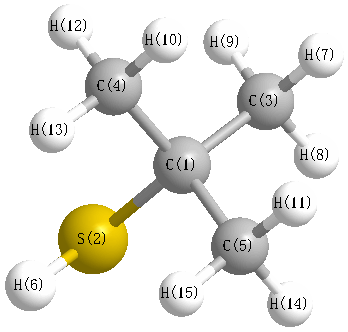
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 96.108 C1 C3 H7 109.385
C1 C3 H8 111.091 C1 C3 H9 111.091
C1 C4 H10 109.814 C1 C4 H12 110.874
C1 C4 H13 110.982 C1 C5 H11 109.814
C1 C5 H14 110.874 C1 C5 H15 110.982
S2 C1 C3 105.438 S2 C1 C4 109.828
S2 C1 C5 109.828 C3 C1 C4 110.447
C3 C1 C5 110.447 C4 C1 C5 110.723
H7 C3 H8 108.343 H7 C3 H9 108.343
H8 C3 H9 108.506 H10 C4 H12 108.756
H10 C4 H13 108.110 H11 C5 H14 108.756
H11 C5 H15 108.110 H12 C4 H13 108.225
H14 C5 H15 108.225
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability