All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)
using model chemistry: QCISD(T)=FULL/aug-cc-pVDZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at QCISD(T)=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -555.728933 |
| Energy at 298.15K | |
| HF Energy | -554.847578 |
| Nuclear repulsion energy | 242.916524 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/aug-cc-pVDZ
Geometric Data calculated at QCISD(T)=FULL/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.354 |
-0.007 |
0.000 |
| S2 |
-1.511 |
0.089 |
0.000 |
| C3 |
0.838 |
1.455 |
0.000 |
| C4 |
0.838 |
-0.734 |
1.266 |
| C5 |
0.838 |
-0.734 |
-1.266 |
| H6 |
-1.728 |
-1.251 |
0.000 |
| H7 |
1.944 |
1.476 |
0.000 |
| H8 |
0.481 |
1.992 |
-0.897 |
| H9 |
0.481 |
1.992 |
0.897 |
| H10 |
1.945 |
-0.762 |
1.283 |
| H11 |
1.945 |
-0.762 |
-1.283 |
| H12 |
0.483 |
-0.219 |
2.175 |
| H13 |
0.468 |
-1.774 |
1.291 |
| H14 |
0.483 |
-0.219 |
-2.175 |
| H15 |
0.468 |
-1.774 |
-1.291 |
Atom - Atom Distances (Å)
| |
C1 |
S2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.8678 | 1.5398 | 1.5382 | 1.5382 | 2.4258 | 2.1742 | 2.1940 | 2.1940 | 2.1795 | 2.1795 | 2.1891 | 2.1919 | 2.1891 | 2.1919 |
S2 | 1.8678 | | 2.7181 | 2.7929 | 2.7929 | 1.3572 | 3.7236 | 2.8974 | 2.8974 | 3.7841 | 3.7841 | 2.9669 | 3.0091 | 2.9669 | 3.0091 | C3 | 1.5398 | 2.7181 | | 2.5284 | 2.5284 | 3.7297 | 1.1058 | 1.1045 | 1.1045 | 2.7907 | 2.7907 | 2.7674 | 3.4975 | 2.7674 | 3.4975 | C4 | 1.5382 | 2.7929 | 2.5284 | | 2.5321 | 2.9084 | 2.7765 | 3.4975 | 2.7735 | 1.1074 | 2.7796 | 1.1034 | 1.1050 | 3.4975 | 2.7855 | C5 | 1.5382 | 2.7929 | 2.5284 | 2.5321 | | 2.9084 | 2.7765 | 2.7735 | 3.4975 | 2.7796 | 1.1074 | 3.4975 | 2.7855 | 1.1034 | 1.1050 | H6 | 2.4258 | 1.3572 | 3.7297 | 2.9084 | 2.9084 | | 4.5744 | 4.0250 | 4.0250 | 3.9219 | 3.9219 | 3.2690 | 2.6008 | 3.2690 | 2.6008 | H7 | 2.1742 | 3.7236 | 1.1058 | 2.7765 | 2.7765 | 4.5744 | | 1.7918 | 1.7918 | 2.5803 | 2.5803 | 3.1207 | 3.7963 | 3.1207 | 3.7963 | H8 | 2.1940 | 2.8974 | 1.1045 | 3.4975 | 2.7735 | 4.0250 | 1.7918 | | 1.7932 | 3.8055 | 3.1430 | 3.7844 | 4.3554 | 2.5536 | 3.7867 | H9 | 2.1940 | 2.8974 | 1.1045 | 2.7735 | 3.4975 | 4.0250 | 1.7918 | 1.7932 | | 3.1430 | 3.8055 | 2.5536 | 3.7867 | 3.7844 | 4.3554 | H10 | 2.1795 | 3.7841 | 2.7907 | 1.1074 | 2.7796 | 3.9219 | 2.5803 | 3.8055 | 3.1430 | | 2.5669 | 1.7968 | 1.7908 | 3.7940 | 3.1361 | H11 | 2.1795 | 3.7841 | 2.7907 | 2.7796 | 1.1074 | 3.9219 | 2.5803 | 3.1430 | 3.8055 | 2.5669 | | 3.7940 | 3.1361 | 1.7968 | 1.7908 | H12 | 2.1891 | 2.9669 | 2.7674 | 1.1034 | 3.4975 | 3.2690 | 3.1207 | 3.7844 | 2.5536 | 1.7968 | 3.7940 | | 1.7892 | 4.3500 | 3.7991 | H13 | 2.1919 | 3.0091 | 3.4975 | 1.1050 | 2.7855 | 2.6008 | 3.7963 | 4.3554 | 3.7867 | 1.7908 | 3.1361 | 1.7892 | | 3.7991 | 2.5821 | H14 | 2.1891 | 2.9669 | 2.7674 | 3.4975 | 1.1034 | 3.2690 | 3.1207 | 2.5536 | 3.7844 | 3.7940 | 1.7968 | 4.3500 | 3.7991 | | 1.7892 | H15 | 2.1919 | 3.0091 | 3.4975 | 2.7855 | 1.1050 | 2.6008 | 3.7963 | 3.7867 | 4.3554 | 3.1361 | 1.7908 | 3.7991 | 2.5821 | 1.7892 | |
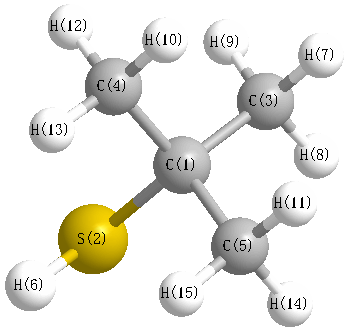 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
S2 |
H6 |
96.275 |
|
C1 |
C3 |
H7 |
109.443 |
| C1 |
C3 |
H8 |
111.070 |
|
C1 |
C3 |
H9 |
111.070 |
| C1 |
C4 |
H10 |
109.870 |
|
C1 |
C4 |
H12 |
110.862 |
| C1 |
C4 |
H13 |
110.987 |
|
C1 |
C5 |
H11 |
109.870 |
| C1 |
C5 |
H14 |
110.862 |
|
C1 |
C5 |
H15 |
110.987 |
| S2 |
C1 |
C3 |
105.412 |
|
S2 |
C1 |
C4 |
109.791 |
| S2 |
C1 |
C5 |
109.791 |
|
C3 |
C1 |
C4 |
110.459 |
| C3 |
C1 |
C5 |
110.459 |
|
C4 |
C1 |
C5 |
110.792 |
| H7 |
C3 |
H8 |
108.319 |
|
H7 |
C3 |
H9 |
108.319 |
| H8 |
C3 |
H9 |
108.537 |
|
H10 |
C4 |
H12 |
108.726 |
| H10 |
C4 |
H13 |
108.086 |
|
H11 |
C5 |
H14 |
108.726 |
| H11 |
C5 |
H15 |
108.086 |
|
H12 |
C4 |
H13 |
108.229 |
| H14 |
C5 |
H15 |
108.229 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
