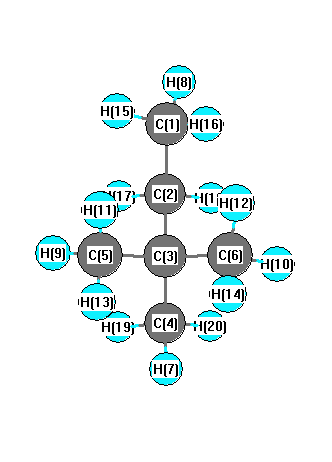
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3138 |
3007 |
52.29 |
|
|
|
| 2 |
A' |
3133 |
3002 |
55.65 |
|
|
|
| 3 |
A' |
3128 |
2997 |
70.46 |
|
|
|
| 4 |
A' |
3121 |
2991 |
0.99 |
|
|
|
| 5 |
A' |
3061 |
2933 |
22.31 |
|
|
|
| 6 |
A' |
3051 |
2924 |
13.88 |
|
|
|
| 7 |
A' |
3041 |
2914 |
46.13 |
|
|
|
| 8 |
A' |
3037 |
2910 |
24.77 |
|
|
|
| 9 |
A' |
1503 |
1441 |
13.17 |
|
|
|
| 10 |
A' |
1495 |
1433 |
7.71 |
|
|
|
| 11 |
A' |
1486 |
1424 |
5.79 |
|
|
|
| 12 |
A' |
1471 |
1410 |
0.06 |
|
|
|
| 13 |
A' |
1462 |
1401 |
0.13 |
|
|
|
| 14 |
A' |
1416 |
1357 |
6.32 |
|
|
|
| 15 |
A' |
1402 |
1343 |
5.72 |
|
|
|
| 16 |
A' |
1385 |
1327 |
10.36 |
|
|
|
| 17 |
A' |
1361 |
1305 |
0.99 |
|
|
|
| 18 |
A' |
1283 |
1229 |
2.14 |
|
|
|
| 19 |
A' |
1244 |
1192 |
5.81 |
|
|
|
| 20 |
A' |
1097 |
1051 |
0.99 |
|
|
|
| 21 |
A' |
1046 |
1002 |
3.45 |
|
|
|
| 22 |
A' |
1005 |
963 |
2.64 |
|
|
|
| 23 |
A' |
942 |
903 |
1.01 |
|
|
|
| 24 |
A' |
889 |
852 |
0.23 |
|
|
|
| 25 |
A' |
727 |
697 |
0.30 |
|
|
|
| 26 |
A' |
488 |
467 |
0.33 |
|
|
|
| 27 |
A' |
409 |
392 |
0.05 |
|
|
|
| 28 |
A' |
355 |
341 |
0.01 |
|
|
|
| 29 |
A' |
269 |
258 |
0.05 |
|
|
|
| 30 |
A' |
255 |
244 |
0.04 |
|
|
|
| 31 |
A" |
3143 |
3011 |
30.79 |
|
|
|
| 32 |
A" |
3131 |
3001 |
33.57 |
|
|
|
| 33 |
A" |
3124 |
2994 |
30.43 |
|
|
|
| 34 |
A" |
3121 |
2991 |
0.08 |
|
|
|
| 35 |
A" |
3077 |
2949 |
19.37 |
|
|
|
| 36 |
A" |
3044 |
2917 |
40.31 |
|
|
|
| 37 |
A" |
1497 |
1434 |
0.17 |
|
|
|
| 38 |
A" |
1485 |
1423 |
11.37 |
|
|
|
| 39 |
A" |
1466 |
1405 |
0.11 |
|
|
|
| 40 |
A" |
1463 |
1402 |
0.25 |
|
|
|
| 41 |
A" |
1388 |
1330 |
12.19 |
|
|
|
| 42 |
A" |
1332 |
1276 |
0.22 |
|
|
|
| 43 |
A" |
1247 |
1195 |
1.10 |
|
|
|
| 44 |
A" |
1097 |
1051 |
0.19 |
|
|
|
| 45 |
A" |
995 |
953 |
0.33 |
|
|
|
| 46 |
A" |
947 |
908 |
0.14 |
|
|
|
| 47 |
A" |
945 |
906 |
0.25 |
|
|
|
| 48 |
A" |
782 |
749 |
2.34 |
|
|
|
| 49 |
A" |
423 |
406 |
0.04 |
|
|
|
| 50 |
A" |
338 |
324 |
0.01 |
|
|
|
| 51 |
A" |
286 |
274 |
0.00 |
|
|
|
| 52 |
A" |
251 |
241 |
0.00 |
|
|
|
| 53 |
A" |
204 |
196 |
0.01 |
|
|
|
| 54 |
A" |
91 |
87 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41288.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 39566.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.521 |
|
|
|
| 2 |
C |
1.429 |
|
|
|
| 3 |
C |
-3.871 |
|
|
|
| 4 |
C |
1.410 |
|
|
|
| 5 |
C |
1.327 |
|
|
|
| 6 |
C |
1.327 |
|
|
|
| 7 |
H |
-0.132 |
|
|
|
| 8 |
H |
-0.133 |
|
|
|
| 9 |
H |
-0.095 |
|
|
|
| 10 |
H |
-0.095 |
|
|
|
| 11 |
H |
-0.161 |
|
|
|
| 12 |
H |
-0.161 |
|
|
|
| 13 |
H |
-0.112 |
|
|
|
| 14 |
H |
-0.112 |
|
|
|
| 15 |
H |
-0.129 |
|
|
|
| 16 |
H |
-0.129 |
|
|
|
| 17 |
H |
-0.342 |
|
|
|
| 18 |
H |
-0.342 |
|
|
|
| 19 |
H |
-0.100 |
|
|
|
| 20 |
H |
-0.100 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.057 |
0.011 |
0.000 |
0.058 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.009 |
0.236 |
0.000 |
| y |
0.236 |
-42.576 |
0.000 |
| z |
0.000 |
0.000 |
-42.072 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.315 |
0.236 |
0.000 |
| y |
0.236 |
-0.536 |
0.000 |
| z |
0.000 |
0.000 |
0.221 |
|
| Polar |
|---|
| 3z2-r2 | 0.442 |
|---|
| x2-y2 | 0.567 |
|---|
| xy | 0.236 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.791 |
-0.510 |
0.000 |
| y |
-0.510 |
11.398 |
0.000 |
| z |
0.000 |
0.000 |
11.003 |
<r2> (average value of r
2) Å
2
| <r2> |
195.935 |
| (<r2>)1/2 |
13.998 |