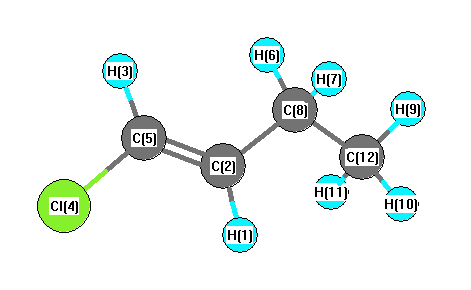
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3234 |
3099 |
8.95 |
|
|
|
| 2 |
A |
3191 |
3058 |
6.12 |
|
|
|
| 3 |
A |
3147 |
3016 |
28.80 |
|
|
|
| 4 |
A |
3141 |
3010 |
33.49 |
|
|
|
| 5 |
A |
3101 |
2971 |
8.66 |
|
|
|
| 6 |
A |
3058 |
2930 |
30.21 |
|
|
|
| 7 |
A |
3048 |
2921 |
23.93 |
|
|
|
| 8 |
A |
1718 |
1647 |
23.14 |
|
|
|
| 9 |
A |
1482 |
1420 |
6.39 |
|
|
|
| 10 |
A |
1474 |
1412 |
7.19 |
|
|
|
| 11 |
A |
1460 |
1399 |
3.78 |
|
|
|
| 12 |
A |
1393 |
1335 |
2.69 |
|
|
|
| 13 |
A |
1347 |
1291 |
1.54 |
|
|
|
| 14 |
A |
1307 |
1252 |
6.08 |
|
|
|
| 15 |
A |
1289 |
1235 |
9.82 |
|
|
|
| 16 |
A |
1241 |
1189 |
8.64 |
|
|
|
| 17 |
A |
1145 |
1097 |
0.15 |
|
|
|
| 18 |
A |
1089 |
1044 |
2.34 |
|
|
|
| 19 |
A |
1052 |
1008 |
10.95 |
|
|
|
| 20 |
A |
961 |
921 |
44.03 |
|
|
|
| 21 |
A |
919 |
881 |
7.53 |
|
|
|
| 22 |
A |
844 |
809 |
55.89 |
|
|
|
| 23 |
A |
820 |
786 |
5.24 |
|
|
|
| 24 |
A |
760 |
728 |
5.89 |
|
|
|
| 25 |
A |
443 |
424 |
1.87 |
|
|
|
| 26 |
A |
381 |
365 |
3.10 |
|
|
|
| 27 |
A |
291 |
279 |
0.29 |
|
|
|
| 28 |
A |
198 |
189 |
0.18 |
|
|
|
| 29 |
A |
166 |
159 |
0.24 |
|
|
|
| 30 |
A |
99 |
95 |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21898.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 20985.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.468 |
|
|
|
| 2 |
C |
1.713 |
|
|
|
| 3 |
H |
-0.524 |
|
|
|
| 4 |
Cl |
-0.219 |
|
|
|
| 5 |
C |
-0.308 |
|
|
|
| 6 |
H |
-0.221 |
|
|
|
| 7 |
H |
-0.111 |
|
|
|
| 8 |
C |
0.167 |
|
|
|
| 9 |
H |
-0.148 |
|
|
|
| 10 |
H |
-0.108 |
|
|
|
| 11 |
H |
-0.111 |
|
|
|
| 12 |
C |
0.338 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.075 |
0.506 |
-0.036 |
2.137 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.871 |
0.092 |
0.634 |
| y |
0.092 |
-37.952 |
-1.779 |
| z |
0.634 |
-1.779 |
-38.347 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.722 |
0.092 |
0.634 |
| y |
0.092 |
1.157 |
-1.779 |
| z |
0.634 |
-1.779 |
0.565 |
|
| Polar |
|---|
| 3z2-r2 | 1.130 |
|---|
| x2-y2 | -1.920 |
|---|
| xy | 0.092 |
|---|
| xz | 0.634 |
|---|
| yz | -1.779 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.740 |
-0.023 |
0.521 |
| y |
-0.023 |
8.244 |
-0.335 |
| z |
0.521 |
-0.335 |
7.912 |
<r2> (average value of r
2) Å
2
| <r2> |
237.633 |
| (<r2>)1/2 |
15.415 |