Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -638.666608 |
| Energy at 298.15K | -638.671479 |
| HF Energy | -638.666608 |
| Nuclear repulsion energy | 156.870242 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3122 |
2992 |
15.34 |
|
|
|
| 2 |
A' |
3093 |
2964 |
15.26 |
|
|
|
| 3 |
A' |
1493 |
1430 |
2.69 |
|
|
|
| 4 |
A' |
1472 |
1410 |
6.07 |
|
|
|
| 5 |
A' |
1396 |
1338 |
2.32 |
|
|
|
| 6 |
A' |
1260 |
1208 |
5.14 |
|
|
|
| 7 |
A' |
1099 |
1054 |
20.66 |
|
|
|
| 8 |
A' |
1053 |
1009 |
116.38 |
|
|
|
| 9 |
A' |
787 |
754 |
43.13 |
|
|
|
| 10 |
A' |
386 |
369 |
2.50 |
|
|
|
| 11 |
A' |
242 |
232 |
11.42 |
|
|
|
| 12 |
A" |
3198 |
3065 |
5.91 |
|
|
|
| 13 |
A" |
3156 |
3024 |
10.37 |
|
|
|
| 14 |
A" |
1287 |
1234 |
0.01 |
|
|
|
| 15 |
A" |
1204 |
1154 |
1.14 |
|
|
|
| 16 |
A" |
1050 |
1006 |
1.18 |
|
|
|
| 17 |
A" |
789 |
756 |
0.96 |
|
|
|
| 18 |
A" |
124 |
119 |
9.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13104.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 12557.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.620 |
0.000 |
| C2 |
0.983 |
-0.528 |
0.000 |
| Cl3 |
-1.665 |
-0.045 |
0.000 |
| F4 |
2.272 |
0.004 |
0.000 |
| H5 |
0.110 |
1.238 |
0.896 |
| H6 |
0.110 |
1.238 |
-0.896 |
| H7 |
0.871 |
-1.149 |
0.897 |
| H8 |
0.871 |
-1.149 |
-0.897 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5120 | 1.7932 | 2.3540 | 1.0936 | 1.0936 | 2.1663 | 2.1663 |
C2 | 1.5120 | | 2.6923 | 1.3937 | 2.1645 | 2.1645 | 1.0967 | 1.0967 | Cl3 | 1.7932 | 2.6923 | | 3.9372 | 2.3663 | 2.3663 | 2.9075 | 2.9075 | F4 | 2.3540 | 1.3937 | 3.9372 | | 2.6460 | 2.6460 | 2.0237 | 2.0237 | H5 | 1.0936 | 2.1645 | 2.3663 | 2.6460 | | 1.7914 | 2.5053 | 3.0807 | H6 | 1.0936 | 2.1645 | 2.3663 | 2.6460 | 1.7914 | | 3.0807 | 2.5053 | H7 | 2.1663 | 1.0967 | 2.9075 | 2.0237 | 2.5053 | 3.0807 | | 1.7942 | H8 | 2.1663 | 1.0967 | 2.9075 | 2.0237 | 3.0807 | 2.5053 | 1.7942 | |
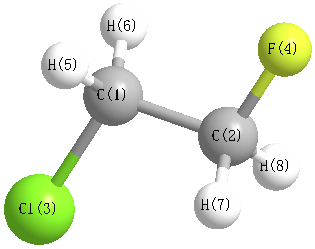 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.152 |
|
C1 |
C2 |
H7 |
111.289 |
| C1 |
C2 |
H8 |
111.289 |
|
C2 |
C1 |
Cl3 |
108.788 |
| C2 |
C1 |
H5 |
111.335 |
|
C2 |
C1 |
H6 |
111.335 |
| Cl3 |
C1 |
H5 |
107.626 |
|
Cl3 |
C1 |
H6 |
107.626 |
| F4 |
C2 |
H7 |
108.111 |
|
F4 |
C2 |
H8 |
108.111 |
| H5 |
C1 |
H6 |
109.979 |
|
H7 |
C2 |
H8 |
109.769 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.088 |
|
|
|
| 2 |
C |
1.005 |
|
|
|
| 3 |
Cl |
-0.077 |
|
|
|
| 4 |
F |
-0.447 |
|
|
|
| 5 |
H |
-0.022 |
|
|
|
| 6 |
H |
-0.022 |
|
|
|
| 7 |
H |
-0.175 |
|
|
|
| 8 |
H |
-0.175 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.024 |
0.076 |
0.000 |
0.079 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.004 |
-0.707 |
0.000 |
| y |
-0.707 |
-29.020 |
0.000 |
| z |
0.000 |
0.000 |
-29.628 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.680 |
-0.707 |
0.000 |
| y |
-0.707 |
4.796 |
0.000 |
| z |
0.000 |
0.000 |
3.884 |
|
| Polar |
|---|
| 3z2-r2 | 7.768 |
|---|
| x2-y2 | -8.984 |
|---|
| xy | -0.707 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.518 |
0.247 |
0.000 |
| y |
0.247 |
5.698 |
0.000 |
| z |
0.000 |
0.000 |
5.253 |
<r2> (average value of r
2) Å
2
| <r2> |
133.988 |
| (<r2>)1/2 |
11.575 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -638.665891 |
| Energy at 298.15K | -638.670868 |
| HF Energy | -638.665891 |
| Nuclear repulsion energy | 160.557144 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3183 |
3050 |
3.60 |
|
|
|
| 2 |
A |
3138 |
3007 |
16.25 |
|
|
|
| 3 |
A |
3110 |
2980 |
13.12 |
|
|
|
| 4 |
A |
3067 |
2939 |
25.11 |
|
|
|
| 5 |
A |
1473 |
1411 |
4.55 |
|
|
|
| 6 |
A |
1439 |
1379 |
12.35 |
|
|
|
| 7 |
A |
1405 |
1347 |
6.92 |
|
|
|
| 8 |
A |
1314 |
1260 |
26.57 |
|
|
|
| 9 |
A |
1266 |
1214 |
0.44 |
|
|
|
| 10 |
A |
1209 |
1159 |
5.14 |
|
|
|
| 11 |
A |
1119 |
1073 |
60.57 |
|
|
|
| 12 |
A |
1053 |
1009 |
30.11 |
|
|
|
| 13 |
A |
977 |
937 |
9.97 |
|
|
|
| 14 |
A |
851 |
815 |
10.00 |
|
|
|
| 15 |
A |
692 |
664 |
29.37 |
|
|
|
| 16 |
A |
463 |
443 |
12.65 |
|
|
|
| 17 |
A |
285 |
274 |
1.08 |
|
|
|
| 18 |
A |
126 |
121 |
2.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13086.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 12540.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.087 |
0.834 |
-0.299 |
| C2 |
1.195 |
0.413 |
0.366 |
| Cl3 |
-1.441 |
-0.285 |
0.064 |
| F4 |
1.689 |
-0.751 |
-0.203 |
| H5 |
-0.381 |
1.825 |
0.065 |
| H6 |
0.027 |
0.860 |
-1.387 |
| H7 |
1.951 |
1.200 |
0.222 |
| H8 |
1.052 |
0.232 |
1.439 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5043 | 1.7942 | 2.3822 | 1.0954 | 1.0942 | 2.1344 | 2.1631 |
C2 | 1.5043 | | 2.7439 | 1.3869 | 2.1371 | 2.1536 | 1.1004 | 1.0973 | Cl3 | 1.7942 | 2.7439 | | 3.1759 | 2.3607 | 2.3606 | 3.7060 | 2.8932 | F4 | 2.3822 | 1.3869 | 3.1759 | | 3.3152 | 2.5997 | 2.0145 | 2.0170 | H5 | 1.0954 | 2.1371 | 2.3607 | 3.3152 | | 1.7904 | 2.4187 | 2.5450 | H6 | 1.0942 | 2.1536 | 2.3606 | 2.5997 | 1.7904 | | 2.5310 | 3.0709 | H7 | 2.1344 | 1.1004 | 3.7060 | 2.0145 | 2.4187 | 2.5310 | | 1.7960 | H8 | 2.1631 | 1.0973 | 2.8932 | 2.0170 | 2.5450 | 3.0709 | 1.7960 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.896 |
|
C1 |
C2 |
H7 |
109.071 |
| C1 |
C2 |
H8 |
111.542 |
|
C2 |
C1 |
Cl3 |
112.284 |
| C2 |
C1 |
H5 |
109.576 |
|
C2 |
C1 |
H6 |
110.966 |
| Cl3 |
C1 |
H5 |
107.070 |
|
Cl3 |
C1 |
H6 |
107.126 |
| F4 |
C2 |
H7 |
107.619 |
|
F4 |
C2 |
H8 |
108.005 |
| H5 |
C1 |
H6 |
109.705 |
|
H7 |
C2 |
H8 |
109.620 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.089 |
|
|
|
| 2 |
C |
0.955 |
|
|
|
| 3 |
Cl |
-0.075 |
|
|
|
| 4 |
F |
-0.436 |
|
|
|
| 5 |
H |
-0.000 |
|
|
|
| 6 |
H |
-0.016 |
|
|
|
| 7 |
H |
-0.180 |
|
|
|
| 8 |
H |
-0.160 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.815 |
2.679 |
0.310 |
2.817 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.599 |
2.511 |
1.105 |
| y |
2.511 |
-29.644 |
-0.576 |
| z |
1.105 |
-0.576 |
-29.562 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.997 |
2.511 |
1.105 |
| y |
2.511 |
1.937 |
-0.576 |
| z |
1.105 |
-0.576 |
2.060 |
|
| Polar |
|---|
| 3z2-r2 | 4.119 |
|---|
| x2-y2 | -3.956 |
|---|
| xy | 2.511 |
|---|
| xz | 1.105 |
|---|
| yz | -0.576 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.068 |
0.632 |
-0.027 |
| y |
0.632 |
6.027 |
-0.083 |
| z |
-0.027 |
-0.083 |
5.453 |
<r2> (average value of r
2) Å
2
| <r2> |
117.025 |
| (<r2>)1/2 |
10.818 |
