Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -637.733458 |
| Energy at 298.15K | -637.738304 |
| HF Energy | -637.025638 |
| Nuclear repulsion energy | 155.304690 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3124 |
3051 |
|
|
|
|
| 2 |
A' |
3087 |
3015 |
|
|
|
|
| 3 |
A' |
1569 |
1533 |
|
|
|
|
| 4 |
A' |
1538 |
1502 |
|
|
|
|
| 5 |
A' |
1457 |
1424 |
|
|
|
|
| 6 |
A' |
1323 |
1292 |
|
|
|
|
| 7 |
A' |
1108 |
1082 |
|
|
|
|
| 8 |
A' |
1078 |
1053 |
|
|
|
|
| 9 |
A' |
804 |
785 |
|
|
|
|
| 10 |
A' |
386 |
377 |
|
|
|
|
| 11 |
A' |
246 |
240 |
|
|
|
|
| 12 |
A" |
3192 |
3118 |
|
|
|
|
| 13 |
A" |
3145 |
3073 |
|
|
|
|
| 14 |
A" |
1325 |
1294 |
|
|
|
|
| 15 |
A" |
1241 |
1212 |
|
|
|
|
| 16 |
A" |
1103 |
1077 |
|
|
|
|
| 17 |
A" |
811 |
793 |
|
|
|
|
| 18 |
A" |
139 |
136 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13336.7 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 13028.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.629 |
0.000 |
| C2 |
0.986 |
-0.533 |
0.000 |
| Cl3 |
-1.680 |
-0.051 |
0.000 |
| F4 |
2.296 |
0.012 |
0.000 |
| H5 |
0.109 |
1.248 |
0.904 |
| H6 |
0.109 |
1.248 |
-0.904 |
| H7 |
0.879 |
-1.154 |
0.905 |
| H8 |
0.879 |
-1.154 |
-0.905 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5246 | 1.8126 | 2.3777 | 1.1006 | 1.1006 | 2.1847 | 2.1847 |
C2 | 1.5246 | | 2.7097 | 1.4185 | 2.1812 | 2.1812 | 1.1029 | 1.1029 | Cl3 | 1.8126 | 2.7097 | | 3.9766 | 2.3886 | 2.3886 | 2.9302 | 2.9302 | F4 | 2.3777 | 1.4185 | 3.9766 | | 2.6695 | 2.6695 | 2.0459 | 2.0459 | H5 | 1.1006 | 2.1812 | 2.3886 | 2.6695 | | 1.8077 | 2.5221 | 3.1038 | H6 | 1.1006 | 2.1812 | 2.3886 | 2.6695 | 1.8077 | | 3.1038 | 2.5221 | H7 | 2.1847 | 1.1029 | 2.9302 | 2.0459 | 2.5221 | 3.1038 | | 1.8104 | H8 | 2.1847 | 1.1029 | 2.9302 | 2.0459 | 3.1038 | 2.5221 | 1.8104 | |
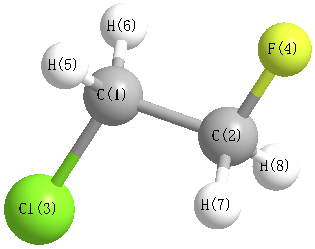 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.725 |
|
C1 |
C2 |
H7 |
111.492 |
| C1 |
C2 |
H8 |
111.492 |
|
C2 |
C1 |
Cl3 |
108.271 |
| C2 |
C1 |
H5 |
111.354 |
|
C2 |
C1 |
H6 |
111.354 |
| Cl3 |
C1 |
H5 |
107.637 |
|
Cl3 |
C1 |
H6 |
107.637 |
| F4 |
C2 |
H7 |
107.816 |
|
F4 |
C2 |
H8 |
107.816 |
| H5 |
C1 |
H6 |
110.418 |
|
H7 |
C2 |
H8 |
110.323 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -637.732297 |
| Energy at 298.15K | -637.737245 |
| HF Energy | -637.024212 |
| Nuclear repulsion energy | 159.333843 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3176 |
3103 |
|
|
|
|
| 2 |
A |
3131 |
3059 |
|
|
|
|
| 3 |
A |
3109 |
3037 |
|
|
|
|
| 4 |
A |
3064 |
2994 |
|
|
|
|
| 5 |
A |
1553 |
1517 |
|
|
|
|
| 6 |
A |
1515 |
1480 |
|
|
|
|
| 7 |
A |
1457 |
1424 |
|
|
|
|
| 8 |
A |
1381 |
1349 |
|
|
|
|
| 9 |
A |
1293 |
1263 |
|
|
|
|
| 10 |
A |
1246 |
1218 |
|
|
|
|
| 11 |
A |
1134 |
1108 |
|
|
|
|
| 12 |
A |
1096 |
1071 |
|
|
|
|
| 13 |
A |
1004 |
981 |
|
|
|
|
| 14 |
A |
879 |
859 |
|
|
|
|
| 15 |
A |
711 |
694 |
|
|
|
|
| 16 |
A |
472 |
461 |
|
|
|
|
| 17 |
A |
290 |
284 |
|
|
|
|
| 18 |
A |
139 |
136 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13324.7 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 13016.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.087 |
0.854 |
-0.305 |
| C2 |
1.203 |
0.426 |
0.372 |
| Cl3 |
-1.442 |
-0.289 |
0.066 |
| F4 |
1.681 |
-0.770 |
-0.206 |
| H5 |
-0.389 |
1.849 |
0.063 |
| H6 |
0.033 |
0.874 |
-1.399 |
| H7 |
1.978 |
1.200 |
0.216 |
| H8 |
1.058 |
0.242 |
1.450 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5186 | 1.8106 | 2.4029 | 1.1027 | 1.1013 | 2.1578 | 2.1827 |
C2 | 1.5186 | | 2.7567 | 1.4117 | 2.1576 | 2.1698 | 1.1062 | 1.1034 | Cl3 | 1.8106 | 2.7567 | | 3.1712 | 2.3830 | 2.3817 | 3.7327 | 2.9058 | F4 | 2.4029 | 1.4117 | 3.1712 | | 3.3490 | 2.6159 | 2.0362 | 2.0388 | H5 | 1.1027 | 2.1576 | 2.3830 | 3.3490 | | 1.8072 | 2.4592 | 2.5688 | H6 | 1.1013 | 2.1698 | 2.3817 | 2.6159 | 1.8072 | | 2.5493 | 3.0933 | H7 | 2.1578 | 1.1062 | 3.7327 | 2.0362 | 2.4592 | 2.5493 | | 1.8132 | H8 | 2.1827 | 1.1034 | 2.9058 | 2.0388 | 2.5688 | 3.0933 | 1.8132 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.116 |
|
C1 |
C2 |
H7 |
109.582 |
| C1 |
C2 |
H8 |
111.727 |
|
C2 |
C1 |
Cl3 |
111.498 |
| C2 |
C1 |
H5 |
109.774 |
|
C2 |
C1 |
H6 |
110.820 |
| Cl3 |
C1 |
H5 |
107.257 |
|
Cl3 |
C1 |
H6 |
107.238 |
| F4 |
C2 |
H7 |
107.320 |
|
F4 |
C2 |
H8 |
107.684 |
| H5 |
C1 |
H6 |
110.165 |
|
H7 |
C2 |
H8 |
110.299 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
