return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6O (Pentadienal)

using model chemistry: QCISD(T)/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/aug-cc-pVDZ
 hartrees
Energy at 0K-268.644261
Energy at 298.15K-268.649710
HF Energy-267.683188
Nuclear repulsion energy191.443883
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3237 3165        
2 A' 3174 3104        
3 A' 3157 3087        
4 A' 3136 3066        
5 A' 3131 3061        
6 A' 2913 2848        
7 A' 1709 1671        
8 A' 1677 1640        
9 A' 1617 1581        
10 A' 1439 1407        
11 A' 1384 1354        
12 A' 1301 1272        
13 A' 1297 1268        
14 A' 1243 1215        
15 A' 1180 1154        
16 A' 1117 1092        
17 A' 949 928        
18 A' 588 575        
19 A' 423 414        
20 A' 377 369        
21 A' 147 144        
22 A" 1014 991        
23 A" 987 966        
24 A" 958 937        
25 A" 925 905        
26 A" 841 822        
27 A" 633 618        
28 A" 277 270        
29 A" 205 200        
30 A" 95 93        

Unscaled Zero Point Vibrational Energy (zpe) 20564.5 cm-1
Scaled (by 0.9778) Zero Point Vibrational Energy (zpe) 20108.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/aug-cc-pVDZ
ABC
0.93355 0.04376 0.04180

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.132 -1.592 0.000
O2 -1.125 -2.822 0.000
C3 0.090 -0.754 0.000
C4 0.000 0.609 0.000
C5 1.157 1.513 0.000
C6 1.037 2.870 0.000
H7 -2.099 -1.025 0.000
H8 1.058 -1.274 0.000
H9 -0.999 1.075 0.000
H10 2.154 1.050 0.000
H11 0.050 3.352 0.000
H12 1.917 3.522 0.000

Atom - Atom Distances (Å)
  C1 O2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
C11.22991.48212.47573.85834.96121.12082.21242.67124.21705.08335.9541
O21.22992.39893.61134.89966.08852.04392.67563.89985.07426.28467.0356
C31.48212.39891.36612.50583.74512.20571.09842.12872.74144.10554.6495
C42.47573.61131.36611.46862.48672.66032.16001.10202.19902.74263.4868
C53.85834.89962.50581.46861.36174.12892.78932.20011.09942.14572.1473
C64.96126.08853.74512.48671.36175.00034.14382.71332.13541.09771.0957
H71.12082.04392.20572.66034.12895.00033.16602.37154.73254.87616.0667
H82.21242.67561.09842.16002.78934.14383.16603.12222.57004.73404.8722
H92.67123.89982.12871.10202.20012.71332.37153.12223.15312.50623.8062
H104.21705.07422.74142.19901.09942.13544.73252.57003.15313.11822.4829
H115.08336.28464.10552.74262.14571.09774.87614.73402.50623.11821.8747
H125.95417.03564.64953.48682.14731.09576.06674.87223.80622.48291.8747

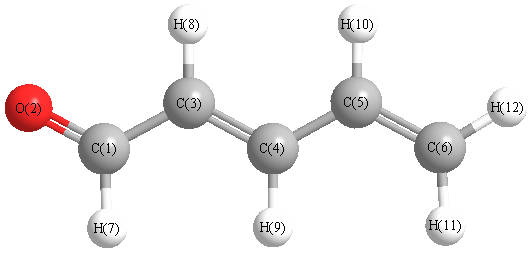
picture of Pentadienal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C4 120.676 C1 C3 H8 117.265
O2 C1 C3 124.121 O2 C1 H7 120.728
C3 C1 H7 115.152 C3 C4 C5 124.208
C3 C4 H9 118.802 C4 C3 H8 122.059
C4 C5 C6 122.909 C4 C5 H10 117.085
C5 C4 H9 116.990 C5 C6 H11 121.118
C5 C6 H12 121.432 C6 C5 H10 120.007
H11 C6 H12 117.450
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability