return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6O (Pentadienal)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-268.645775
Energy at 298.15K 
HF Energy-267.682339
Nuclear repulsion energy191.342390
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3256 3181        
2 A' 3184 3111        
3 A' 3172 3099        
4 A' 3163 3090        
5 A' 3137 3064        
6 A' 2900 2833        
7 A' 1729 1689        
8 A' 1707 1668        
9 A' 1645 1607        
10 A' 1476 1441        
11 A' 1423 1390        
12 A' 1340 1309        
13 A' 1331 1300        
14 A' 1283 1253        
15 A' 1212 1184        
16 A' 1140 1114        
17 A' 974 952        
18 A' 600 586        
19 A' 431 421        
20 A' 382 374        
21 A' 149 145        
22 A" 1029 1005        
23 A" 999 976        
24 A" 973 950        
25 A" 915 894        
26 A" 841 821        
27 A" 639 624        
28 A" 272 266        
29 A" 196 191        
30 A" 92 90        

Unscaled Zero Point Vibrational Energy (zpe) 20793.8 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 20313.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.93330 0.04374 0.04178

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.123 -1.595 0.000
O2 -1.117 -2.830 0.000
C3 0.096 -0.754 0.000
C4 0.000 0.610 0.000
C5 1.151 1.519 0.000
C6 1.022 2.875 0.000
H7 -2.090 -1.026 0.000
H8 1.066 -1.269 0.000
H9 -1.002 1.069 0.000
H10 2.151 1.062 0.000
H11 0.033 3.349 0.000
H12 1.899 3.531 0.000

Atom - Atom Distances (Å)
  C1 O2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
C11.23581.48032.47393.85534.95731.12202.21302.66654.21625.07675.9503
O21.23582.40443.61694.90496.09282.05022.68383.90125.08246.28537.0403
C31.48032.40441.36742.50593.74522.20261.09842.12842.74284.10364.6494
C42.47393.61691.36741.46662.48482.65382.16041.10222.19792.73933.4844
C53.85534.90492.50591.46661.36194.12042.78922.19931.09912.14472.1471
C64.95736.09283.74522.48481.36194.98984.14392.71242.13481.09751.0953
H71.12202.05022.20262.65384.12044.98983.16552.36064.72704.86246.0562
H82.21302.68381.09842.16042.78924.14393.16553.12162.57154.73224.8722
H92.66653.90122.12841.10222.19932.71242.36063.12163.15282.50353.8048
H104.21625.08242.74282.19791.09912.13484.72702.57153.15283.11682.4818
H115.07676.28534.10362.73932.14471.09754.86244.73222.50353.11681.8750
H125.95037.04034.64943.48442.14711.09536.05624.87223.80482.48181.8750

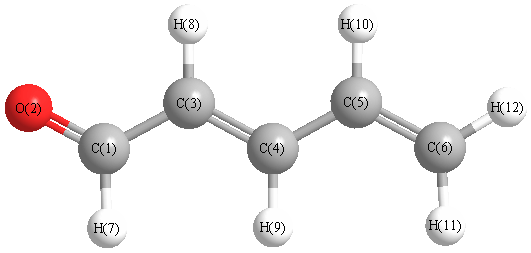
picture of Pentadienal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C4 120.569 C1 C3 H8 117.458
O2 C1 C3 124.314 O2 C1 H7 120.740
C3 C1 H7 114.946 C3 C4 C5 124.279
C3 C4 H9 118.648 C4 C3 H8 121.973
C4 C5 C6 122.879 C4 C5 H10 117.168
C5 C4 H9 117.073 C5 C6 H11 121.029
C5 C6 H12 121.430 C6 C5 H10 119.952
H11 C6 H12 117.541
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability