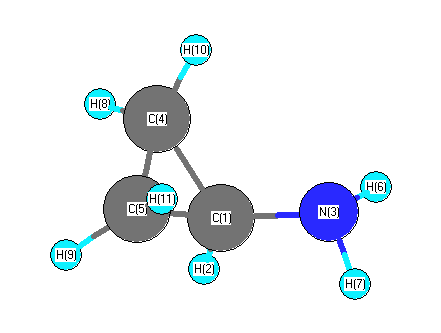
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3527 |
3380 |
0.90 |
|
|
|
| 2 |
A |
3268 |
3132 |
15.39 |
|
|
|
| 3 |
A |
3168 |
3036 |
6.13 |
|
|
|
| 4 |
A |
3105 |
2975 |
45.72 |
|
|
|
| 5 |
A |
1648 |
1579 |
16.56 |
|
|
|
| 6 |
A |
1507 |
1444 |
22.62 |
|
|
|
| 7 |
A |
1407 |
1349 |
19.70 |
|
|
|
| 8 |
A |
1254 |
1202 |
0.63 |
|
|
|
| 9 |
A |
1182 |
1133 |
4.50 |
|
|
|
| 10 |
A |
1049 |
1005 |
10.56 |
|
|
|
| 11 |
A |
1015 |
972 |
7.00 |
|
|
|
| 12 |
A |
844 |
809 |
33.17 |
|
|
|
| 13 |
A |
807 |
773 |
106.82 |
|
|
|
| 14 |
A |
771 |
738 |
4.39 |
|
|
|
| 15 |
A |
409 |
392 |
5.69 |
|
|
|
| 16 |
A |
3612 |
3462 |
2.31 |
|
|
|
| 17 |
A |
3255 |
3120 |
0.21 |
|
|
|
| 18 |
A |
3162 |
3030 |
20.95 |
|
|
|
| 19 |
A |
1443 |
1383 |
2.93 |
|
|
|
| 20 |
A |
1263 |
1210 |
1.10 |
|
|
|
| 21 |
A |
1187 |
1137 |
0.30 |
|
|
|
| 22 |
A |
1118 |
1071 |
1.24 |
|
|
|
| 23 |
A |
1053 |
1009 |
1.20 |
|
|
|
| 24 |
A |
914 |
876 |
10.57 |
|
|
|
| 25 |
A |
837 |
802 |
2.67 |
|
|
|
| 26 |
A |
397 |
381 |
5.34 |
|
|
|
| 27 |
A |
265 |
254 |
29.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21733.2 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 20826.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.895 |
|
|
|
| 2 |
H |
-0.405 |
|
|
|
| 3 |
N |
-0.298 |
|
|
|
| 4 |
C |
0.788 |
|
|
|
| 5 |
C |
0.788 |
|
|
|
| 6 |
H |
-0.020 |
|
|
|
| 7 |
H |
-0.020 |
|
|
|
| 8 |
H |
-0.447 |
|
|
|
| 9 |
H |
-0.447 |
|
|
|
| 10 |
H |
-0.416 |
|
|
|
| 11 |
H |
-0.416 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.794 |
0.960 |
0.000 |
1.246 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.990 |
0.645 |
0.000 |
| y |
0.645 |
-23.817 |
0.000 |
| z |
0.000 |
0.000 |
-24.927 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.618 |
0.645 |
0.000 |
| y |
0.645 |
2.141 |
0.000 |
| z |
0.000 |
0.000 |
0.476 |
|
| Polar |
|---|
| 3z2-r2 | 0.953 |
|---|
| x2-y2 | -3.173 |
|---|
| xy | 0.645 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.521 |
0.461 |
0.000 |
| y |
0.461 |
7.240 |
0.000 |
| z |
0.000 |
0.000 |
6.702 |
<r2> (average value of r
2) Å
2
| <r2> |
77.687 |
| (<r2>)1/2 |
8.814 |