return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (Methyl cyclopropyl ketone)

using model chemistry: QCISD(T)/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/aug-cc-pVDZ
 hartrees
Energy at 0K-269.860857
Energy at 298.15K-269.869398
HF Energy-268.863279
Nuclear repulsion energy228.303125
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3242 3170        
2 A' 3169 3098        
3 A' 3146 3076        
4 A' 3136 3066        
5 A' 3017 2950        
6 A' 1721 1683        
7 A' 1478 1446        
8 A' 1447 1415        
9 A' 1410 1379        
10 A' 1359 1329        
11 A' 1206 1180        
12 A' 1186 1160        
13 A' 1092 1067        
14 A' 1037 1014        
15 A' 962 941        
16 A' 910 890        
17 A' 790 773        
18 A' 741 725        
19 A' 580 567        
20 A' 366 357        
21 A' 232 227        
22 A" 3229 3157        
23 A" 3131 3062        
24 A" 3090 3021        
25 A" 1458 1426        
26 A" 1436 1404        
27 A" 1190 1164        
28 A" 1103 1079        
29 A" 1062 1038        
30 A" 1027 1004        
31 A" 860 841        
32 A" 818 800        
33 A" 585 572        
34 A" 253 247        
35 A" 140 137        
36 A" 66 64        

Unscaled Zero Point Vibrational Energy (zpe) 25837.6 cm-1
Scaled (by 0.9778) Zero Point Vibrational Energy (zpe) 25264.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/aug-cc-pVDZ
ABC
0.23544 0.08637 0.07549

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -1.494 -0.555 0.000
C2 -0.279 -0.763 0.000
C3 0.299 -2.173 0.000
C4 0.710 0.369 0.000
C5 0.299 1.643 0.754
C6 0.299 1.643 -0.754
H7 -0.515 -2.915 0.000
H8 0.940 -2.310 -0.892
H9 0.940 -2.310 0.892
H10 1.772 0.098 0.000
H11 -0.673 1.595 1.255
H12 1.094 2.185 1.277
H13 -0.673 1.595 -1.255
H14 1.094 2.185 -1.277

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
O11.23302.41542.39022.93512.93512.55523.13113.13113.33092.62093.97902.62093.9790
C21.23301.52361.50342.58692.58692.16442.16232.16232.22462.69993.49352.69993.4935
C32.41541.52362.57463.88953.88951.10121.10681.10682.70694.08824.60964.08824.6096
C42.39021.50342.57461.53671.53673.50452.83292.83291.09562.23432.25282.23432.2528
C52.93512.58693.88951.53671.50884.69064.32994.00722.26361.09481.09452.23312.2476
C62.93512.58693.88951.53671.50884.69064.00724.32992.26362.23312.24761.09481.0945
H72.55522.16441.10123.50454.69064.69061.81021.81023.78254.68335.49734.68335.4973
H83.13112.16231.10682.83294.32994.00721.81021.78322.69984.73924.99314.24064.5140
H93.13112.16231.10682.83294.00724.32991.81021.78322.69984.24064.51404.73924.9931
H103.33092.22462.70691.09562.26362.26363.78252.69982.69983.12942.53843.12942.5384
H112.62092.69994.08822.23431.09482.23314.68334.73924.24063.12941.86292.51073.1437
H123.97903.49354.60962.25281.09452.24765.49734.99314.51402.53841.86293.14372.5542
H132.62092.69994.08822.23432.23311.09484.68334.24064.73923.12942.51073.14371.8629
H143.97903.49354.60962.25282.24761.09455.49734.51404.99312.53843.14372.55421.8629

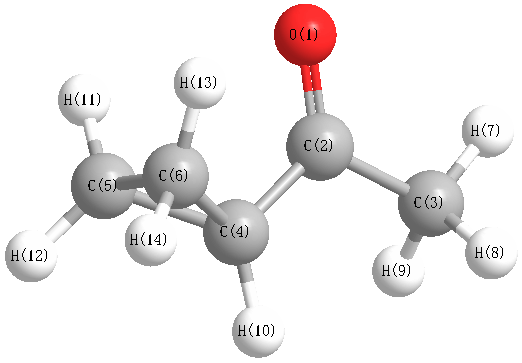
picture of Methyl cyclopropyl ketone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 122.033 O1 C2 C4 121.424
C2 C3 H7 110.057 C2 C3 H8 109.561
C2 C3 H9 109.561 C2 C4 C5 116.623
C2 C4 C6 116.623 C2 C4 H10 116.867
C3 C2 C4 116.543 C4 C5 C6 60.598
C4 C5 H11 115.190 C4 C5 H12 116.779
C4 C6 C5 60.598 C4 C6 H13 115.190
C4 C6 H14 116.779 C5 C4 C6 58.803
C5 C4 H10 117.643 C5 C6 H13 117.230
C5 C6 H14 118.525 C6 C4 H10 117.643
C6 C5 H11 117.230 C6 C5 H12 118.525
H7 C3 H8 110.140 H7 C3 H9 110.140
H8 C3 H9 107.337 H11 C5 H12 116.618
H13 C6 H14 116.618
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability