Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.816903 |
| Energy at 298.15K | -235.829117 |
| HF Energy | -235.816903 |
| Nuclear repulsion energy | 230.081643 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3177 |
3045 |
48.09 |
|
|
|
| 2 |
A |
3157 |
3025 |
12.04 |
|
|
|
| 3 |
A |
3147 |
3016 |
9.06 |
|
|
|
| 4 |
A |
3136 |
3005 |
36.11 |
|
|
|
| 5 |
A |
3129 |
2998 |
57.39 |
|
|
|
| 6 |
A |
3107 |
2977 |
13.40 |
|
|
|
| 7 |
A |
3103 |
2974 |
16.83 |
|
|
|
| 8 |
A |
3086 |
2957 |
5.12 |
|
|
|
| 9 |
A |
3052 |
2925 |
34.23 |
|
|
|
| 10 |
A |
3049 |
2922 |
33.80 |
|
|
|
| 11 |
A |
3045 |
2918 |
30.27 |
|
|
|
| 12 |
A |
3031 |
2905 |
19.03 |
|
|
|
| 13 |
A |
1753 |
1680 |
4.50 |
|
|
|
| 14 |
A |
1487 |
1425 |
7.63 |
|
|
|
| 15 |
A |
1477 |
1416 |
7.81 |
|
|
|
| 16 |
A |
1472 |
1411 |
3.45 |
|
|
|
| 17 |
A |
1470 |
1409 |
2.45 |
|
|
|
| 18 |
A |
1462 |
1401 |
0.58 |
|
|
|
| 19 |
A |
1455 |
1394 |
5.63 |
|
|
|
| 20 |
A |
1431 |
1372 |
11.65 |
|
|
|
| 21 |
A |
1397 |
1339 |
3.84 |
|
|
|
| 22 |
A |
1385 |
1327 |
1.95 |
|
|
|
| 23 |
A |
1366 |
1309 |
0.20 |
|
|
|
| 24 |
A |
1314 |
1259 |
0.01 |
|
|
|
| 25 |
A |
1293 |
1239 |
0.47 |
|
|
|
| 26 |
A |
1277 |
1224 |
2.58 |
|
|
|
| 27 |
A |
1244 |
1192 |
0.15 |
|
|
|
| 28 |
A |
1168 |
1119 |
0.28 |
|
|
|
| 29 |
A |
1116 |
1069 |
1.02 |
|
|
|
| 30 |
A |
1078 |
1033 |
1.93 |
|
|
|
| 31 |
A |
1059 |
1015 |
2.55 |
|
|
|
| 32 |
A |
1048 |
1004 |
2.23 |
|
|
|
| 33 |
A |
1014 |
972 |
6.45 |
|
|
|
| 34 |
A |
1001 |
959 |
1.09 |
|
|
|
| 35 |
A |
929 |
891 |
4.93 |
|
|
|
| 36 |
A |
904 |
866 |
6.14 |
|
|
|
| 37 |
A |
869 |
832 |
3.73 |
|
|
|
| 38 |
A |
741 |
711 |
2.36 |
|
|
|
| 39 |
A |
712 |
682 |
34.54 |
|
|
|
| 40 |
A |
571 |
547 |
7.51 |
|
|
|
| 41 |
A |
472 |
452 |
1.10 |
|
|
|
| 42 |
A |
365 |
350 |
0.00 |
|
|
|
| 43 |
A |
283 |
271 |
0.04 |
|
|
|
| 44 |
A |
242 |
232 |
0.00 |
|
|
|
| 45 |
A |
198 |
190 |
0.02 |
|
|
|
| 46 |
A |
120 |
115 |
0.36 |
|
|
|
| 47 |
A |
90 |
86 |
0.02 |
|
|
|
| 48 |
A |
49 |
47 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36264.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 34752.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.482 |
0.941 |
-0.078 |
| C2 |
1.995 |
-0.473 |
-0.138 |
| C3 |
0.767 |
-0.928 |
0.137 |
| C4 |
-0.436 |
-0.138 |
0.552 |
| C5 |
-1.589 |
-0.250 |
-0.450 |
| C6 |
-2.835 |
0.506 |
-0.010 |
| H7 |
1.698 |
1.649 |
0.208 |
| H8 |
3.305 |
1.037 |
0.644 |
| H9 |
2.882 |
1.254 |
-1.053 |
| H10 |
2.745 |
-1.210 |
-0.438 |
| H11 |
0.600 |
-2.005 |
0.035 |
| H12 |
-0.182 |
0.920 |
0.698 |
| H13 |
-0.787 |
-0.508 |
1.530 |
| H14 |
-1.248 |
0.125 |
-1.426 |
| H15 |
-1.833 |
-1.312 |
-0.600 |
| H16 |
-3.643 |
0.407 |
-0.745 |
| H17 |
-3.210 |
0.127 |
0.952 |
| H18 |
-2.627 |
1.577 |
0.115 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4964 | 2.5461 | 3.1746 | 4.2575 | 5.3351 | 1.0949 | 1.0997 | 1.0996 | 2.1965 | 3.4975 | 2.7749 | 3.9209 | 4.0494 | 4.8958 | 6.1844 | 5.8408 | 5.1515 |
C2 | 1.4964 | | 1.3388 | 2.5495 | 3.6042 | 4.9299 | 2.1708 | 2.1470 | 2.1465 | 1.0934 | 2.0794 | 2.7164 | 3.2441 | 3.5409 | 3.9464 | 5.7387 | 5.3511 | 5.0622 | C3 | 2.5461 | 1.3388 | | 1.4979 | 2.5202 | 3.8793 | 2.7419 | 3.2508 | 3.2644 | 2.0793 | 1.0941 | 2.1519 | 2.1288 | 2.7592 | 2.7294 | 4.6911 | 4.1937 | 4.2179 | C4 | 3.1746 | 2.5495 | 1.4979 | | 1.5317 | 2.5465 | 2.8053 | 3.9231 | 3.9408 | 3.4999 | 2.1967 | 1.0979 | 1.1026 | 2.1550 | 2.1584 | 3.5021 | 2.8144 | 2.8162 | C5 | 4.2575 | 3.6042 | 2.5202 | 1.5317 | | 1.5229 | 3.8528 | 5.1776 | 4.7555 | 4.4384 | 2.8469 | 2.1600 | 2.1520 | 1.0995 | 1.1002 | 2.1771 | 2.1761 | 2.1762 | C6 | 5.3351 | 4.9299 | 3.8793 | 2.5465 | 1.5229 | | 4.6802 | 6.1980 | 5.8598 | 5.8533 | 4.2549 | 2.7766 | 2.7553 | 2.1609 | 2.1583 | 1.0972 | 1.0987 | 1.0986 | H7 | 1.0949 | 2.1708 | 2.7419 | 2.8053 | 3.8528 | 4.6802 | | 1.7744 | 1.7750 | 3.1129 | 3.8198 | 2.0754 | 3.5466 | 3.6982 | 4.6792 | 5.5661 | 5.1916 | 4.3261 | H8 | 1.0997 | 2.1470 | 3.2508 | 3.9231 | 5.1776 | 6.1980 | 1.7744 | | 1.7628 | 2.5567 | 4.1166 | 3.4900 | 4.4636 | 5.0849 | 5.7858 | 7.1141 | 6.5854 | 5.9799 | H9 | 1.0996 | 2.1465 | 3.2644 | 3.9408 | 4.7555 | 5.8598 | 1.7750 | 1.7628 | | 2.5432 | 4.1250 | 3.5454 | 4.8213 | 4.2984 | 5.3877 | 6.5875 | 6.5115 | 5.6405 | H10 | 2.1965 | 1.0934 | 2.0793 | 3.4999 | 4.4384 | 5.8533 | 3.1129 | 2.5567 | 2.5432 | | 2.3360 | 3.7940 | 4.1040 | 4.3247 | 4.5819 | 6.5965 | 6.2589 | 6.0765 | H11 | 3.4975 | 2.0794 | 1.0941 | 2.1967 | 2.8469 | 4.2549 | 3.8198 | 4.1166 | 4.1250 | 2.3360 | | 3.0992 | 2.5296 | 3.1764 | 2.6084 | 4.9426 | 4.4606 | 4.8215 | H12 | 2.7749 | 2.7164 | 2.1519 | 1.0979 | 2.1600 | 2.7766 | 2.0754 | 3.4900 | 3.5454 | 3.7940 | 3.0992 | | 1.7600 | 2.5061 | 3.0649 | 3.7847 | 3.1396 | 2.5974 | H13 | 3.9209 | 3.2441 | 2.1288 | 1.1026 | 2.1520 | 2.7553 | 3.5466 | 4.4636 | 4.8213 | 4.1040 | 2.5296 | 1.7600 | | 3.0584 | 2.5060 | 3.7643 | 2.5702 | 3.1199 | H14 | 4.0494 | 3.5409 | 2.7592 | 2.1550 | 1.0995 | 2.1609 | 3.6982 | 5.0849 | 4.2984 | 4.3247 | 3.1764 | 2.5061 | 3.0584 | | 1.7579 | 2.5057 | 3.0822 | 2.5263 | H15 | 4.8958 | 3.9464 | 2.7294 | 2.1584 | 1.1002 | 2.1583 | 4.6792 | 5.7858 | 5.3877 | 4.5819 | 2.6084 | 3.0649 | 2.5060 | 1.7579 | | 2.5005 | 2.5250 | 3.0805 | H16 | 6.1844 | 5.7387 | 4.6911 | 3.5021 | 2.1771 | 1.0972 | 5.5661 | 7.1141 | 6.5875 | 6.5965 | 4.9426 | 3.7847 | 3.7643 | 2.5057 | 2.5005 | | 1.7733 | 1.7727 | H17 | 5.8408 | 5.3511 | 4.1937 | 2.8144 | 2.1761 | 1.0987 | 5.1916 | 6.5854 | 6.5115 | 6.2589 | 4.4606 | 3.1396 | 2.5702 | 3.0822 | 2.5250 | 1.7733 | | 1.7726 | H18 | 5.1515 | 5.0622 | 4.2179 | 2.8162 | 2.1762 | 1.0986 | 4.3261 | 5.9799 | 5.6405 | 6.0765 | 4.8215 | 2.5974 | 3.1199 | 2.5263 | 3.0805 | 1.7727 | 1.7726 | |
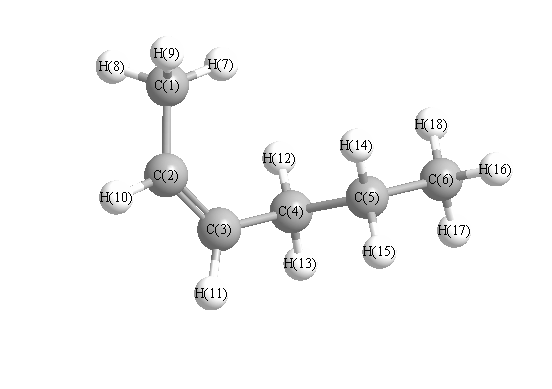 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.721 |
|
C1 |
C2 |
H10 |
115.132 |
| C2 |
C1 |
H7 |
112.891 |
|
C2 |
C1 |
H8 |
110.656 |
| C2 |
C1 |
H9 |
110.627 |
|
C2 |
C3 |
C4 |
127.906 |
| C2 |
C3 |
H11 |
117.097 |
|
C3 |
C2 |
H10 |
117.146 |
| C3 |
C4 |
C5 |
112.579 |
|
C3 |
C4 |
H12 |
111.057 |
| C3 |
C4 |
H13 |
108.946 |
|
C4 |
C3 |
H11 |
114.990 |
| C4 |
C5 |
C6 |
112.948 |
|
C4 |
C5 |
H14 |
108.869 |
| C4 |
C5 |
H15 |
109.087 |
|
C5 |
C4 |
H12 |
109.343 |
| C5 |
C4 |
H13 |
108.457 |
|
C5 |
C6 |
H16 |
111.346 |
| C5 |
C6 |
H17 |
111.179 |
|
C5 |
C6 |
H18 |
111.190 |
| C6 |
C5 |
H14 |
109.926 |
|
C6 |
C5 |
H15 |
109.677 |
| H7 |
C1 |
H8 |
107.904 |
|
H7 |
C1 |
H9 |
107.959 |
| H8 |
C1 |
H9 |
106.547 |
|
H12 |
C4 |
H13 |
106.225 |
| H14 |
C5 |
H16 |
93.951 |
|
H16 |
C6 |
H17 |
107.717 |
| H16 |
C6 |
H18 |
107.670 |
|
H17 |
C6 |
H18 |
107.556 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.122 |
|
|
|
| 2 |
C |
0.374 |
|
|
|
| 3 |
C |
1.186 |
|
|
|
| 4 |
C |
0.507 |
|
|
|
| 5 |
C |
0.366 |
|
|
|
| 6 |
C |
0.397 |
|
|
|
| 7 |
H |
-0.302 |
|
|
|
| 8 |
H |
0.003 |
|
|
|
| 9 |
H |
0.027 |
|
|
|
| 10 |
H |
-0.577 |
|
|
|
| 11 |
H |
-0.649 |
|
|
|
| 12 |
H |
-0.366 |
|
|
|
| 13 |
H |
-0.189 |
|
|
|
| 14 |
H |
-0.244 |
|
|
|
| 15 |
H |
-0.281 |
|
|
|
| 16 |
H |
-0.137 |
|
|
|
| 17 |
H |
-0.103 |
|
|
|
| 18 |
H |
-0.134 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.078 |
0.251 |
-0.020 |
0.263 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.140 |
0.963 |
-0.572 |
| y |
0.963 |
-39.433 |
-0.028 |
| z |
-0.572 |
-0.028 |
-41.329 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.242 |
0.963 |
-0.572 |
| y |
0.963 |
0.801 |
-0.028 |
| z |
-0.572 |
-0.028 |
-2.043 |
|
| Polar |
|---|
| 3z2-r2 | -4.086 |
|---|
| x2-y2 | 0.293 |
|---|
| xy | 0.963 |
|---|
| xz | -0.572 |
|---|
| yz | -0.028 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.736 |
0.560 |
-0.470 |
| y |
0.560 |
10.683 |
0.049 |
| z |
-0.470 |
0.049 |
9.562 |
<r2> (average value of r
2) Å
2
| <r2> |
261.876 |
| (<r2>)1/2 |
16.183 |
