Jump to
S1C2
Vibrational Frequencies calculated at M06-2X/aug-cc-pVDZ
Geometric Data calculated at M06-2X/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at M06-2X/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -260.985031 |
| Energy at 298.15K | -260.990119 |
| HF Energy | -260.985031 |
| Nuclear repulsion energy | 127.995648 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3545 |
3545 |
43.37 |
|
|
|
| 2 |
A' |
1602 |
1602 |
88.24 |
|
|
|
| 3 |
A' |
1474 |
1474 |
216.74 |
|
|
|
| 4 |
A' |
1071 |
1071 |
19.56 |
|
|
|
| 5 |
A' |
833 |
833 |
86.27 |
|
|
|
| 6 |
A' |
744 |
744 |
94.28 |
|
|
|
| 7 |
A' |
650 |
650 |
130.00 |
|
|
|
| 8 |
A" |
3685 |
3685 |
74.70 |
|
|
|
| 9 |
A" |
1763 |
1763 |
434.93 |
|
|
|
| 10 |
A" |
1269 |
1269 |
56.32 |
|
|
|
| 11 |
A" |
590 |
590 |
0.53 |
|
|
|
| 12 |
A" |
435 |
435 |
26.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8830.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8830.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.070 |
-1.240 |
0.000 |
| N2 |
0.008 |
0.140 |
0.000 |
| O3 |
0.008 |
0.681 |
1.085 |
| O4 |
0.008 |
0.681 |
-1.085 |
| H5 |
-0.330 |
-1.595 |
-0.862 |
| H6 |
-0.330 |
-1.595 |
0.862 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
O3 |
O4 |
H5 |
H6 |
| N1 | | 1.3810 | 2.2065 | 2.2065 | 1.0148 | 1.0148 |
N2 | 1.3810 | | 1.2120 | 1.2120 | 1.9667 | 1.9667 | O3 | 2.2065 | 1.2120 | | 2.1692 | 3.0141 | 2.3115 | O4 | 2.2065 | 1.2120 | 2.1692 | | 2.3115 | 3.0141 | H5 | 1.0148 | 1.9667 | 3.0141 | 2.3115 | | 1.7248 | H6 | 1.0148 | 1.9667 | 2.3115 | 3.0141 | 1.7248 | |
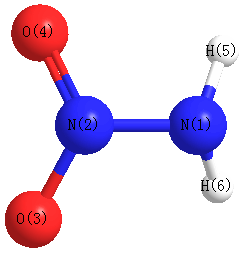 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
N2 |
O3 |
116.478 |
|
N1 |
N2 |
O4 |
116.478 |
| N2 |
N1 |
H5 |
109.398 |
|
N2 |
N1 |
H6 |
109.398 |
| O3 |
N2 |
O4 |
126.985 |
|
H5 |
N1 |
H6 |
116.377 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
0.168 |
|
|
|
| 2 |
N |
0.913 |
|
|
|
| 3 |
O |
-0.595 |
|
|
|
| 4 |
O |
-0.595 |
|
|
|
| 5 |
H |
0.054 |
|
|
|
| 6 |
H |
0.054 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.208 |
-3.669 |
0.000 |
3.863 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.603 |
2.258 |
0.000 |
| y |
2.258 |
-20.616 |
0.000 |
| z |
0.000 |
0.000 |
-24.470 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.061 |
2.258 |
0.000 |
| y |
2.258 |
2.921 |
0.000 |
| z |
0.000 |
0.000 |
-2.860 |
|
| Polar |
|---|
| 3z2-r2 | -5.720 |
|---|
| x2-y2 | -1.988 |
|---|
| xy | 2.258 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.833 |
0.022 |
0.000 |
| y |
0.022 |
4.828 |
0.000 |
| z |
0.000 |
0.000 |
5.156 |
<r2> (average value of r
2) Å
2
| <r2> |
58.055 |
| (<r2>)1/2 |
7.619 |
