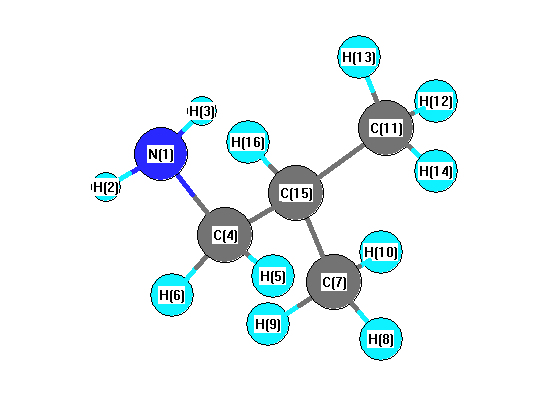
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3621 |
3470 |
1.58 |
|
|
|
| 2 |
A |
3538 |
3391 |
0.59 |
|
|
|
| 3 |
A |
3136 |
3005 |
42.36 |
|
|
|
| 4 |
A |
3134 |
3003 |
24.52 |
|
|
|
| 5 |
A |
3124 |
2994 |
65.76 |
|
|
|
| 6 |
A |
3119 |
2989 |
10.75 |
|
|
|
| 7 |
A |
3088 |
2959 |
32.37 |
|
|
|
| 8 |
A |
3058 |
2931 |
19.55 |
|
|
|
| 9 |
A |
3046 |
2919 |
22.28 |
|
|
|
| 10 |
A |
3041 |
2914 |
30.68 |
|
|
|
| 11 |
A |
2963 |
2840 |
69.23 |
|
|
|
| 12 |
A |
1647 |
1578 |
32.72 |
|
|
|
| 13 |
A |
1497 |
1435 |
16.28 |
|
|
|
| 14 |
A |
1487 |
1425 |
0.82 |
|
|
|
| 15 |
A |
1484 |
1423 |
5.97 |
|
|
|
| 16 |
A |
1473 |
1412 |
1.65 |
|
|
|
| 17 |
A |
1465 |
1404 |
1.05 |
|
|
|
| 18 |
A |
1422 |
1363 |
8.18 |
|
|
|
| 19 |
A |
1399 |
1341 |
3.77 |
|
|
|
| 20 |
A |
1390 |
1332 |
10.25 |
|
|
|
| 21 |
A |
1360 |
1304 |
0.39 |
|
|
|
| 22 |
A |
1336 |
1281 |
0.47 |
|
|
|
| 23 |
A |
1298 |
1244 |
5.84 |
|
|
|
| 24 |
A |
1226 |
1175 |
0.30 |
|
|
|
| 25 |
A |
1197 |
1147 |
0.77 |
|
|
|
| 26 |
A |
1168 |
1119 |
3.90 |
|
|
|
| 27 |
A |
1105 |
1059 |
20.49 |
|
|
|
| 28 |
A |
1096 |
1050 |
1.57 |
|
|
|
| 29 |
A |
968 |
928 |
0.71 |
|
|
|
| 30 |
A |
950 |
910 |
0.72 |
|
|
|
| 31 |
A |
921 |
882 |
9.02 |
|
|
|
| 32 |
A |
915 |
876 |
19.34 |
|
|
|
| 33 |
A |
831 |
796 |
81.44 |
|
|
|
| 34 |
A |
809 |
776 |
11.43 |
|
|
|
| 35 |
A |
487 |
466 |
5.93 |
|
|
|
| 36 |
A |
415 |
397 |
0.49 |
|
|
|
| 37 |
A |
363 |
348 |
0.17 |
|
|
|
| 38 |
A |
274 |
262 |
16.89 |
|
|
|
| 39 |
A |
261 |
250 |
1.70 |
|
|
|
| 40 |
A |
231 |
222 |
16.69 |
|
|
|
| 41 |
A |
213 |
204 |
11.84 |
|
|
|
| 42 |
A |
117 |
112 |
3.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32834.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 31465.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.167 |
|
|
|
| 2 |
H |
-0.043 |
|
|
|
| 3 |
H |
0.008 |
|
|
|
| 4 |
C |
0.983 |
|
|
|
| 5 |
H |
-0.243 |
|
|
|
| 6 |
H |
-0.337 |
|
|
|
| 7 |
C |
0.747 |
|
|
|
| 8 |
H |
-0.061 |
|
|
|
| 9 |
H |
-0.141 |
|
|
|
| 10 |
H |
-0.144 |
|
|
|
| 11 |
C |
0.835 |
|
|
|
| 12 |
H |
-0.154 |
|
|
|
| 13 |
H |
-0.175 |
|
|
|
| 14 |
H |
-0.062 |
|
|
|
| 15 |
C |
-0.694 |
|
|
|
| 16 |
H |
-0.351 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.110 |
-0.163 |
1.067 |
1.085 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.035 |
0.664 |
-3.155 |
| y |
0.664 |
-33.378 |
0.507 |
| z |
-3.155 |
0.507 |
-35.834 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.571 |
0.664 |
-3.155 |
| y |
0.664 |
1.057 |
0.507 |
| z |
-3.155 |
0.507 |
-2.628 |
|
| Polar |
|---|
| 3z2-r2 | -5.255 |
|---|
| x2-y2 | 0.343 |
|---|
| xy | 0.664 |
|---|
| xz | -3.155 |
|---|
| yz | 0.507 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.085 |
-0.107 |
0.059 |
| y |
-0.107 |
9.256 |
-0.084 |
| z |
0.059 |
-0.084 |
8.346 |
<r2> (average value of r
2) Å
2
| <r2> |
150.143 |
| (<r2>)1/2 |
12.253 |