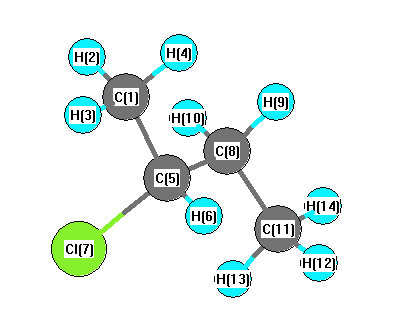
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3166 |
3034 |
16.62 |
|
|
|
| 2 |
A |
3156 |
3024 |
20.85 |
|
|
|
| 3 |
A |
3143 |
3012 |
28.62 |
|
|
|
| 4 |
A |
3139 |
3008 |
25.58 |
|
|
|
| 5 |
A |
3113 |
2983 |
10.40 |
|
|
|
| 6 |
A |
3094 |
2965 |
1.49 |
|
|
|
| 7 |
A |
3060 |
2933 |
11.17 |
|
|
|
| 8 |
A |
3059 |
2932 |
41.55 |
|
|
|
| 9 |
A |
3048 |
2921 |
15.75 |
|
|
|
| 10 |
A |
1484 |
1422 |
5.96 |
|
|
|
| 11 |
A |
1477 |
1415 |
7.81 |
|
|
|
| 12 |
A |
1474 |
1413 |
3.55 |
|
|
|
| 13 |
A |
1464 |
1403 |
10.17 |
|
|
|
| 14 |
A |
1453 |
1393 |
0.73 |
|
|
|
| 15 |
A |
1403 |
1344 |
1.11 |
|
|
|
| 16 |
A |
1397 |
1338 |
13.25 |
|
|
|
| 17 |
A |
1384 |
1326 |
0.49 |
|
|
|
| 18 |
A |
1312 |
1257 |
11.17 |
|
|
|
| 19 |
A |
1305 |
1250 |
5.90 |
|
|
|
| 20 |
A |
1251 |
1198 |
14.39 |
|
|
|
| 21 |
A |
1179 |
1130 |
8.83 |
|
|
|
| 22 |
A |
1141 |
1093 |
2.25 |
|
|
|
| 23 |
A |
1088 |
1042 |
2.66 |
|
|
|
| 24 |
A |
1057 |
1013 |
3.70 |
|
|
|
| 25 |
A |
1000 |
958 |
9.68 |
|
|
|
| 26 |
A |
967 |
926 |
5.52 |
|
|
|
| 27 |
A |
862 |
826 |
8.97 |
|
|
|
| 28 |
A |
790 |
757 |
11.24 |
|
|
|
| 29 |
A |
621 |
595 |
23.23 |
|
|
|
| 30 |
A |
462 |
443 |
1.20 |
|
|
|
| 31 |
A |
388 |
372 |
2.29 |
|
|
|
| 32 |
A |
327 |
313 |
1.29 |
|
|
|
| 33 |
A |
248 |
238 |
0.16 |
|
|
|
| 34 |
A |
230 |
221 |
0.07 |
|
|
|
| 35 |
A |
211 |
202 |
1.08 |
|
|
|
| 36 |
A |
112 |
107 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27030.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 25903.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.695 |
|
|
|
| 2 |
H |
-0.048 |
|
|
|
| 3 |
H |
-0.121 |
|
|
|
| 4 |
H |
-0.095 |
|
|
|
| 5 |
C |
-0.361 |
|
|
|
| 6 |
H |
-0.242 |
|
|
|
| 7 |
Cl |
-0.166 |
|
|
|
| 8 |
C |
0.742 |
|
|
|
| 9 |
H |
-0.235 |
|
|
|
| 10 |
H |
-0.261 |
|
|
|
| 11 |
C |
0.434 |
|
|
|
| 12 |
H |
-0.116 |
|
|
|
| 13 |
H |
-0.108 |
|
|
|
| 14 |
H |
-0.118 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.851 |
2.092 |
0.369 |
2.288 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.039 |
-1.283 |
-0.338 |
| y |
-1.283 |
-40.504 |
0.409 |
| z |
-0.338 |
0.409 |
-39.312 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.131 |
-1.283 |
-0.338 |
| y |
-1.283 |
-0.828 |
0.409 |
| z |
-0.338 |
0.409 |
0.959 |
|
| Polar |
|---|
| 3z2-r2 | 1.919 |
|---|
| x2-y2 | 0.464 |
|---|
| xy | -1.283 |
|---|
| xz | -0.338 |
|---|
| yz | 0.409 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.475 |
0.090 |
0.057 |
| y |
0.090 |
10.493 |
0.044 |
| z |
0.057 |
0.044 |
8.419 |
<r2> (average value of r
2) Å
2
| <r2> |
181.584 |
| (<r2>)1/2 |
13.475 |