Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -232.440651 |
| Energy at 298.15K | -232.448906 |
| Nuclear repulsion energy | 176.209245 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3189 |
3056 |
8.87 |
|
|
|
| 2 |
A |
3163 |
3031 |
19.62 |
|
|
|
| 3 |
A |
3155 |
3024 |
20.71 |
|
|
|
| 4 |
A |
3134 |
3004 |
8.75 |
|
|
|
| 5 |
A |
3081 |
2953 |
8.23 |
|
|
|
| 6 |
A |
3073 |
2945 |
22.49 |
|
|
|
| 7 |
A |
3059 |
2932 |
8.28 |
|
|
|
| 8 |
A |
3045 |
2918 |
17.25 |
|
|
|
| 9 |
A |
1811 |
1736 |
166.77 |
|
|
|
| 10 |
A |
1477 |
1416 |
10.47 |
|
|
|
| 11 |
A |
1469 |
1408 |
8.58 |
|
|
|
| 12 |
A |
1456 |
1395 |
9.27 |
|
|
|
| 13 |
A |
1444 |
1383 |
22.98 |
|
|
|
| 14 |
A |
1432 |
1372 |
1.49 |
|
|
|
| 15 |
A |
1400 |
1342 |
13.03 |
|
|
|
| 16 |
A |
1386 |
1328 |
58.20 |
|
|
|
| 17 |
A |
1357 |
1300 |
9.85 |
|
|
|
| 18 |
A |
1273 |
1220 |
0.01 |
|
|
|
| 19 |
A |
1194 |
1144 |
56.40 |
|
|
|
| 20 |
A |
1119 |
1072 |
0.44 |
|
|
|
| 21 |
A |
1112 |
1066 |
0.94 |
|
|
|
| 22 |
A |
1014 |
972 |
4.58 |
|
|
|
| 23 |
A |
949 |
909 |
4.86 |
|
|
|
| 24 |
A |
935 |
896 |
5.97 |
|
|
|
| 25 |
A |
780 |
747 |
3.33 |
|
|
|
| 26 |
A |
743 |
712 |
2.58 |
|
|
|
| 27 |
A |
593 |
568 |
8.89 |
|
|
|
| 28 |
A |
476 |
456 |
0.26 |
|
|
|
| 29 |
A |
402 |
385 |
4.00 |
|
|
|
| 30 |
A |
245 |
235 |
5.69 |
|
|
|
| 31 |
A |
196 |
188 |
0.17 |
|
|
|
| 32 |
A |
109 |
105 |
0.19 |
|
|
|
| 33 |
A |
57 |
55 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24662.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 23634.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.878 |
-0.504 |
0.030 |
| C2 |
0.526 |
0.166 |
-0.018 |
| C3 |
-0.681 |
-0.747 |
-0.052 |
| C4 |
-2.011 |
-0.022 |
0.043 |
| O5 |
0.417 |
1.375 |
-0.020 |
| H6 |
2.668 |
0.232 |
-0.138 |
| H7 |
2.018 |
-0.964 |
1.019 |
| H8 |
1.945 |
-1.312 |
-0.710 |
| H9 |
-2.844 |
-0.734 |
-0.007 |
| H10 |
-2.088 |
0.535 |
0.984 |
| H11 |
-2.122 |
0.701 |
-0.773 |
| H12 |
-0.614 |
-1.331 |
-0.985 |
| H13 |
-0.566 |
-1.488 |
0.754 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5099 | 2.5722 | 3.9193 | 2.3811 | 1.0931 | 1.0994 | 1.0978 | 4.7273 | 4.2095 | 4.2538 | 2.8145 | 2.7325 |
C2 | 1.5099 | | 1.5143 | 2.5453 | 1.2134 | 2.1461 | 2.1396 | 2.1634 | 3.4879 | 2.8237 | 2.8048 | 2.1155 | 2.1276 | C3 | 2.5722 | 1.5143 | | 1.5178 | 2.3891 | 3.4904 | 2.9123 | 2.7664 | 2.1627 | 2.1668 | 2.1656 | 1.1020 | 1.1012 | C4 | 3.9193 | 2.5453 | 1.5178 | | 2.8017 | 4.6896 | 4.2517 | 4.2296 | 1.0965 | 1.0960 | 1.0955 | 2.1731 | 2.1782 | O5 | 2.3811 | 1.2134 | 2.3891 | 2.8017 | | 2.5273 | 3.0186 | 3.1678 | 3.8827 | 2.8259 | 2.7320 | 3.0517 | 3.1244 | H6 | 1.0931 | 2.1461 | 3.4904 | 4.6896 | 2.5273 | | 1.7865 | 1.7987 | 5.5970 | 4.8958 | 4.8540 | 3.7323 | 3.7704 | H7 | 1.0994 | 2.1396 | 2.9123 | 4.2517 | 3.0186 | 1.7865 | | 1.7655 | 4.9740 | 4.3711 | 4.8082 | 3.3280 | 2.6504 | H8 | 1.0978 | 2.1634 | 2.7664 | 4.2296 | 3.1678 | 1.7987 | 1.7655 | | 4.8748 | 4.7488 | 4.5386 | 2.5741 | 2.9126 | H9 | 4.7273 | 3.4879 | 2.1627 | 1.0965 | 3.8827 | 5.5970 | 4.9740 | 4.8748 | | 1.7784 | 1.7796 | 2.5066 | 2.5166 | H10 | 4.2095 | 2.8237 | 2.1668 | 1.0960 | 2.8259 | 4.8958 | 4.3711 | 4.7488 | 1.7784 | | 1.7650 | 3.0868 | 2.5421 | H11 | 4.2538 | 2.8048 | 2.1656 | 1.0955 | 2.7320 | 4.8540 | 4.8082 | 4.5386 | 1.7796 | 1.7650 | | 2.5390 | 3.0891 | H12 | 2.8145 | 2.1155 | 1.1020 | 2.1731 | 3.0517 | 3.7323 | 3.3280 | 2.5741 | 2.5066 | 3.0868 | 2.5390 | | 1.7458 | H13 | 2.7325 | 2.1276 | 1.1012 | 2.1782 | 3.1244 | 3.7704 | 2.6504 | 2.9126 | 2.5166 | 2.5421 | 3.0891 | 1.7458 | |
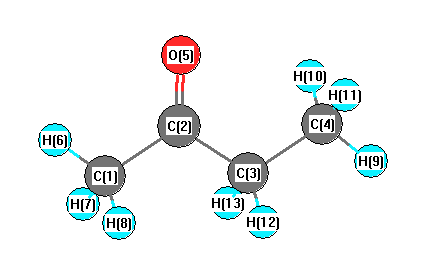 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
116.539 |
|
C1 |
C2 |
O5 |
121.556 |
| C2 |
C1 |
H6 |
110.035 |
|
C2 |
C1 |
H7 |
109.155 |
| C2 |
C1 |
H8 |
111.136 |
|
C2 |
C3 |
C4 |
114.159 |
| C2 |
C3 |
H12 |
106.859 |
|
C2 |
C3 |
H13 |
107.827 |
| C3 |
C2 |
O5 |
121.904 |
|
C3 |
C4 |
H9 |
110.602 |
| C3 |
C4 |
H10 |
110.959 |
|
C3 |
C4 |
H11 |
110.892 |
| C4 |
C3 |
H12 |
111.092 |
|
C4 |
C3 |
H13 |
111.550 |
| H6 |
C1 |
H7 |
109.140 |
|
H6 |
C1 |
H8 |
110.365 |
| H7 |
C1 |
H8 |
106.935 |
|
H9 |
C4 |
H10 |
108.414 |
| H9 |
C4 |
H11 |
108.565 |
|
H10 |
C4 |
H11 |
107.298 |
| H12 |
C3 |
H13 |
104.814 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.381 |
|
|
|
| 2 |
C |
0.170 |
|
|
|
| 3 |
C |
0.332 |
|
|
|
| 4 |
C |
0.462 |
|
|
|
| 5 |
O |
-0.611 |
|
|
|
| 6 |
H |
-0.143 |
|
|
|
| 7 |
H |
0.013 |
|
|
|
| 8 |
H |
-0.033 |
|
|
|
| 9 |
H |
-0.117 |
|
|
|
| 10 |
H |
-0.105 |
|
|
|
| 11 |
H |
-0.092 |
|
|
|
| 12 |
H |
-0.110 |
|
|
|
| 13 |
H |
-0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.179 |
-2.952 |
0.025 |
2.958 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.010 |
-1.194 |
0.118 |
| y |
-1.194 |
-35.511 |
0.031 |
| z |
0.118 |
0.031 |
-30.614 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.053 |
-1.194 |
0.118 |
| y |
-1.194 |
-5.199 |
0.031 |
| z |
0.118 |
0.031 |
2.146 |
|
| Polar |
|---|
| 3z2-r2 | 4.292 |
|---|
| x2-y2 | 5.501 |
|---|
| xy | -1.194 |
|---|
| xz | 0.118 |
|---|
| yz | 0.031 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.165 |
-0.203 |
0.039 |
| y |
-0.203 |
8.318 |
0.005 |
| z |
0.039 |
0.005 |
6.497 |
<r2> (average value of r
2) Å
2
| <r2> |
136.777 |
| (<r2>)1/2 |
11.695 |
