return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (Methyl glyoxal)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-266.541530
Energy at 298.15K-266.545557
HF Energy-265.672915
Nuclear repulsion energy157.916383
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3188 3114        
2 A' 3071 3000        
3 A' 2978 2910        
4 A' 1747 1707        
5 A' 1723 1684        
6 A' 1506 1472        
7 A' 1434 1401        
8 A' 1363 1332        
9 A' 1282 1252        
10 A' 1028 1004        
11 A' 798 780        
12 A' 570 556        
13 A' 477 466        
14 A' 248 242        
15 A" 3146 3073        
16 A" 1509 1474        
17 A" 1075 1050        
18 A" 913 892        
19 A" 462 451        
20 A" 135 132        
21 A" 107 105        

Unscaled Zero Point Vibrational Energy (zpe) 14379.4 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 14047.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.29795 0.14430 0.09903

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.848 -0.733 0.000
C2 0.000 0.556 0.000
C3 1.505 0.396 0.000
O4 -0.354 -1.859 0.000
O5 -0.597 1.637 0.000
H6 -1.950 -0.549 0.000
H7 1.984 1.386 0.000
H8 1.814 -0.187 0.887
H9 1.814 -0.187 -0.887

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.54322.61001.22852.38331.11813.53742.85842.8584
C21.54321.51402.44051.23452.24162.15102.15172.1517
C32.61001.51402.92232.44143.58261.10041.10531.1053
O41.22852.44052.92233.50372.06483.99982.87802.8780
O52.38331.23452.44143.50372.57062.59343.15053.1505
H61.11812.24163.58262.06482.57064.38483.88453.8845
H73.53742.15101.10043.99982.59344.38481.81421.8142
H82.85842.15171.10532.87803.15053.88451.81421.7743
H92.85842.15171.10532.87803.15053.88451.81421.7743

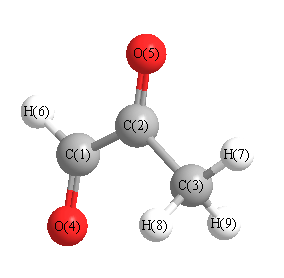
picture of Methyl glyoxal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 117.231 C1 C2 O5 117.763
C2 C1 O4 123.005 C2 C1 H6 113.812
C2 C3 H7 109.708 C2 C3 H8 109.481
C2 C3 H9 109.481 C3 C2 O5 125.006
O4 C1 H6 123.183 H7 C3 H8 110.675
H7 C3 H9 110.675 H8 C3 H9 106.770
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability