Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -1073.052386 |
| Energy at 298.15K | |
| HF Energy | -1073.052386 |
| Nuclear repulsion energy | 266.117213 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3129 |
2998 |
5.43 |
80.55 |
0.15 |
0.26 |
| 2 |
A |
2976 |
2851 |
45.62 |
135.80 |
0.28 |
0.43 |
| 3 |
A |
1859 |
1781 |
150.73 |
20.19 |
0.39 |
0.56 |
| 4 |
A |
1384 |
1326 |
12.33 |
2.61 |
0.35 |
0.52 |
| 5 |
A |
1254 |
1202 |
10.86 |
2.07 |
0.74 |
0.85 |
| 6 |
A |
1200 |
1150 |
9.98 |
4.81 |
0.68 |
0.81 |
| 7 |
A |
1038 |
995 |
14.62 |
1.43 |
0.40 |
0.57 |
| 8 |
A |
940 |
901 |
6.40 |
3.47 |
0.52 |
0.68 |
| 9 |
A |
820 |
785 |
80.59 |
6.00 |
0.69 |
0.82 |
| 10 |
A |
654 |
627 |
37.10 |
13.27 |
0.07 |
0.14 |
| 11 |
A |
616 |
591 |
30.90 |
6.14 |
0.38 |
0.55 |
| 12 |
A |
342 |
328 |
1.68 |
3.23 |
0.13 |
0.23 |
| 13 |
A |
275 |
264 |
3.65 |
3.22 |
0.62 |
0.77 |
| 14 |
A |
215 |
206 |
3.01 |
1.01 |
0.64 |
0.78 |
| 15 |
A |
88 |
85 |
10.39 |
1.35 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 8394.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 8044.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.094 |
-0.023 |
0.519 |
| C2 |
-0.708 |
-1.282 |
0.197 |
| H3 |
0.225 |
0.062 |
1.604 |
| Cl4 |
1.732 |
-0.258 |
-0.168 |
| Cl5 |
-0.676 |
1.451 |
-0.058 |
| O6 |
-1.788 |
-1.286 |
-0.319 |
| H7 |
-0.181 |
-2.213 |
0.500 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5270 | 1.0960 | 1.7920 | 1.7597 | 2.4167 | 2.2070 |
C2 | 1.5270 | | 2.1584 | 2.6711 | 2.7445 | 1.1968 | 1.1120 | H3 | 1.0960 | 2.1584 | | 2.3480 | 2.3457 | 3.0936 | 2.5610 | Cl4 | 1.7920 | 2.6711 | 2.3480 | | 2.9539 | 3.6702 | 2.8155 | Cl5 | 1.7597 | 2.7445 | 2.3457 | 2.9539 | | 2.9660 | 3.7385 | O6 | 2.4167 | 1.1968 | 3.0936 | 3.6702 | 2.9660 | | 2.0277 | H7 | 2.2070 | 1.1120 | 2.5610 | 2.8155 | 3.7385 | 2.0277 | |
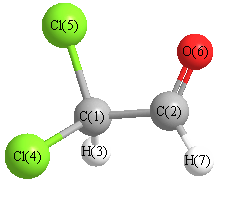 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.612 |
|
C1 |
C2 |
H7 |
112.553 |
| C2 |
C1 |
H3 |
109.646 |
|
C2 |
C1 |
Cl4 |
106.912 |
| C2 |
C1 |
Cl5 |
113.045 |
|
H3 |
C1 |
Cl4 |
106.269 |
| H3 |
C1 |
Cl5 |
108.194 |
|
Cl4 |
C1 |
Cl5 |
112.546 |
| O6 |
C2 |
H7 |
122.833 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.724 |
|
|
|
| 2 |
C |
1.036 |
|
|
|
| 3 |
H |
0.135 |
|
|
|
| 4 |
Cl |
0.056 |
|
|
|
| 5 |
Cl |
0.059 |
|
|
|
| 6 |
O |
-0.437 |
|
|
|
| 7 |
H |
-0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.345 |
-0.724 |
1.955 |
2.481 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.734 |
-2.225 |
-1.368 |
| y |
-2.225 |
-41.679 |
-1.671 |
| z |
-1.368 |
-1.671 |
-40.328 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.730 |
-2.225 |
-1.368 |
| y |
-2.225 |
2.352 |
-1.671 |
| z |
-1.368 |
-1.671 |
4.378 |
|
| Polar |
|---|
| 3z2-r2 | 8.756 |
|---|
| x2-y2 | -6.054 |
|---|
| xy | -2.225 |
|---|
| xz | -1.368 |
|---|
| yz | -1.671 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.670 |
-0.531 |
0.218 |
| y |
-0.531 |
8.666 |
-0.216 |
| z |
0.218 |
-0.216 |
6.466 |
<r2> (average value of r
2) Å
2
| <r2> |
185.405 |
| (<r2>)1/2 |
13.616 |
