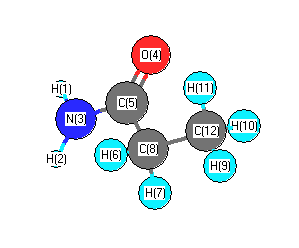
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3961 |
3607 |
50.59 |
|
|
|
| 2 |
A |
3820 |
3478 |
51.14 |
|
|
|
| 3 |
A |
3271 |
2979 |
35.48 |
|
|
|
| 4 |
A |
3251 |
2961 |
35.04 |
|
|
|
| 5 |
A |
3207 |
2920 |
14.99 |
|
|
|
| 6 |
A |
3184 |
2900 |
25.13 |
|
|
|
| 7 |
A |
3170 |
2887 |
36.12 |
|
|
|
| 8 |
A |
1923 |
1751 |
413.21 |
|
|
|
| 9 |
A |
1761 |
1604 |
121.58 |
|
|
|
| 10 |
A |
1598 |
1455 |
13.81 |
|
|
|
| 11 |
A |
1586 |
1444 |
5.91 |
|
|
|
| 12 |
A |
1572 |
1431 |
5.10 |
|
|
|
| 13 |
A |
1543 |
1405 |
35.89 |
|
|
|
| 14 |
A |
1514 |
1379 |
38.05 |
|
|
|
| 15 |
A |
1388 |
1264 |
0.03 |
|
|
|
| 16 |
A |
1375 |
1252 |
148.02 |
|
|
|
| 17 |
A |
1213 |
1105 |
2.78 |
|
|
|
| 18 |
A |
1195 |
1088 |
0.50 |
|
|
|
| 19 |
A |
1143 |
1041 |
9.39 |
|
|
|
| 20 |
A |
1072 |
977 |
2.79 |
|
|
|
| 21 |
A |
876 |
798 |
8.69 |
|
|
|
| 22 |
A |
855 |
778 |
1.33 |
|
|
|
| 23 |
A |
661 |
602 |
9.05 |
|
|
|
| 24 |
A |
648 |
590 |
17.60 |
|
|
|
| 25 |
A |
534 |
486 |
5.35 |
|
|
|
| 26 |
A |
474 |
431 |
4.65 |
|
|
|
| 27 |
A |
271 |
247 |
10.44 |
|
|
|
| 28 |
A |
236 |
215 |
0.12 |
|
|
|
| 29 |
A |
188 |
172 |
228.93 |
|
|
|
| 30 |
A |
31 |
28 |
0.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23761.2 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 21636.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.046 |
|
|
|
| 2 |
H |
-0.048 |
|
|
|
| 3 |
N |
-0.176 |
|
|
|
| 4 |
O |
-0.738 |
|
|
|
| 5 |
C |
0.682 |
|
|
|
| 6 |
H |
-0.032 |
|
|
|
| 7 |
H |
-0.043 |
|
|
|
| 8 |
C |
0.117 |
|
|
|
| 9 |
H |
-0.058 |
|
|
|
| 10 |
H |
-0.058 |
|
|
|
| 11 |
H |
-0.044 |
|
|
|
| 12 |
C |
0.446 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.645 |
-3.835 |
0.304 |
3.901 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.088 |
2.440 |
-0.653 |
| y |
2.440 |
-33.860 |
-0.396 |
| z |
-0.653 |
-0.396 |
-31.670 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.678 |
2.440 |
-0.653 |
| y |
2.440 |
-4.981 |
-0.396 |
| z |
-0.653 |
-0.396 |
-1.697 |
|
| Polar |
|---|
| 3z2-r2 | -3.393 |
|---|
| x2-y2 | 7.772 |
|---|
| xy | 2.440 |
|---|
| xz | -0.653 |
|---|
| yz | -0.396 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
128.021 |
| (<r2>)1/2 |
11.315 |