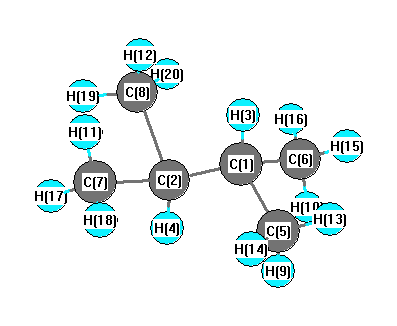
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3150 |
3019 |
0.00 |
|
|
|
| 2 |
Ag |
3125 |
2994 |
0.00 |
|
|
|
| 3 |
Ag |
3050 |
2923 |
0.00 |
|
|
|
| 4 |
Ag |
3010 |
2884 |
0.00 |
|
|
|
| 5 |
Ag |
1506 |
1443 |
0.00 |
|
|
|
| 6 |
Ag |
1486 |
1424 |
0.00 |
|
|
|
| 7 |
Ag |
1405 |
1346 |
0.00 |
|
|
|
| 8 |
Ag |
1368 |
1311 |
0.00 |
|
|
|
| 9 |
Ag |
1211 |
1161 |
0.00 |
|
|
|
| 10 |
Ag |
1188 |
1139 |
0.00 |
|
|
|
| 11 |
Ag |
961 |
921 |
0.00 |
|
|
|
| 12 |
Ag |
767 |
735 |
0.00 |
|
|
|
| 13 |
Ag |
501 |
480 |
0.00 |
|
|
|
| 14 |
Ag |
381 |
365 |
0.00 |
|
|
|
| 15 |
Ag |
236 |
226 |
0.00 |
|
|
|
| 16 |
Au |
3150 |
3019 |
54.98 |
|
|
|
| 17 |
Au |
3120 |
2990 |
2.34 |
|
|
|
| 18 |
Au |
3045 |
2918 |
50.77 |
|
|
|
| 19 |
Au |
1484 |
1422 |
4.67 |
|
|
|
| 20 |
Au |
1461 |
1400 |
1.27 |
|
|
|
| 21 |
Au |
1387 |
1330 |
14.20 |
|
|
|
| 22 |
Au |
1328 |
1273 |
1.57 |
|
|
|
| 23 |
Au |
1105 |
1059 |
1.83 |
|
|
|
| 24 |
Au |
968 |
927 |
0.78 |
|
|
|
| 25 |
Au |
919 |
880 |
1.97 |
|
|
|
| 26 |
Au |
295 |
282 |
0.00 |
|
|
|
| 27 |
Au |
193 |
185 |
0.01 |
|
|
|
| 28 |
Au |
68 |
65 |
0.00 |
|
|
|
| 29 |
Bg |
3137 |
3007 |
0.00 |
|
|
|
| 30 |
Bg |
3120 |
2990 |
0.00 |
|
|
|
| 31 |
Bg |
3044 |
2917 |
0.00 |
|
|
|
| 32 |
Bg |
1470 |
1409 |
0.00 |
|
|
|
| 33 |
Bg |
1465 |
1404 |
0.00 |
|
|
|
| 34 |
Bg |
1387 |
1329 |
0.00 |
|
|
|
| 35 |
Bg |
1342 |
1286 |
0.00 |
|
|
|
| 36 |
Bg |
1184 |
1135 |
0.00 |
|
|
|
| 37 |
Bg |
965 |
925 |
0.00 |
|
|
|
| 38 |
Bg |
912 |
874 |
0.00 |
|
|
|
| 39 |
Bg |
430 |
412 |
0.00 |
|
|
|
| 40 |
Bg |
193 |
185 |
0.00 |
|
|
|
| 41 |
Bu |
3140 |
3009 |
76.91 |
|
|
|
| 42 |
Bu |
3126 |
2995 |
138.97 |
|
|
|
| 43 |
Bu |
3048 |
2921 |
76.16 |
|
|
|
| 44 |
Bu |
3025 |
2898 |
31.01 |
|
|
|
| 45 |
Bu |
1490 |
1427 |
23.48 |
|
|
|
| 46 |
Bu |
1482 |
1420 |
14.99 |
|
|
|
| 47 |
Bu |
1400 |
1342 |
17.26 |
|
|
|
| 48 |
Bu |
1289 |
1235 |
4.25 |
|
|
|
| 49 |
Bu |
1174 |
1125 |
7.42 |
|
|
|
| 50 |
Bu |
988 |
947 |
5.30 |
|
|
|
| 51 |
Bu |
889 |
852 |
2.29 |
|
|
|
| 52 |
Bu |
410 |
393 |
0.08 |
|
|
|
| 53 |
Bu |
350 |
335 |
0.10 |
|
|
|
| 54 |
Bu |
214 |
205 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41269.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 39548.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.407 |
|
|
|
| 2 |
C |
-0.407 |
|
|
|
| 3 |
H |
-0.467 |
|
|
|
| 4 |
H |
-0.467 |
|
|
|
| 5 |
C |
0.828 |
|
|
|
| 6 |
C |
0.828 |
|
|
|
| 7 |
C |
0.828 |
|
|
|
| 8 |
C |
0.828 |
|
|
|
| 9 |
H |
-0.067 |
|
|
|
| 10 |
H |
-0.067 |
|
|
|
| 11 |
H |
-0.067 |
|
|
|
| 12 |
H |
-0.067 |
|
|
|
| 13 |
H |
-0.157 |
|
|
|
| 14 |
H |
-0.167 |
|
|
|
| 15 |
H |
-0.157 |
|
|
|
| 16 |
H |
-0.167 |
|
|
|
| 17 |
H |
-0.157 |
|
|
|
| 18 |
H |
-0.167 |
|
|
|
| 19 |
H |
-0.157 |
|
|
|
| 20 |
H |
-0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.620 |
-0.487 |
0.000 |
| y |
-0.487 |
-42.336 |
0.000 |
| z |
0.000 |
0.000 |
-42.675 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.885 |
-0.487 |
0.000 |
| y |
-0.487 |
-0.188 |
0.000 |
| z |
0.000 |
0.000 |
-0.697 |
|
| Polar |
|---|
| 3z2-r2 | -1.395 |
|---|
| x2-y2 | 0.715 |
|---|
| xy | -0.487 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.100 |
0.335 |
0.000 |
| y |
0.335 |
12.308 |
0.000 |
| z |
0.000 |
0.000 |
11.839 |
<r2> (average value of r
2) Å
2
| <r2> |
207.763 |
| (<r2>)1/2 |
14.414 |