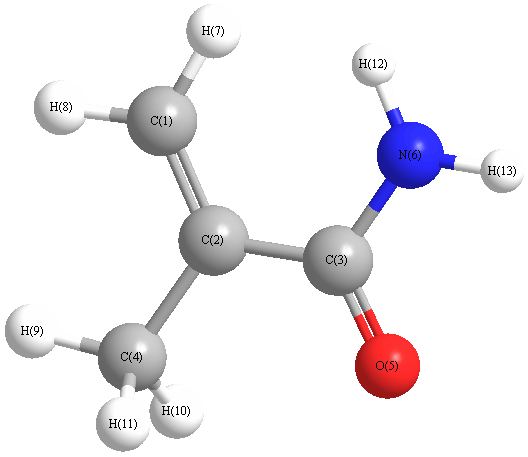
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3950 |
3597 |
46.02 |
|
|
|
| 2 |
A |
3813 |
3473 |
48.84 |
|
|
|
| 3 |
A |
3391 |
3088 |
10.08 |
|
|
|
| 4 |
A |
3305 |
3010 |
6.05 |
|
|
|
| 5 |
A |
3271 |
2978 |
15.96 |
|
|
|
| 6 |
A |
3247 |
2957 |
21.68 |
|
|
|
| 7 |
A |
3181 |
2896 |
23.25 |
|
|
|
| 8 |
A |
1921 |
1749 |
402.19 |
|
|
|
| 9 |
A |
1836 |
1672 |
91.15 |
|
|
|
| 10 |
A |
1759 |
1602 |
116.24 |
|
|
|
| 11 |
A |
1594 |
1451 |
20.26 |
|
|
|
| 12 |
A |
1572 |
1432 |
8.79 |
|
|
|
| 13 |
A |
1554 |
1415 |
12.09 |
|
|
|
| 14 |
A |
1532 |
1395 |
22.20 |
|
|
|
| 15 |
A |
1491 |
1357 |
82.81 |
|
|
|
| 16 |
A |
1326 |
1207 |
62.03 |
|
|
|
| 17 |
A |
1197 |
1090 |
8.96 |
|
|
|
| 18 |
A |
1157 |
1053 |
0.59 |
|
|
|
| 19 |
A |
1101 |
1003 |
9.59 |
|
|
|
| 20 |
A |
1075 |
979 |
40.10 |
|
|
|
| 21 |
A |
1018 |
927 |
2.89 |
|
|
|
| 22 |
A |
901 |
821 |
25.19 |
|
|
|
| 23 |
A |
803 |
731 |
5.20 |
|
|
|
| 24 |
A |
738 |
672 |
12.94 |
|
|
|
| 25 |
A |
622 |
566 |
19.48 |
|
|
|
| 26 |
A |
585 |
533 |
21.77 |
|
|
|
| 27 |
A |
546 |
498 |
5.13 |
|
|
|
| 28 |
A |
409 |
372 |
44.94 |
|
|
|
| 29 |
A |
383 |
349 |
150.98 |
|
|
|
| 30 |
A |
353 |
321 |
5.48 |
|
|
|
| 31 |
A |
283 |
257 |
13.62 |
|
|
|
| 32 |
A |
197 |
180 |
0.21 |
|
|
|
| 33 |
A |
70 |
63 |
4.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25089.0 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 22846.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.181 |
|
|
|
| 2 |
H |
-0.016 |
|
|
|
| 3 |
H |
-0.028 |
|
|
|
| 4 |
C |
0.238 |
|
|
|
| 5 |
H |
-0.128 |
|
|
|
| 6 |
H |
0.076 |
|
|
|
| 7 |
H |
0.084 |
|
|
|
| 8 |
C |
1.069 |
|
|
|
| 9 |
H |
-0.407 |
|
|
|
| 10 |
H |
-0.386 |
|
|
|
| 11 |
C |
0.591 |
|
|
|
| 12 |
O |
-0.809 |
|
|
|
| 13 |
C |
-0.102 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.627 |
3.223 |
1.594 |
3.650 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.036 |
-4.160 |
-2.759 |
| y |
-4.160 |
-38.329 |
-0.435 |
| z |
-2.759 |
-0.435 |
-37.875 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.065 |
-4.160 |
-2.759 |
| y |
-4.160 |
-3.373 |
-0.435 |
| z |
-2.759 |
-0.435 |
-2.692 |
|
| Polar |
|---|
| 3z2-r2 | -5.384 |
|---|
| x2-y2 | 6.292 |
|---|
| xy | -4.160 |
|---|
| xz | -2.759 |
|---|
| yz | -0.435 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
163.314 |
| (<r2>)1/2 |
12.779 |