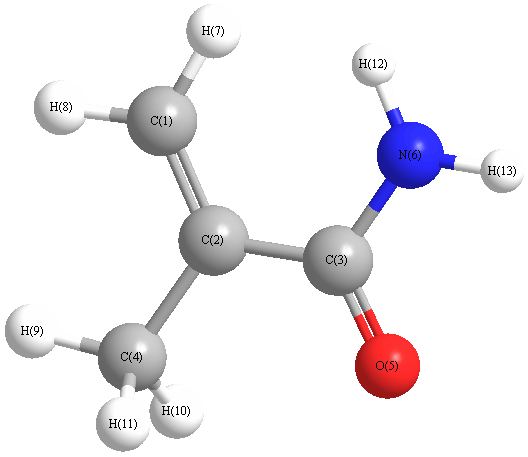
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3764 |
3607 |
38.70 |
|
|
|
| 2 |
A |
3623 |
3472 |
35.77 |
|
|
|
| 3 |
A |
3264 |
3128 |
8.78 |
|
|
|
| 4 |
A |
3171 |
3038 |
5.27 |
|
|
|
| 5 |
A |
3169 |
3037 |
11.44 |
|
|
|
| 6 |
A |
3136 |
3005 |
8.87 |
|
|
|
| 7 |
A |
3063 |
2935 |
14.29 |
|
|
|
| 8 |
A |
1776 |
1702 |
295.69 |
|
|
|
| 9 |
A |
1716 |
1644 |
71.17 |
|
|
|
| 10 |
A |
1614 |
1546 |
117.62 |
|
|
|
| 11 |
A |
1475 |
1413 |
25.70 |
|
|
|
| 12 |
A |
1449 |
1388 |
13.56 |
|
|
|
| 13 |
A |
1437 |
1377 |
16.64 |
|
|
|
| 14 |
A |
1416 |
1357 |
27.74 |
|
|
|
| 15 |
A |
1381 |
1323 |
42.22 |
|
|
|
| 16 |
A |
1241 |
1189 |
44.41 |
|
|
|
| 17 |
A |
1095 |
1049 |
1.07 |
|
|
|
| 18 |
A |
1058 |
1014 |
0.49 |
|
|
|
| 19 |
A |
1015 |
972 |
7.55 |
|
|
|
| 20 |
A |
963 |
923 |
8.10 |
|
|
|
| 21 |
A |
955 |
916 |
31.99 |
|
|
|
| 22 |
A |
824 |
790 |
11.79 |
|
|
|
| 23 |
A |
759 |
728 |
3.78 |
|
|
|
| 24 |
A |
676 |
648 |
7.43 |
|
|
|
| 25 |
A |
579 |
554 |
13.86 |
|
|
|
| 26 |
A |
567 |
544 |
5.91 |
|
|
|
| 27 |
A |
506 |
485 |
3.60 |
|
|
|
| 28 |
A |
377 |
362 |
8.26 |
|
|
|
| 29 |
A |
326 |
312 |
30.73 |
|
|
|
| 30 |
A |
315 |
302 |
129.35 |
|
|
|
| 31 |
A |
259 |
248 |
22.95 |
|
|
|
| 32 |
A |
173 |
166 |
0.30 |
|
|
|
| 33 |
A |
69 |
66 |
4.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23603.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 22619.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.054 |
|
|
|
| 2 |
H |
-0.028 |
|
|
|
| 3 |
H |
-0.048 |
|
|
|
| 4 |
C |
0.141 |
|
|
|
| 5 |
H |
0.043 |
|
|
|
| 6 |
H |
0.029 |
|
|
|
| 7 |
H |
-0.193 |
|
|
|
| 8 |
C |
0.870 |
|
|
|
| 9 |
H |
-0.546 |
|
|
|
| 10 |
H |
-0.526 |
|
|
|
| 11 |
C |
0.525 |
|
|
|
| 12 |
O |
-0.658 |
|
|
|
| 13 |
C |
0.443 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.456 |
3.116 |
1.431 |
3.459 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.564 |
-4.595 |
-2.391 |
| y |
-4.595 |
-37.673 |
-0.422 |
| z |
-2.391 |
-0.422 |
-37.705 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.125 |
-4.595 |
-2.391 |
| y |
-4.595 |
-3.038 |
-0.422 |
| z |
-2.391 |
-0.422 |
-3.086 |
|
| Polar |
|---|
| 3z2-r2 | -6.173 |
|---|
| x2-y2 | 6.109 |
|---|
| xy | -4.595 |
|---|
| xz | -2.391 |
|---|
| yz | -0.422 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.055 |
0.337 |
-0.186 |
| y |
0.337 |
11.102 |
-0.105 |
| z |
-0.186 |
-0.105 |
6.814 |
<r2> (average value of r
2) Å
2
| <r2> |
163.618 |
| (<r2>)1/2 |
12.791 |